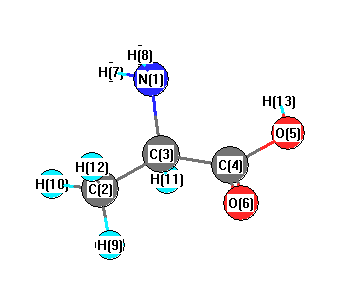
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3490 |
3433 |
3.69 |
|
|
|
| 2 |
A |
3396 |
3340 |
0.22 |
|
|
|
| 3 |
A |
3120 |
3069 |
302.63 |
|
|
|
| 4 |
A |
3094 |
3043 |
1.19 |
|
|
|
| 5 |
A |
3061 |
3010 |
25.74 |
|
|
|
| 6 |
A |
2990 |
2940 |
25.67 |
|
|
|
| 7 |
A |
2984 |
2935 |
2.20 |
|
|
|
| 8 |
A |
1819 |
1789 |
254.72 |
|
|
|
| 9 |
A |
1644 |
1617 |
31.94 |
|
|
|
| 10 |
A |
1481 |
1456 |
15.27 |
|
|
|
| 11 |
A |
1471 |
1447 |
5.83 |
|
|
|
| 12 |
A |
1443 |
1419 |
373.50 |
|
|
|
| 13 |
A |
1379 |
1356 |
12.16 |
|
|
|
| 14 |
A |
1359 |
1336 |
7.25 |
|
|
|
| 15 |
A |
1284 |
1263 |
5.59 |
|
|
|
| 16 |
A |
1212 |
1192 |
11.81 |
|
|
|
| 17 |
A |
1190 |
1170 |
13.39 |
|
|
|
| 18 |
A |
1124 |
1106 |
7.04 |
|
|
|
| 19 |
A |
1076 |
1058 |
44.89 |
|
|
|
| 20 |
A |
998 |
982 |
3.39 |
|
|
|
| 21 |
A |
942 |
926 |
81.53 |
|
|
|
| 22 |
A |
914 |
899 |
45.66 |
|
|
|
| 23 |
A |
860 |
845 |
84.80 |
|
|
|
| 24 |
A |
780 |
767 |
5.79 |
|
|
|
| 25 |
A |
709 |
698 |
5.59 |
|
|
|
| 26 |
A |
555 |
546 |
3.40 |
|
|
|
| 27 |
A |
513 |
504 |
0.50 |
|
|
|
| 28 |
A |
402 |
395 |
5.74 |
|
|
|
| 29 |
A |
362 |
356 |
19.49 |
|
|
|
| 30 |
A |
307 |
302 |
17.24 |
|
|
|
| 31 |
A |
267 |
262 |
12.27 |
|
|
|
| 32 |
A |
225 |
222 |
0.33 |
|
|
|
| 33 |
A |
79 |
77 |
1.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23262.8 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 22878.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.749 |
|
|
|
| 2 |
C |
-0.490 |
|
|
|
| 3 |
C |
-0.129 |
|
|
|
| 4 |
C |
0.565 |
|
|
|
| 5 |
O |
-0.546 |
|
|
|
| 6 |
O |
-0.437 |
|
|
|
| 7 |
H |
0.325 |
|
|
|
| 8 |
H |
0.327 |
|
|
|
| 9 |
H |
0.202 |
|
|
|
| 10 |
H |
0.164 |
|
|
|
| 11 |
H |
0.193 |
|
|
|
| 12 |
H |
0.156 |
|
|
|
| 13 |
H |
0.419 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.946 |
2.052 |
0.266 |
5.361 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.297 |
0.235 |
-0.350 |
| y |
0.235 |
-36.970 |
-0.069 |
| z |
-0.350 |
-0.069 |
-33.327 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.148 |
0.235 |
-0.350 |
| y |
0.235 |
-1.158 |
-0.069 |
| z |
-0.350 |
-0.069 |
4.306 |
|
| Polar |
|---|
| 3z2-r2 | 8.613 |
|---|
| x2-y2 | -1.327 |
|---|
| xy | 0.235 |
|---|
| xz | -0.350 |
|---|
| yz | -0.069 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.177 |
0.285 |
0.131 |
| y |
0.285 |
7.551 |
-0.101 |
| z |
0.131 |
-0.101 |
5.239 |
<r2> (average value of r
2) Å
2
| <r2> |
163.149 |
| (<r2>)1/2 |
12.773 |