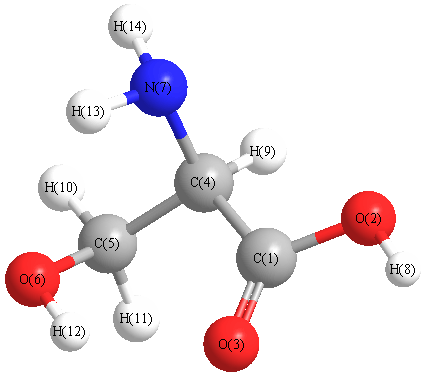
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3711 |
3550 |
4.44 |
|
|
|
| 2 |
A |
3705 |
3545 |
152.70 |
|
|
|
| 3 |
A |
3597 |
3441 |
2.69 |
|
|
|
| 4 |
A |
3505 |
3354 |
0.63 |
|
|
|
| 5 |
A |
3138 |
3003 |
23.52 |
|
|
|
| 6 |
A |
3061 |
2928 |
13.17 |
|
|
|
| 7 |
A |
2982 |
2853 |
68.62 |
|
|
|
| 8 |
A |
1824 |
1745 |
239.68 |
|
|
|
| 9 |
A |
1703 |
1630 |
35.71 |
|
|
|
| 10 |
A |
1534 |
1467 |
1.29 |
|
|
|
| 11 |
A |
1466 |
1402 |
55.89 |
|
|
|
| 12 |
A |
1406 |
1346 |
11.74 |
|
|
|
| 13 |
A |
1401 |
1340 |
26.21 |
|
|
|
| 14 |
A |
1389 |
1329 |
57.33 |
|
|
|
| 15 |
A |
1334 |
1276 |
11.17 |
|
|
|
| 16 |
A |
1249 |
1195 |
7.59 |
|
|
|
| 17 |
A |
1215 |
1163 |
38.57 |
|
|
|
| 18 |
A |
1197 |
1145 |
174.65 |
|
|
|
| 19 |
A |
1162 |
1112 |
38.76 |
|
|
|
| 20 |
A |
1123 |
1075 |
111.80 |
|
|
|
| 21 |
A |
1038 |
993 |
41.85 |
|
|
|
| 22 |
A |
1003 |
959 |
1.62 |
|
|
|
| 23 |
A |
896 |
857 |
195.03 |
|
|
|
| 24 |
A |
816 |
781 |
32.29 |
|
|
|
| 25 |
A |
745 |
713 |
42.46 |
|
|
|
| 26 |
A |
660 |
631 |
89.17 |
|
|
|
| 27 |
A |
593 |
567 |
185.95 |
|
|
|
| 28 |
A |
567 |
543 |
29.17 |
|
|
|
| 29 |
A |
534 |
511 |
31.46 |
|
|
|
| 30 |
A |
424 |
405 |
8.39 |
|
|
|
| 31 |
A |
315 |
302 |
5.05 |
|
|
|
| 32 |
A |
299 |
286 |
9.40 |
|
|
|
| 33 |
A |
253 |
242 |
30.51 |
|
|
|
| 34 |
A |
230 |
220 |
13.15 |
|
|
|
| 35 |
A |
166 |
159 |
1.98 |
|
|
|
| 36 |
A |
28 |
26 |
0.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25134.3 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 24046.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.595 |
|
|
|
| 2 |
O |
-0.578 |
|
|
|
| 3 |
O |
-0.487 |
|
|
|
| 4 |
C |
-0.152 |
|
|
|
| 5 |
C |
-0.067 |
|
|
|
| 6 |
O |
-0.639 |
|
|
|
| 7 |
N |
-0.732 |
|
|
|
| 8 |
H |
0.428 |
|
|
|
| 9 |
H |
0.219 |
|
|
|
| 10 |
H |
0.138 |
|
|
|
| 11 |
H |
0.192 |
|
|
|
| 12 |
H |
0.428 |
|
|
|
| 13 |
H |
0.335 |
|
|
|
| 14 |
H |
0.320 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.635 |
0.285 |
0.722 |
3.717 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.276 |
-2.292 |
-0.806 |
| y |
-2.292 |
-44.101 |
0.951 |
| z |
-0.806 |
0.951 |
-38.740 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.145 |
-2.292 |
-0.806 |
| y |
-2.292 |
-4.093 |
0.951 |
| z |
-0.806 |
0.951 |
3.948 |
|
| Polar |
|---|
| 3z2-r2 | 7.896 |
|---|
| x2-y2 | 2.825 |
|---|
| xy | -2.292 |
|---|
| xz | -0.806 |
|---|
| yz | 0.951 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.767 |
0.211 |
0.002 |
| y |
0.211 |
7.477 |
-0.039 |
| z |
0.002 |
-0.039 |
5.428 |
<r2> (average value of r
2) Å
2
| <r2> |
215.930 |
| (<r2>)1/2 |
14.695 |