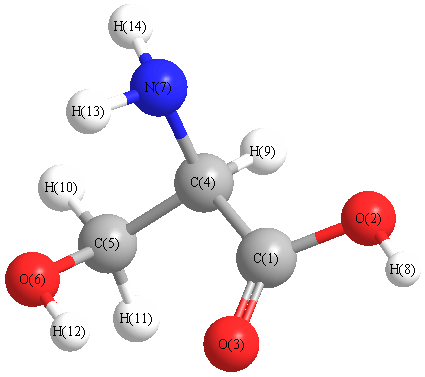
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3730 |
3548 |
17.60 |
|
|
|
| 2 |
A |
3723 |
3541 |
152.55 |
|
|
|
| 3 |
A |
3614 |
3437 |
3.24 |
|
|
|
| 4 |
A |
3522 |
3349 |
0.81 |
|
|
|
| 5 |
A |
3151 |
2996 |
22.83 |
|
|
|
| 6 |
A |
3074 |
2923 |
12.57 |
|
|
|
| 7 |
A |
2993 |
2846 |
67.78 |
|
|
|
| 8 |
A |
1842 |
1751 |
244.87 |
|
|
|
| 9 |
A |
1708 |
1625 |
37.75 |
|
|
|
| 10 |
A |
1537 |
1462 |
1.40 |
|
|
|
| 11 |
A |
1475 |
1403 |
56.06 |
|
|
|
| 12 |
A |
1413 |
1344 |
29.47 |
|
|
|
| 13 |
A |
1408 |
1339 |
51.84 |
|
|
|
| 14 |
A |
1400 |
1331 |
25.25 |
|
|
|
| 15 |
A |
1340 |
1274 |
11.32 |
|
|
|
| 16 |
A |
1255 |
1194 |
8.47 |
|
|
|
| 17 |
A |
1225 |
1165 |
51.88 |
|
|
|
| 18 |
A |
1208 |
1149 |
161.20 |
|
|
|
| 19 |
A |
1173 |
1115 |
35.22 |
|
|
|
| 20 |
A |
1137 |
1081 |
108.25 |
|
|
|
| 21 |
A |
1046 |
995 |
42.62 |
|
|
|
| 22 |
A |
1010 |
961 |
0.95 |
|
|
|
| 23 |
A |
905 |
860 |
194.57 |
|
|
|
| 24 |
A |
823 |
782 |
33.14 |
|
|
|
| 25 |
A |
752 |
715 |
44.00 |
|
|
|
| 26 |
A |
664 |
632 |
88.94 |
|
|
|
| 27 |
A |
600 |
571 |
187.94 |
|
|
|
| 28 |
A |
573 |
545 |
25.14 |
|
|
|
| 29 |
A |
540 |
514 |
33.95 |
|
|
|
| 30 |
A |
426 |
406 |
8.23 |
|
|
|
| 31 |
A |
317 |
301 |
5.17 |
|
|
|
| 32 |
A |
302 |
287 |
9.57 |
|
|
|
| 33 |
A |
254 |
241 |
25.14 |
|
|
|
| 34 |
A |
235 |
223 |
18.95 |
|
|
|
| 35 |
A |
170 |
161 |
1.60 |
|
|
|
| 36 |
A |
30 |
29 |
0.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25287.9 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 24048.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.598 |
|
|
|
| 2 |
O |
-0.580 |
|
|
|
| 3 |
O |
-0.487 |
|
|
|
| 4 |
C |
-0.159 |
|
|
|
| 5 |
C |
-0.069 |
|
|
|
| 6 |
O |
-0.641 |
|
|
|
| 7 |
N |
-0.734 |
|
|
|
| 8 |
H |
0.431 |
|
|
|
| 9 |
H |
0.221 |
|
|
|
| 10 |
H |
0.139 |
|
|
|
| 11 |
H |
0.194 |
|
|
|
| 12 |
H |
0.431 |
|
|
|
| 13 |
H |
0.337 |
|
|
|
| 14 |
H |
0.321 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.658 |
0.263 |
0.732 |
3.739 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.360 |
-2.374 |
-0.754 |
| y |
-2.374 |
-44.254 |
0.961 |
| z |
-0.754 |
0.961 |
-38.814 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.174 |
-2.374 |
-0.754 |
| y |
-2.374 |
-4.167 |
0.961 |
| z |
-0.754 |
0.961 |
3.993 |
|
| Polar |
|---|
| 3z2-r2 | 7.986 |
|---|
| x2-y2 | 2.894 |
|---|
| xy | -2.374 |
|---|
| xz | -0.754 |
|---|
| yz | 0.961 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.718 |
0.209 |
0.013 |
| y |
0.209 |
7.419 |
-0.029 |
| z |
0.013 |
-0.029 |
5.408 |
<r2> (average value of r
2) Å
2
| <r2> |
214.784 |
| (<r2>)1/2 |
14.656 |