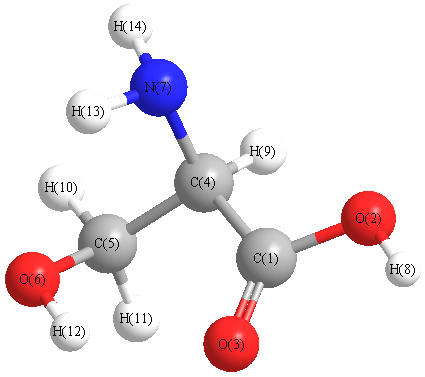
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3546 |
3488 |
24.15 |
|
|
|
| 2 |
A |
3518 |
3460 |
112.02 |
|
|
|
| 3 |
A |
3475 |
3418 |
0.70 |
|
|
|
| 4 |
A |
3386 |
3330 |
0.24 |
|
|
|
| 5 |
A |
3052 |
3002 |
23.74 |
|
|
|
| 6 |
A |
2978 |
2929 |
12.28 |
|
|
|
| 7 |
A |
2879 |
2832 |
76.99 |
|
|
|
| 8 |
A |
1745 |
1716 |
201.64 |
|
|
|
| 9 |
A |
1655 |
1628 |
30.35 |
|
|
|
| 10 |
A |
1488 |
1463 |
1.07 |
|
|
|
| 11 |
A |
1437 |
1413 |
75.00 |
|
|
|
| 12 |
A |
1359 |
1337 |
14.23 |
|
|
|
| 13 |
A |
1356 |
1334 |
4.55 |
|
|
|
| 14 |
A |
1333 |
1311 |
48.08 |
|
|
|
| 15 |
A |
1286 |
1265 |
10.90 |
|
|
|
| 16 |
A |
1208 |
1188 |
5.71 |
|
|
|
| 17 |
A |
1177 |
1158 |
11.07 |
|
|
|
| 18 |
A |
1153 |
1134 |
141.14 |
|
|
|
| 19 |
A |
1126 |
1108 |
83.76 |
|
|
|
| 20 |
A |
1080 |
1062 |
109.33 |
|
|
|
| 21 |
A |
1006 |
990 |
37.77 |
|
|
|
| 22 |
A |
966 |
950 |
3.74 |
|
|
|
| 23 |
A |
879 |
865 |
179.58 |
|
|
|
| 24 |
A |
785 |
772 |
27.56 |
|
|
|
| 25 |
A |
722 |
710 |
47.01 |
|
|
|
| 26 |
A |
650 |
639 |
63.70 |
|
|
|
| 27 |
A |
596 |
586 |
185.47 |
|
|
|
| 28 |
A |
550 |
541 |
3.81 |
|
|
|
| 29 |
A |
523 |
515 |
43.51 |
|
|
|
| 30 |
A |
406 |
399 |
8.64 |
|
|
|
| 31 |
A |
307 |
302 |
3.24 |
|
|
|
| 32 |
A |
295 |
290 |
9.39 |
|
|
|
| 33 |
A |
243 |
239 |
15.68 |
|
|
|
| 34 |
A |
227 |
223 |
24.88 |
|
|
|
| 35 |
A |
165 |
162 |
1.08 |
|
|
|
| 36 |
A |
33 |
33 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24294.1 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 23893.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.534 |
|
|
|
| 2 |
O |
-0.529 |
|
|
|
| 3 |
O |
-0.446 |
|
|
|
| 4 |
C |
-0.145 |
|
|
|
| 5 |
C |
-0.072 |
|
|
|
| 6 |
O |
-0.592 |
|
|
|
| 7 |
N |
-0.682 |
|
|
|
| 8 |
H |
0.405 |
|
|
|
| 9 |
H |
0.203 |
|
|
|
| 10 |
H |
0.124 |
|
|
|
| 11 |
H |
0.178 |
|
|
|
| 12 |
H |
0.403 |
|
|
|
| 13 |
H |
0.315 |
|
|
|
| 14 |
H |
0.303 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.638 |
0.193 |
0.693 |
3.709 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.490 |
-2.298 |
-0.480 |
| y |
-2.298 |
-44.341 |
0.610 |
| z |
-0.480 |
0.610 |
-38.952 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.156 |
-2.298 |
-0.480 |
| y |
-2.298 |
-4.120 |
0.610 |
| z |
-0.480 |
0.610 |
3.963 |
|
| Polar |
|---|
| 3z2-r2 | 7.926 |
|---|
| x2-y2 | 2.851 |
|---|
| xy | -2.298 |
|---|
| xz | -0.480 |
|---|
| yz | 0.610 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.393 |
0.153 |
0.098 |
| y |
0.153 |
7.860 |
0.017 |
| z |
0.098 |
0.017 |
5.583 |
<r2> (average value of r
2) Å
2
| <r2> |
218.066 |
| (<r2>)1/2 |
14.767 |