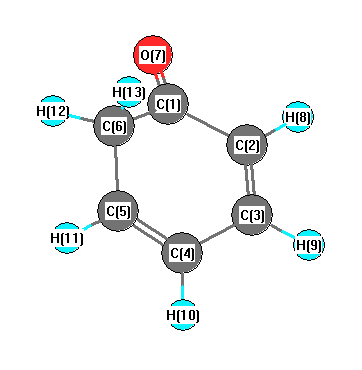
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3237 |
3078 |
0.42 |
|
|
|
| 2 |
A1 |
3200 |
3043 |
8.21 |
|
|
|
| 3 |
A1 |
3026 |
2877 |
9.56 |
|
|
|
| 4 |
A1 |
1798 |
1710 |
287.86 |
|
|
|
| 5 |
A1 |
1737 |
1652 |
0.71 |
|
|
|
| 6 |
A1 |
1454 |
1383 |
17.94 |
|
|
|
| 7 |
A1 |
1431 |
1361 |
3.71 |
|
|
|
| 8 |
A1 |
1203 |
1144 |
5.84 |
|
|
|
| 9 |
A1 |
970 |
923 |
5.49 |
|
|
|
| 10 |
A1 |
905 |
861 |
9.90 |
|
|
|
| 11 |
A1 |
789 |
750 |
1.03 |
|
|
|
| 12 |
A1 |
507 |
482 |
2.94 |
|
|
|
| 13 |
A2 |
1213 |
1154 |
0.00 |
|
|
|
| 14 |
A2 |
1016 |
966 |
0.00 |
|
|
|
| 15 |
A2 |
754 |
717 |
0.00 |
|
|
|
| 16 |
A2 |
361 |
343 |
0.00 |
|
|
|
| 17 |
B1 |
3045 |
2896 |
6.41 |
|
|
|
| 18 |
B1 |
1034 |
983 |
0.08 |
|
|
|
| 19 |
B1 |
957 |
910 |
22.52 |
|
|
|
| 20 |
B1 |
862 |
820 |
40.22 |
|
|
|
| 21 |
B1 |
578 |
549 |
21.53 |
|
|
|
| 22 |
B1 |
316 |
301 |
3.67 |
|
|
|
| 23 |
B1 |
131 |
124 |
0.79 |
|
|
|
| 24 |
B2 |
3235 |
3077 |
16.43 |
|
|
|
| 25 |
B2 |
3199 |
3042 |
15.48 |
|
|
|
| 26 |
B2 |
1701 |
1617 |
3.29 |
|
|
|
| 27 |
B2 |
1446 |
1375 |
36.13 |
|
|
|
| 28 |
B2 |
1402 |
1333 |
2.61 |
|
|
|
| 29 |
B2 |
1291 |
1227 |
26.47 |
|
|
|
| 30 |
B2 |
1151 |
1095 |
8.60 |
|
|
|
| 31 |
B2 |
1013 |
963 |
6.34 |
|
|
|
| 32 |
B2 |
579 |
551 |
0.76 |
|
|
|
| 33 |
B2 |
454 |
432 |
15.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22995.9 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 21869.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.420 |
|
|
|
| 2 |
C |
-0.420 |
|
|
|
| 3 |
C |
-0.187 |
|
|
|
| 4 |
C |
-0.187 |
|
|
|
| 5 |
C |
-0.138 |
|
|
|
| 6 |
C |
-0.138 |
|
|
|
| 7 |
O |
-0.498 |
|
|
|
| 8 |
H |
0.184 |
|
|
|
| 9 |
H |
0.184 |
|
|
|
| 10 |
H |
0.180 |
|
|
|
| 11 |
H |
0.180 |
|
|
|
| 12 |
H |
0.211 |
|
|
|
| 13 |
H |
0.211 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.507 |
4.507 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.453 |
0.000 |
0.000 |
| y |
0.000 |
-35.528 |
0.000 |
| z |
0.000 |
0.000 |
-44.306 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.536 |
0.000 |
0.000 |
| y |
0.000 |
7.352 |
0.000 |
| z |
0.000 |
0.000 |
-5.816 |
|
| Polar |
|---|
| 3z2-r2 | -11.631 |
|---|
| x2-y2 | -5.926 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.310 |
0.000 |
0.000 |
| y |
0.000 |
8.925 |
0.000 |
| z |
0.000 |
0.000 |
13.453 |
<r2> (average value of r
2) Å
2
| <r2> |
186.733 |
| (<r2>)1/2 |
13.665 |