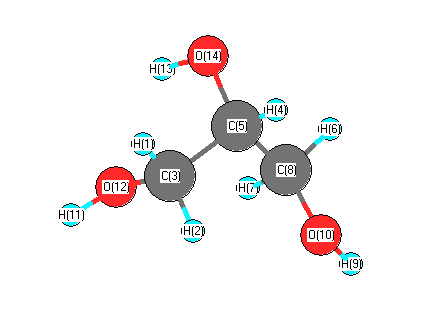
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3817 |
3652 |
33.62 |
|
|
|
| 2 |
A |
3806 |
3642 |
22.04 |
|
|
|
| 3 |
A |
3722 |
3561 |
47.27 |
|
|
|
| 4 |
A |
3092 |
2958 |
50.29 |
|
|
|
| 5 |
A |
3091 |
2958 |
42.74 |
|
|
|
| 6 |
A |
3059 |
2927 |
39.50 |
|
|
|
| 7 |
A |
3033 |
2902 |
39.53 |
|
|
|
| 8 |
A |
3019 |
2889 |
52.54 |
|
|
|
| 9 |
A |
1545 |
1478 |
6.93 |
|
|
|
| 10 |
A |
1532 |
1465 |
10.27 |
|
|
|
| 11 |
A |
1473 |
1410 |
1.22 |
|
|
|
| 12 |
A |
1465 |
1402 |
38.08 |
|
|
|
| 13 |
A |
1439 |
1377 |
41.56 |
|
|
|
| 14 |
A |
1384 |
1324 |
6.77 |
|
|
|
| 15 |
A |
1344 |
1286 |
47.22 |
|
|
|
| 16 |
A |
1291 |
1235 |
20.62 |
|
|
|
| 17 |
A |
1259 |
1205 |
15.38 |
|
|
|
| 18 |
A |
1233 |
1180 |
0.59 |
|
|
|
| 19 |
A |
1206 |
1153 |
36.59 |
|
|
|
| 20 |
A |
1153 |
1103 |
35.14 |
|
|
|
| 21 |
A |
1119 |
1070 |
91.24 |
|
|
|
| 22 |
A |
1103 |
1055 |
80.41 |
|
|
|
| 23 |
A |
1079 |
1032 |
76.54 |
|
|
|
| 24 |
A |
1012 |
968 |
16.15 |
|
|
|
| 25 |
A |
954 |
912 |
8.46 |
|
|
|
| 26 |
A |
835 |
799 |
13.01 |
|
|
|
| 27 |
A |
657 |
628 |
28.97 |
|
|
|
| 28 |
A |
489 |
468 |
125.45 |
|
|
|
| 29 |
A |
487 |
466 |
11.01 |
|
|
|
| 30 |
A |
405 |
387 |
5.19 |
|
|
|
| 31 |
A |
278 |
266 |
4.68 |
|
|
|
| 32 |
A |
256 |
245 |
44.81 |
|
|
|
| 33 |
A |
231 |
221 |
97.42 |
|
|
|
| 34 |
A |
220 |
211 |
89.79 |
|
|
|
| 35 |
A |
166 |
159 |
25.45 |
|
|
|
| 36 |
A |
99 |
95 |
2.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26176.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 25043.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.157 |
|
|
0.047 |
| 2 |
H |
0.178 |
|
|
0.080 |
| 3 |
C |
-0.114 |
|
|
-0.033 |
| 4 |
H |
0.174 |
|
|
0.053 |
| 5 |
C |
0.080 |
|
|
0.327 |
| 6 |
H |
0.158 |
|
|
0.026 |
| 7 |
H |
0.154 |
|
|
0.031 |
| 8 |
C |
-0.097 |
|
|
0.107 |
| 9 |
H |
0.410 |
|
|
0.428 |
| 10 |
O |
-0.636 |
|
|
-0.654 |
| 11 |
H |
0.417 |
|
|
0.430 |
| 12 |
O |
-0.650 |
|
|
-0.605 |
| 13 |
H |
0.419 |
|
|
0.397 |
| 14 |
O |
-0.648 |
|
|
-0.633 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.331 |
-2.145 |
-0.720 |
2.286 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.319 |
-2.153 |
-0.734 |
2.297 |
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
173.393 |
| (<r2>)1/2 |
13.168 |