return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H8O3 (1,2,3-Propanetriol)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-344.411006
Energy at 298.15K-344.421261
HF Energy-344.106720
Nuclear repulsion energy265.456958
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3799 3606 32.92      
2 A 3789 3597 21.58      
3 A 3722 3533 44.48      
4 A 3126 2967 35.31      
5 A 3121 2962 56.06      
6 A 3096 2939 30.56      
7 A 3065 2909 36.33      
8 A 3054 2899 47.97      
9 A 1570 1490 6.46      
10 A 1557 1478 8.98      
11 A 1491 1415 0.70      
12 A 1482 1406 27.58      
13 A 1457 1383 47.35      
14 A 1404 1333 9.82      
15 A 1355 1286 51.44      
16 A 1303 1236 17.49      
17 A 1268 1204 17.44      
18 A 1244 1181 0.68      
19 A 1215 1153 39.05      
20 A 1161 1102 17.22      
21 A 1123 1066 86.65      
22 A 1089 1034 95.51      
23 A 1080 1025 66.90      
24 A 1014 963 18.43      
25 A 965 916 9.04      
26 A 842 799 15.03      
27 A 659 626 29.80      
28 A 491 466 9.00      
29 A 488 463 130.59      
30 A 411 391 5.37      
31 A 288 273 3.05      
32 A 260 247 32.90      
33 A 239 227 114.90      
34 A 229 217 81.86      
35 A 173 165 32.23      
36 A 100 95 2.61      

Unscaled Zero Point Vibrational Energy (zpe) 26363.8 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 25024.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.15590 0.09253 0.07020

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -1.720 -0.478 1.299
H2 -0.213 -1.416 1.151
C3 -0.860 -0.691 0.643
H4 0.276 0.945 1.416
C5 -0.073 0.601 0.432
H6 1.666 1.330 -0.580
H7 0.791 0.028 -1.428
C8 1.140 0.376 -0.451
H9 2.725 -0.760 -0.346
O10 1.954 -0.594 0.216
H11 -1.770 -1.996 -0.490
O12 -1.290 -1.166 -0.632
H13 -1.613 1.793 0.330
O14 -0.837 1.615 -0.221

Atom - Atom Distances (Å)
  H1 H2 C3 H4 C5 H6 H7 C8 H9 O10 H11 O12 H13 O14
H11.78111.10202.45412.15144.27403.74143.45944.74783.83182.34642.09502.47132.7332
H21.78111.09702.42612.14633.75173.12192.75833.36232.49942.33442.09843.59653.3852
C31.10201.09702.13641.52743.45902.74452.51663.71972.84801.95261.42752.61432.4622
H42.45412.42612.13641.09902.46273.03212.13363.46512.57364.05753.33252.33852.0896
C52.15142.14631.52741.09902.14042.12961.51663.20712.36293.23582.39541.95071.4275
H64.27403.75173.45902.46272.14041.78361.09782.35482.10274.78273.86983.43522.5445
H73.74143.12192.74453.03212.12961.78361.09482.35182.10773.39572.52783.46282.5739
C83.45942.75832.51662.13361.51661.09781.09481.95281.43113.75362.88373.19372.3440
H94.74783.36233.71973.46513.20712.35482.35181.95280.96844.66334.04575.07924.2825
O103.83182.49942.84802.57362.36292.10272.10771.43110.96844.04053.40174.29423.5859
H112.34642.33441.95264.05753.23584.78273.39573.75364.66334.04050.96863.87963.7386
O122.09502.09841.42753.33252.39543.86982.52782.88374.04573.40170.96863.12872.8475
H132.47133.59652.61432.33851.95073.43523.46283.19375.07924.29423.87963.12870.9693
O142.73323.38522.46222.08961.42752.54452.57392.34404.28253.58593.73862.84750.9693

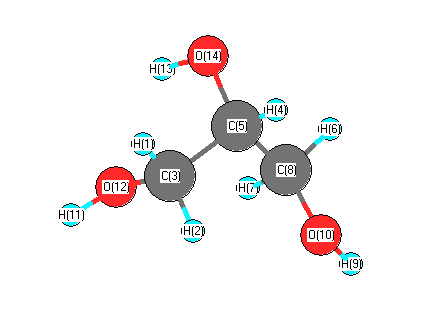
picture of 1,2,3-Propanetriol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C3 H2 108.178 H1 C3 C5 108.727
H1 C3 O12 111.183 H2 C3 C5 108.628
H2 C3 O12 111.782 C3 C5 H4 107.749
C3 C5 C8 111.532 C3 C5 O14 112.826
C3 O12 H11 107.611 H4 C5 C8 108.257
H4 C5 O14 110.928 C5 C3 O12 108.270
C5 C8 H6 108.856 C5 C8 H7 108.185
C5 C8 O10 106.529 C5 O14 H13 107.406
H6 C8 H7 108.876 H6 C8 O10 111.823
H7 C8 O10 112.434 C8 C5 O14 105.489
C8 O10 H9 107.368
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 H 0.146      
2 H 0.170      
3 C -0.055      
4 H 0.162      
5 C 0.133      
6 H 0.147      
7 H 0.146      
8 C -0.038      
9 H 0.416      
10 O -0.683      
11 H 0.422      
12 O -0.698      
13 H 0.430      
14 O -0.700      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 6.772 0.092 0.048
y 0.092 6.101 -0.111
z 0.048 -0.111 5.865


<r2> (average value of r2) Å2
<r2> 173.237
(<r2>)1/2 13.162