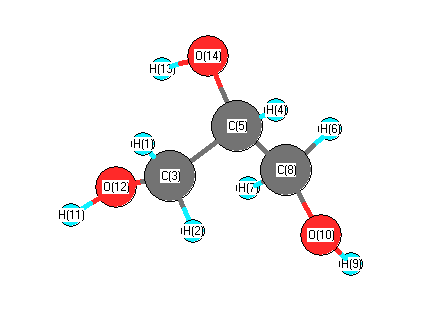
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3840 |
3652 |
37.70 |
|
|
|
| 2 |
A |
3829 |
3641 |
24.64 |
|
|
|
| 3 |
A |
3740 |
3557 |
50.54 |
|
|
|
| 4 |
A |
3107 |
2954 |
38.04 |
|
|
|
| 5 |
A |
3105 |
2953 |
52.22 |
|
|
|
| 6 |
A |
3074 |
2923 |
38.76 |
|
|
|
| 7 |
A |
3047 |
2898 |
38.13 |
|
|
|
| 8 |
A |
3032 |
2883 |
49.82 |
|
|
|
| 9 |
A |
1548 |
1472 |
7.32 |
|
|
|
| 10 |
A |
1535 |
1459 |
11.03 |
|
|
|
| 11 |
A |
1481 |
1408 |
0.80 |
|
|
|
| 12 |
A |
1475 |
1403 |
42.94 |
|
|
|
| 13 |
A |
1445 |
1374 |
42.48 |
|
|
|
| 14 |
A |
1388 |
1320 |
6.49 |
|
|
|
| 15 |
A |
1352 |
1286 |
44.35 |
|
|
|
| 16 |
A |
1296 |
1233 |
20.68 |
|
|
|
| 17 |
A |
1264 |
1202 |
16.11 |
|
|
|
| 18 |
A |
1238 |
1177 |
0.70 |
|
|
|
| 19 |
A |
1210 |
1150 |
38.43 |
|
|
|
| 20 |
A |
1162 |
1105 |
41.93 |
|
|
|
| 21 |
A |
1130 |
1075 |
107.56 |
|
|
|
| 22 |
A |
1117 |
1063 |
64.49 |
|
|
|
| 23 |
A |
1089 |
1035 |
71.15 |
|
|
|
| 24 |
A |
1020 |
970 |
14.55 |
|
|
|
| 25 |
A |
960 |
913 |
9.01 |
|
|
|
| 26 |
A |
844 |
803 |
12.69 |
|
|
|
| 27 |
A |
662 |
629 |
28.97 |
|
|
|
| 28 |
A |
497 |
472 |
124.49 |
|
|
|
| 29 |
A |
492 |
468 |
11.98 |
|
|
|
| 30 |
A |
408 |
388 |
5.48 |
|
|
|
| 31 |
A |
285 |
271 |
3.79 |
|
|
|
| 32 |
A |
259 |
246 |
41.59 |
|
|
|
| 33 |
A |
235 |
223 |
100.60 |
|
|
|
| 34 |
A |
224 |
213 |
88.30 |
|
|
|
| 35 |
A |
170 |
162 |
28.70 |
|
|
|
| 36 |
A |
101 |
96 |
2.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26328.3 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 25038.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.159 |
|
|
|
| 2 |
H |
0.180 |
|
|
|
| 3 |
C |
-0.116 |
|
|
|
| 4 |
H |
0.176 |
|
|
|
| 5 |
C |
0.076 |
|
|
|
| 6 |
H |
0.160 |
|
|
|
| 7 |
H |
0.155 |
|
|
|
| 8 |
C |
-0.099 |
|
|
|
| 9 |
H |
0.412 |
|
|
|
| 10 |
O |
-0.639 |
|
|
|
| 11 |
H |
0.420 |
|
|
|
| 12 |
O |
-0.654 |
|
|
|
| 13 |
H |
0.421 |
|
|
|
| 14 |
O |
-0.650 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.305 |
-2.176 |
-0.724 |
2.314 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.718 |
4.219 |
-3.210 |
| y |
4.219 |
-37.907 |
-0.313 |
| z |
-3.210 |
-0.313 |
-35.385 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.927 |
4.219 |
-3.210 |
| y |
4.219 |
-5.855 |
-0.313 |
| z |
-3.210 |
-0.313 |
-2.072 |
|
| Polar |
|---|
| 3z2-r2 | -4.144 |
|---|
| x2-y2 | 9.188 |
|---|
| xy | 4.219 |
|---|
| xz | -3.210 |
|---|
| yz | -0.313 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.835 |
0.111 |
0.018 |
| y |
0.111 |
6.138 |
-0.130 |
| z |
0.018 |
-0.130 |
5.910 |
<r2> (average value of r
2) Å
2
| <r2> |
172.219 |
| (<r2>)1/2 |
13.123 |