Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -549.875751 |
| Energy at 298.15K | |
| HF Energy | -548.930560 |
| Nuclear repulsion energy | 356.076245 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
1941 |
1793 |
0.00 |
|
|
|
| 2 |
A1g |
736 |
680 |
0.00 |
|
|
|
| 3 |
A2g |
651 |
602 |
0.00 |
|
|
|
| 4 |
A2u |
403i |
372i |
93.08 |
|
|
|
| 5 |
B1g |
1617 |
1494 |
0.00 |
|
|
|
| 6 |
B1g |
646 |
597 |
0.00 |
|
|
|
| 7 |
B2g |
530 |
490 |
0.00 |
|
|
|
| 8 |
B2g |
167 |
154 |
0.00 |
|
|
|
| 9 |
B2u |
339 |
313 |
0.00 |
|
|
|
| 10 |
B2u |
298i |
275i |
0.00 |
|
|
|
| 11 |
Eg |
797i |
736i |
0.00 |
|
|
|
| 11 |
Eg |
797i |
736i |
0.00 |
|
|
|
| 12 |
Eu |
1765 |
1630 |
138.65 |
|
|
|
| 12 |
Eu |
1765 |
1630 |
138.65 |
|
|
|
| 13 |
Eu |
1007 |
930 |
3.86 |
|
|
|
| 13 |
Eu |
1007 |
930 |
3.86 |
|
|
|
| 14 |
Eu |
204 |
189 |
20.61 |
|
|
|
| 14 |
Eu |
204 |
189 |
20.61 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5142.1 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 4749.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is D4h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
1.004 |
0.000 |
| C2 |
1.004 |
0.000 |
0.000 |
| C3 |
0.000 |
-1.004 |
0.000 |
| C4 |
-1.004 |
0.000 |
0.000 |
| F5 |
0.000 |
2.313 |
0.000 |
| F6 |
2.313 |
0.000 |
0.000 |
| F7 |
0.000 |
-2.313 |
0.000 |
| F8 |
-2.313 |
0.000 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
| C1 | | 1.4204 | 2.0087 | 1.4204 | 1.3083 | 2.5213 | 3.3169 | 2.5213 |
C2 | 1.4204 | | 1.4204 | 2.0087 | 2.5213 | 1.3083 | 2.5213 | 3.3169 | C3 | 2.0087 | 1.4204 | | 1.4204 | 3.3169 | 2.5213 | 1.3083 | 2.5213 | C4 | 1.4204 | 2.0087 | 1.4204 | | 2.5213 | 3.3169 | 2.5213 | 1.3083 | F5 | 1.3083 | 2.5213 | 3.3169 | 2.5213 | | 3.2705 | 4.6252 | 3.2705 | F6 | 2.5213 | 1.3083 | 2.5213 | 3.3169 | 3.2705 | | 3.2705 | 4.6252 | F7 | 3.3169 | 2.5213 | 1.3083 | 2.5213 | 4.6252 | 3.2705 | | 3.2705 | F8 | 2.5213 | 3.3169 | 2.5213 | 1.3083 | 3.2705 | 4.6252 | 3.2705 | |
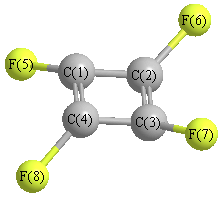 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
90.000 |
|
C1 |
C2 |
F6 |
135.000 |
| C1 |
C4 |
C3 |
90.000 |
|
C1 |
C4 |
F8 |
135.000 |
| C2 |
C1 |
C4 |
90.000 |
|
C2 |
C1 |
F5 |
135.000 |
| C2 |
C3 |
C4 |
90.000 |
|
C2 |
C3 |
F7 |
135.000 |
| C3 |
C2 |
F6 |
135.000 |
|
C3 |
C4 |
F8 |
135.000 |
| C4 |
C1 |
F5 |
135.000 |
|
C4 |
C3 |
F7 |
135.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -549.961253 |
| Energy at 298.15K | -549.961206 |
| HF Energy | -549.007801 |
| Nuclear repulsion energy | 354.722672 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
2035 |
1880 |
0.00 |
|
|
|
| 2 |
Ag |
1348 |
1245 |
0.00 |
|
|
|
| 3 |
Ag |
688 |
635 |
0.00 |
|
|
|
| 4 |
Ag |
241 |
223 |
0.00 |
|
|
|
| 5 |
Ag |
91 |
84 |
0.00 |
|
|
|
| 6 |
Au |
1444 |
1333 |
466.35 |
|
|
|
| 7 |
Au |
996 |
920 |
68.28 |
|
|
|
| 8 |
Au |
596 |
551 |
10.72 |
|
|
|
| 9 |
Au |
238 |
219 |
0.27 |
|
|
|
| 10 |
Au |
172 |
159 |
0.01 |
|
|
|
| 11 |
Bg |
1480 |
1367 |
0.00 |
|
|
|
| 12 |
Bg |
802 |
741 |
0.00 |
|
|
|
| 13 |
Bg |
524 |
484 |
0.00 |
|
|
|
| 14 |
Bg |
499 |
461 |
0.00 |
|
|
|
| 15 |
Bu |
2035 |
1880 |
99.24 |
|
|
|
| 16 |
Bu |
1044 |
964 |
190.38 |
|
|
|
| 17 |
Bu |
319 |
294 |
9.56 |
|
|
|
| 18 |
Bu |
218 |
202 |
1.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7384.0 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 6820.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.043 |
0.781 |
0.655 |
| C2 |
-0.043 |
-0.781 |
0.655 |
| C3 |
-0.043 |
-0.781 |
-0.655 |
| C4 |
0.043 |
0.781 |
-0.655 |
| F5 |
-0.043 |
1.643 |
1.637 |
| F6 |
0.043 |
-1.643 |
1.637 |
| F7 |
0.043 |
-1.643 |
-1.637 |
| F8 |
-0.043 |
1.643 |
-1.637 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
| C1 | | 1.5644 | 2.0401 | 1.3094 | 1.3100 | 2.6157 | 3.3361 | 2.4503 |
C2 | 1.5644 | | 1.3094 | 2.0401 | 2.6157 | 1.3100 | 2.4503 | 3.3361 | C3 | 2.0401 | 1.3094 | | 1.5644 | 3.3361 | 2.4503 | 1.3100 | 2.6157 | C4 | 1.3094 | 2.0401 | 1.5644 | | 2.4503 | 3.3361 | 2.6157 | 1.3100 | F5 | 1.3100 | 2.6157 | 3.3361 | 2.4503 | | 3.2874 | 4.6400 | 3.2746 | F6 | 2.6157 | 1.3100 | 2.4503 | 3.3361 | 3.2874 | | 3.2746 | 4.6400 | F7 | 3.3361 | 2.4503 | 1.3100 | 2.6157 | 4.6400 | 3.2746 | | 3.2874 | F8 | 2.4503 | 3.3361 | 2.6157 | 1.3100 | 3.2746 | 4.6400 | 3.2874 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
90.000 |
|
C1 |
C2 |
F6 |
130.804 |
| C1 |
C4 |
C3 |
90.000 |
|
C1 |
C4 |
F8 |
138.595 |
| C2 |
C1 |
C4 |
90.000 |
|
C2 |
C1 |
F5 |
130.804 |
| C2 |
C3 |
C4 |
90.000 |
|
C2 |
C3 |
F7 |
138.595 |
| C3 |
C2 |
F6 |
138.595 |
|
C3 |
C4 |
F8 |
130.804 |
| C4 |
C1 |
F5 |
138.595 |
|
C4 |
C3 |
F7 |
130.804 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
