Jump to
S1C2
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -550.969776 |
| Energy at 298.15K | |
| HF Energy | -550.969776 |
| Nuclear repulsion energy | 350.252807 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is D4h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
1.024 |
0.000 |
| C2 |
1.024 |
0.000 |
0.000 |
| C3 |
0.000 |
-1.024 |
0.000 |
| C4 |
-1.024 |
0.000 |
0.000 |
| F5 |
0.000 |
2.350 |
0.000 |
| F6 |
2.350 |
0.000 |
0.000 |
| F7 |
0.000 |
-2.350 |
0.000 |
| F8 |
-2.350 |
0.000 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
| C1 | | 1.4488 | 2.0489 | 1.4488 | 1.3259 | 2.5639 | 3.3748 | 2.5639 |
C2 | 1.4488 | | 1.4488 | 2.0489 | 2.5639 | 1.3259 | 2.5639 | 3.3748 | C3 | 2.0489 | 1.4488 | | 1.4488 | 3.3748 | 2.5639 | 1.3259 | 2.5639 | C4 | 1.4488 | 2.0489 | 1.4488 | | 2.5639 | 3.3748 | 2.5639 | 1.3259 | F5 | 1.3259 | 2.5639 | 3.3748 | 2.5639 | | 3.3239 | 4.7007 | 3.3239 | F6 | 2.5639 | 1.3259 | 2.5639 | 3.3748 | 3.3239 | | 3.3239 | 4.7007 | F7 | 3.3748 | 2.5639 | 1.3259 | 2.5639 | 4.7007 | 3.3239 | | 3.3239 | F8 | 2.5639 | 3.3748 | 2.5639 | 1.3259 | 3.3239 | 4.7007 | 3.3239 | |
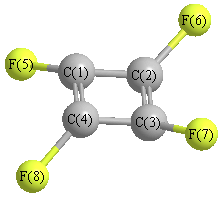 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
90.000 |
|
C1 |
C2 |
F6 |
135.000 |
| C1 |
C4 |
C3 |
90.000 |
|
C1 |
C4 |
F8 |
135.000 |
| C2 |
C1 |
C4 |
90.000 |
|
C2 |
C1 |
F5 |
135.000 |
| C2 |
C3 |
C4 |
90.000 |
|
C2 |
C3 |
F7 |
135.000 |
| C3 |
C2 |
F6 |
135.000 |
|
C3 |
C4 |
F8 |
135.000 |
| C4 |
C1 |
F5 |
135.000 |
|
C4 |
C3 |
F7 |
135.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.045 |
|
|
|
| 2 |
C |
0.336 |
|
|
|
| 3 |
C |
0.045 |
|
|
|
| 4 |
C |
0.336 |
|
|
|
| 5 |
F |
-0.227 |
|
|
|
| 6 |
F |
-0.153 |
|
|
|
| 7 |
F |
-0.227 |
|
|
|
| 8 |
F |
-0.153 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.071 |
0.000 |
0.000 |
| y |
0.000 |
-46.494 |
0.000 |
| z |
0.000 |
0.000 |
-40.002 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.177 |
0.000 |
0.000 |
| y |
0.000 |
-8.457 |
0.000 |
| z |
0.000 |
0.000 |
1.280 |
|
| Polar |
|---|
| 3z2-r2 | 2.559 |
|---|
| x2-y2 | 10.423 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
249.572 |
| (<r2>)1/2 |
15.798 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -551.048887 |
| Energy at 298.15K | -551.048592 |
| HF Energy | -551.048887 |
| Nuclear repulsion energy | 351.919828 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
1782 |
1753 |
0.00 |
|
|
|
| 2 |
Ag |
1164 |
1144 |
0.00 |
|
|
|
| 3 |
Ag |
714 |
702 |
0.00 |
|
|
|
| 4 |
Ag |
327 |
322 |
0.00 |
|
|
|
| 5 |
Ag |
187 |
184 |
0.00 |
|
|
|
| 6 |
Au |
1274 |
1253 |
484.55 |
|
|
|
| 7 |
Au |
891 |
876 |
32.01 |
|
|
|
| 8 |
Au |
546 |
537 |
65.21 |
|
|
|
| 9 |
Au |
233 |
229 |
0.00 |
|
|
|
| 10 |
Au |
145 |
142 |
0.09 |
|
|
|
| 11 |
Bg |
1300 |
1279 |
0.00 |
|
|
|
| 12 |
Bg |
726 |
714 |
0.00 |
|
|
|
| 13 |
Bg |
467 |
460 |
0.00 |
|
|
|
| 14 |
Bg |
292 |
287 |
0.00 |
|
|
|
| 15 |
Bu |
1745 |
1716 |
213.46 |
|
|
|
| 16 |
Bu |
966 |
950 |
179.78 |
|
|
|
| 17 |
Bu |
300 |
295 |
3.29 |
|
|
|
| 18 |
Bu |
172 |
169 |
1.93 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6615.0 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 6505.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.135 |
0.766 |
0.674 |
| C2 |
-0.135 |
-0.766 |
0.674 |
| C3 |
-0.135 |
-0.766 |
-0.674 |
| C4 |
0.135 |
0.766 |
-0.674 |
| F5 |
-0.135 |
1.681 |
1.602 |
| F6 |
0.135 |
-1.681 |
1.602 |
| F7 |
0.135 |
-1.681 |
-1.602 |
| F8 |
-0.135 |
1.681 |
-1.602 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
| C1 | | 1.5548 | 2.0572 | 1.3471 | 1.3319 | 2.6171 | 3.3415 | 2.4679 |
C2 | 1.5548 | | 1.3471 | 2.0572 | 2.6171 | 1.3319 | 2.4679 | 3.3415 | C3 | 2.0572 | 1.3471 | | 1.5548 | 3.3415 | 2.4679 | 1.3319 | 2.6171 | C4 | 1.3471 | 2.0572 | 1.5548 | | 2.4679 | 3.3415 | 2.6171 | 1.3319 | F5 | 1.3319 | 2.6171 | 3.3415 | 2.4679 | | 3.3731 | 4.6527 | 3.2046 | F6 | 2.6171 | 1.3319 | 2.4679 | 3.3415 | 3.3731 | | 3.2046 | 4.6527 | F7 | 3.3415 | 2.4679 | 1.3319 | 2.6171 | 4.6527 | 3.2046 | | 3.3731 | F8 | 2.4679 | 3.3415 | 2.6171 | 1.3319 | 3.2046 | 4.6527 | 3.3731 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
90.000 |
|
C1 |
C2 |
F6 |
129.920 |
| C1 |
C4 |
C3 |
90.000 |
|
C1 |
C4 |
F8 |
134.215 |
| C2 |
C1 |
C4 |
90.000 |
|
C2 |
C1 |
F5 |
129.920 |
| C2 |
C3 |
C4 |
90.000 |
|
C2 |
C3 |
F7 |
134.215 |
| C3 |
C2 |
F6 |
134.215 |
|
C3 |
C4 |
F8 |
129.920 |
| C4 |
C1 |
F5 |
134.215 |
|
C4 |
C3 |
F7 |
129.920 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.194 |
|
|
|
| 2 |
C |
0.194 |
|
|
|
| 3 |
C |
0.194 |
|
|
|
| 4 |
C |
0.194 |
|
|
|
| 5 |
F |
-0.194 |
|
|
|
| 6 |
F |
-0.194 |
|
|
|
| 7 |
F |
-0.194 |
|
|
|
| 8 |
F |
-0.194 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.756 |
-0.165 |
0.000 |
| y |
-0.165 |
-41.792 |
0.000 |
| z |
0.000 |
0.000 |
-41.556 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.918 |
-0.165 |
0.000 |
| y |
-0.165 |
-1.136 |
0.000 |
| z |
0.000 |
0.000 |
-0.782 |
|
| Polar |
|---|
| 3z2-r2 | -1.564 |
|---|
| x2-y2 | 2.036 |
|---|
| xy | -0.165 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.074 |
-0.045 |
0.000 |
| y |
-0.045 |
6.406 |
0.000 |
| z |
0.000 |
0.000 |
7.815 |
<r2> (average value of r
2) Å
2
| <r2> |
245.848 |
| (<r2>)1/2 |
15.680 |
