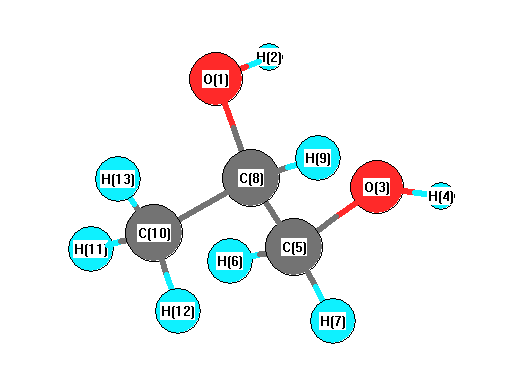
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3614 |
3554 |
8.80 |
|
|
|
| 2 |
A |
3520 |
3462 |
43.74 |
|
|
|
| 3 |
A |
3079 |
3028 |
17.65 |
|
|
|
| 4 |
A |
3069 |
3019 |
29.45 |
|
|
|
| 5 |
A |
3043 |
2993 |
23.99 |
|
|
|
| 6 |
A |
2992 |
2942 |
18.31 |
|
|
|
| 7 |
A |
2952 |
2904 |
52.16 |
|
|
|
| 8 |
A |
2831 |
2785 |
91.26 |
|
|
|
| 9 |
A |
1484 |
1460 |
1.71 |
|
|
|
| 10 |
A |
1469 |
1445 |
4.26 |
|
|
|
| 11 |
A |
1465 |
1441 |
5.46 |
|
|
|
| 12 |
A |
1424 |
1401 |
46.03 |
|
|
|
| 13 |
A |
1386 |
1363 |
44.44 |
|
|
|
| 14 |
A |
1370 |
1347 |
9.59 |
|
|
|
| 15 |
A |
1349 |
1327 |
5.17 |
|
|
|
| 16 |
A |
1328 |
1306 |
51.79 |
|
|
|
| 17 |
A |
1275 |
1254 |
28.02 |
|
|
|
| 18 |
A |
1199 |
1179 |
14.45 |
|
|
|
| 19 |
A |
1151 |
1132 |
22.30 |
|
|
|
| 20 |
A |
1070 |
1053 |
22.87 |
|
|
|
| 21 |
A |
1064 |
1046 |
11.32 |
|
|
|
| 22 |
A |
1022 |
1005 |
107.02 |
|
|
|
| 23 |
A |
921 |
906 |
5.74 |
|
|
|
| 24 |
A |
903 |
888 |
15.29 |
|
|
|
| 25 |
A |
835 |
821 |
11.97 |
|
|
|
| 26 |
A |
591 |
581 |
145.59 |
|
|
|
| 27 |
A |
518 |
510 |
5.59 |
|
|
|
| 28 |
A |
473 |
466 |
8.77 |
|
|
|
| 29 |
A |
364 |
358 |
73.85 |
|
|
|
| 30 |
A |
349 |
343 |
10.44 |
|
|
|
| 31 |
A |
247 |
243 |
2.38 |
|
|
|
| 32 |
A |
220 |
216 |
0.26 |
|
|
|
| 33 |
A |
153 |
151 |
4.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24365.0 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 23963.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.594 |
|
|
|
| 2 |
H |
0.387 |
|
|
|
| 3 |
O |
-0.580 |
|
|
|
| 4 |
H |
0.389 |
|
|
|
| 5 |
C |
-0.115 |
|
|
|
| 6 |
H |
0.170 |
|
|
|
| 7 |
H |
0.139 |
|
|
|
| 8 |
C |
0.090 |
|
|
|
| 9 |
H |
0.108 |
|
|
|
| 10 |
C |
-0.490 |
|
|
|
| 11 |
H |
0.167 |
|
|
|
| 12 |
H |
0.152 |
|
|
|
| 13 |
H |
0.177 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.555 |
-1.609 |
1.468 |
2.247 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.293 |
2.094 |
2.896 |
| y |
2.094 |
-32.553 |
0.185 |
| z |
2.896 |
0.185 |
-29.633 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.200 |
2.094 |
2.896 |
| y |
2.094 |
-1.590 |
0.185 |
| z |
2.896 |
0.185 |
2.790 |
|
| Polar |
|---|
| 3z2-r2 | 5.579 |
|---|
| x2-y2 | 0.260 |
|---|
| xy | 2.094 |
|---|
| xz | 2.896 |
|---|
| yz | 0.185 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.805 |
0.211 |
0.171 |
| y |
0.211 |
6.075 |
-0.116 |
| z |
0.171 |
-0.116 |
5.728 |
<r2> (average value of r
2) Å
2
| <r2> |
132.790 |
| (<r2>)1/2 |
11.523 |