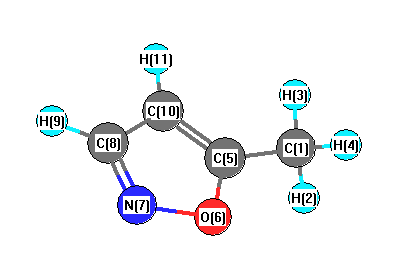
Vibrational Frequencies calculated at B3LYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3292 |
3161 |
0.52 |
|
|
|
| 2 |
A |
3258 |
3128 |
5.02 |
|
|
|
| 3 |
A |
3151 |
3026 |
9.33 |
|
|
|
| 4 |
A |
3115 |
2991 |
9.36 |
|
|
|
| 5 |
A |
3059 |
2937 |
14.28 |
|
|
|
| 6 |
A |
1655 |
1589 |
32.27 |
|
|
|
| 7 |
A |
1531 |
1470 |
41.11 |
|
|
|
| 8 |
A |
1516 |
1456 |
10.78 |
|
|
|
| 9 |
A |
1504 |
1444 |
7.36 |
|
|
|
| 10 |
A |
1442 |
1384 |
1.27 |
|
|
|
| 11 |
A |
1385 |
1330 |
6.53 |
|
|
|
| 12 |
A |
1270 |
1219 |
22.23 |
|
|
|
| 13 |
A |
1224 |
1176 |
8.13 |
|
|
|
| 14 |
A |
1081 |
1038 |
4.10 |
|
|
|
| 15 |
A |
1064 |
1022 |
3.36 |
|
|
|
| 16 |
A |
1032 |
991 |
8.07 |
|
|
|
| 17 |
A |
1007 |
967 |
1.98 |
|
|
|
| 18 |
A |
942 |
904 |
9.02 |
|
|
|
| 19 |
A |
904 |
868 |
3.83 |
|
|
|
| 20 |
A |
904 |
868 |
16.66 |
|
|
|
| 21 |
A |
802 |
771 |
30.26 |
|
|
|
| 22 |
A |
661 |
635 |
0.40 |
|
|
|
| 23 |
A |
649 |
623 |
2.77 |
|
|
|
| 24 |
A |
639 |
614 |
1.06 |
|
|
|
| 25 |
A |
333 |
320 |
2.06 |
|
|
|
| 26 |
A |
250 |
240 |
1.78 |
|
|
|
| 27 |
A |
98 |
94 |
0.80 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18882.9 cm
-1
Scaled (by 0.9603) Zero Point Vibrational Energy (zpe) 18133.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.524 |
-0.449 |
|
-0.624 |
| 2 |
H |
0.185 |
0.131 |
|
0.176 |
| 3 |
H |
0.172 |
0.138 |
|
0.183 |
| 4 |
H |
0.185 |
0.131 |
|
0.176 |
| 5 |
C |
0.413 |
0.473 |
|
0.537 |
| 6 |
O |
-0.363 |
-0.115 |
|
-0.107 |
| 7 |
N |
-0.164 |
-0.381 |
|
-0.382 |
| 8 |
C |
0.054 |
0.306 |
|
0.279 |
| 9 |
H |
0.163 |
0.065 |
|
0.090 |
| 10 |
C |
-0.269 |
-0.473 |
|
-0.535 |
| 11 |
H |
0.147 |
0.173 |
|
0.205 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.896 |
2.583 |
0.000 |
3.205 |
| CHELPG |
1.855 |
2.551 |
0.000 |
3.154 |
| AIM |
|
|
|
|
| ESP |
1.881 |
2.562 |
0.000 |
3.178 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.113 |
-3.537 |
-0.000 |
| y |
-3.537 |
-35.203 |
0.000 |
| z |
-0.000 |
0.000 |
-35.549 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.264 |
-3.537 |
-0.000 |
| y |
-3.537 |
-1.872 |
0.000 |
| z |
-0.000 |
0.000 |
-2.392 |
|
| Polar |
|---|
| 3z2-r2 | -4.783 |
|---|
| x2-y2 | 4.090 |
|---|
| xy | -3.537 |
|---|
| xz | -0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.319 |
-0.408 |
0.000 |
| y |
-0.408 |
7.212 |
-0.001 |
| z |
0.000 |
-0.001 |
3.854 |
<r2> (average value of r
2) Å
2
| <r2> |
136.132 |
| (<r2>)1/2 |
11.668 |