Jump to
S1C2
Energy calculated at MP4=FULL/6-31G*
| | hartrees |
|---|
| Energy at 0K | -259.297198 |
| Energy at 298.15K | |
| HF Energy | -258.230837 |
| Nuclear repulsion energy | 74.365190 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
286 |
275 |
|
|
|
|
| 2 |
A1 |
62 |
60 |
|
|
|
|
| 3 |
A2 |
48i |
47i |
|
|
|
|
| 4 |
B2 |
151i |
145i |
|
|
|
|
| 5 |
B2 |
4458i |
4287i |
|
|
|
|
| 6 |
A1 |
1517 |
1458 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) -1396.7 cm
-1
Scaled (by 0.9616) Zero Point Vibrational Energy (zpe) -1343.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/6-31G*
Point Group is C2v
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP4=FULL/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
7 |
0.000 |
2.296 |
0.638 |
|
| 2 |
7 |
0.000 |
-2.296 |
0.638 |
|
| 3 |
8 |
0.000 |
2.393 |
-0.558 |
|
| 4 |
8 |
0.000 |
-2.393 |
-0.558 |
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
189.418 |
| (<r2>)1/2 |
13.763 |
Jump to
S1C1
Energy calculated at MP4=FULL/6-31G*
| | hartrees |
|---|
| Energy at 0K | -259.297478 |
| Energy at 298.15K | |
| HF Energy | -258.228829 |
| Nuclear repulsion energy | 72.818082 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
1478 |
1421 |
|
|
|
|
| 2 |
Ag |
283 |
272 |
|
|
|
|
| 3 |
Ag |
114i |
110i |
|
|
|
|
| 4 |
Au |
8i |
8i |
|
|
|
|
| 5 |
Bu |
105i |
101i |
|
|
|
|
| 6 |
Bu |
4365i |
4197i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) -1415.7 cm
-1
Scaled (by 0.9616) Zero Point Vibrational Energy (zpe) -1361.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-0.504 |
2.169 |
0.000 |
| N2 |
0.504 |
-2.169 |
0.000 |
| O3 |
0.504 |
2.827 |
0.000 |
| O4 |
-0.504 |
-2.827 |
0.000 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
O3 |
O4 |
| N1 | | 4.4532 | 1.2037 | 4.9958 |
N2 | 4.4532 | | 4.9958 | 1.2037 | O3 | 1.2037 | 4.9958 | | 5.7430 | O4 | 4.9958 | 1.2037 | 5.7430 | |
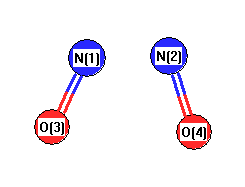 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
N2 |
O4 |
110.064 |
|
N2 |
N1 |
O3 |
110.064 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
