return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12O (3-Pentanol)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-271.915183
Energy at 298.15K-271.928862
Nuclear repulsion energy259.752099
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3974 3671 29.55      
2 A 3273 3023 24.30      
3 A 3248 3000 34.23      
4 A 3239 2992 83.97      
5 A 3237 2990 20.99      
6 A 3214 2969 22.42      
7 A 3205 2960 1.69      
8 A 3172 2930 25.17      
9 A 3169 2927 42.87      
10 A 3163 2922 16.75      
11 A 3162 2920 17.98      
12 A 3103 2866 44.44      
13 A 1615 1491 6.50      
14 A 1610 1487 6.40      
15 A 1602 1479 6.37      
16 A 1598 1476 5.11      
17 A 1588 1467 1.63      
18 A 1579 1459 0.31      
19 A 1545 1427 6.39      
20 A 1521 1405 1.52      
21 A 1511 1396 5.65      
22 A 1500 1386 14.00      
23 A 1465 1353 0.05      
24 A 1414 1306 20.28      
25 A 1396 1289 2.49      
26 A 1364 1260 6.61      
27 A 1336 1234 39.87      
28 A 1257 1161 27.91      
29 A 1231 1137 4.74      
30 A 1165 1076 10.84      
31 A 1139 1052 30.94      
32 A 1109 1024 3.73      
33 A 1088 1005 0.42      
34 A 1048 968 40.18      
35 A 990 915 7.87      
36 A 915 845 4.48      
37 A 829 765 1.50      
38 A 817 754 0.17      
39 A 525 485 3.42      
40 A 502 464 10.43      
41 A 420 388 4.31      
42 A 334 308 36.43      
43 A 313 289 64.00      
44 A 253 234 36.45      
45 A 231 213 5.94      
46 A 198 183 0.53      
47 A 116 107 1.53      
48 A 102 94 0.08      

Unscaled Zero Point Vibrational Energy (zpe) 38189.2 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 35275.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.25104 0.06369 0.05540

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.008 0.022 0.218
C2 -1.272 -0.682 -0.245
C3 1.253 -0.719 -0.208
C4 -2.546 -0.013 0.250
C5 2.540 -0.029 0.226
O6 -0.046 1.329 -0.334
H7 -0.025 0.089 1.312
H8 -1.236 -1.718 0.094
H9 -1.263 -0.703 -1.336
H10 1.239 -0.823 -1.293
H11 1.221 -1.726 0.210
H12 -3.428 -0.544 -0.106
H13 -2.583 -0.001 1.339
H14 -2.600 1.013 -0.100
H15 3.412 -0.621 -0.044
H16 2.650 0.942 -0.253
H17 2.565 0.119 1.306
H18 0.615 1.868 0.101

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 O6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.51811.52352.53842.54901.41941.09672.13262.12402.13322.13723.48082.80842.79323.48992.85192.79581.9525
C21.51812.52521.52223.89592.35622.13831.09071.09052.72402.74102.16492.16632.15854.68814.24404.21513.1909
C31.52352.52523.89141.52362.42902.14402.69872.75691.09031.09094.68514.19804.22622.16722.17062.17152.6831
C42.53841.52223.89145.08662.89682.73762.15582.15374.16704.13901.08851.08981.08595.99635.30695.22103.6818
C52.54903.89591.52365.08662.97482.78824.13864.16602.15212.14955.99915.24275.25511.08791.08861.09042.7061
O61.41942.35622.42902.89682.97482.06123.29862.57162.68353.35243.87203.31702.58383.98082.72493.31300.9581
H71.09672.13832.14402.73762.78822.06122.49273.02803.03572.46223.73982.55953.07893.76263.21462.59042.2456
H82.13261.09072.69872.15584.13863.29862.49271.75282.97482.45992.49422.51233.05904.77744.72144.39194.0356
H92.12401.09052.75692.15374.16602.57163.02801.75282.50473.09922.49513.06432.50264.85024.37994.72293.4935
H102.13322.72401.09034.16702.15212.68353.03572.97482.50471.75414.82334.71274.41962.51432.48693.06613.0945
H112.13722.74101.09094.13902.14953.35242.46222.45993.09921.75414.80724.32704.71232.46683.06182.53193.6472
H123.48082.16494.68511.08855.99913.87203.73982.49422.49514.82334.80721.75921.76366.84026.25806.19224.7121
H132.80842.16634.19801.08985.24273.31702.55952.51233.06434.71274.32701.75921.76096.18355.55025.14943.9055
H142.79322.15854.22621.08595.25512.58383.07893.05902.50264.41964.71231.76361.76096.23055.25255.42753.3329
H153.48994.68812.16725.99631.08793.98083.76264.77744.85022.51432.46686.84026.18356.23051.75141.75743.7473
H162.85194.24402.17065.30691.08862.72493.21464.72144.37992.48693.06186.25805.55025.25251.75141.76582.2638
H172.79584.21512.17155.22101.09043.31302.59044.39194.72293.06612.53196.19225.14945.42751.75741.76582.8839
H181.95253.19092.68313.68182.70610.95812.24564.03563.49353.09453.64724.71213.90553.33293.74732.26382.8839

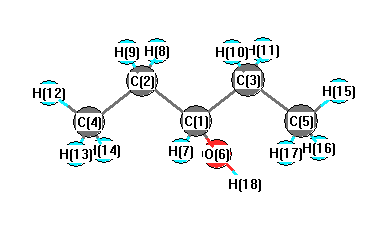
picture of 3-Pentanol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 113.215 C1 C2 H8 108.549
C1 C2 H9 107.893 C1 C3 C5 113.554
C1 C3 H10 108.261 C1 C3 H11 108.534
C1 O6 H18 108.864 C2 C1 C3 112.240
C2 C1 O6 106.611 C2 C1 H7 108.648
C2 C4 H12 110.951 C2 C4 H13 110.982
C2 C4 H14 110.598 C3 C1 O6 111.201
C3 C1 H7 108.732 C3 C5 H15 111.074
C3 C5 H16 111.307 C3 C5 H17 111.269
C4 C2 H8 110.087 C4 C2 H9 109.933
C5 C3 H10 109.728 C5 C3 H11 109.487
O6 C1 H7 109.341 H8 C2 H9 106.947
H10 C3 H11 107.066 H12 C4 H13 107.728
H12 C4 H14 108.399 H13 C4 H14 108.068
H15 C5 H16 107.166 H15 C5 H17 107.571
H16 C5 H17 108.272
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability