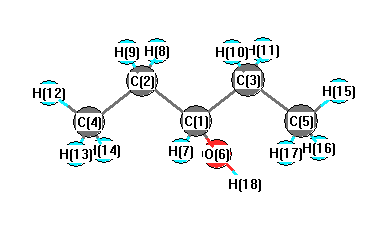
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3798 |
3633 |
12.88 |
|
|
|
| 2 |
A |
3156 |
3019 |
24.16 |
|
|
|
| 3 |
A |
3133 |
2997 |
29.60 |
|
|
|
| 4 |
A |
3127 |
2991 |
52.11 |
|
|
|
| 5 |
A |
3121 |
2986 |
52.46 |
|
|
|
| 6 |
A |
3095 |
2961 |
22.92 |
|
|
|
| 7 |
A |
3088 |
2954 |
1.07 |
|
|
|
| 8 |
A |
3057 |
2925 |
30.35 |
|
|
|
| 9 |
A |
3053 |
2921 |
51.84 |
|
|
|
| 10 |
A |
3048 |
2916 |
28.22 |
|
|
|
| 11 |
A |
3046 |
2914 |
0.93 |
|
|
|
| 12 |
A |
2962 |
2834 |
54.15 |
|
|
|
| 13 |
A |
1538 |
1472 |
7.27 |
|
|
|
| 14 |
A |
1534 |
1468 |
6.24 |
|
|
|
| 15 |
A |
1525 |
1459 |
7.01 |
|
|
|
| 16 |
A |
1521 |
1455 |
5.80 |
|
|
|
| 17 |
A |
1506 |
1441 |
2.45 |
|
|
|
| 18 |
A |
1496 |
1431 |
0.53 |
|
|
|
| 19 |
A |
1455 |
1392 |
6.95 |
|
|
|
| 20 |
A |
1433 |
1371 |
2.17 |
|
|
|
| 21 |
A |
1426 |
1365 |
5.40 |
|
|
|
| 22 |
A |
1415 |
1353 |
14.33 |
|
|
|
| 23 |
A |
1383 |
1323 |
0.15 |
|
|
|
| 24 |
A |
1344 |
1286 |
17.42 |
|
|
|
| 25 |
A |
1322 |
1264 |
3.54 |
|
|
|
| 26 |
A |
1296 |
1240 |
9.15 |
|
|
|
| 27 |
A |
1269 |
1214 |
29.36 |
|
|
|
| 28 |
A |
1184 |
1133 |
25.97 |
|
|
|
| 29 |
A |
1168 |
1118 |
3.27 |
|
|
|
| 30 |
A |
1106 |
1058 |
7.85 |
|
|
|
| 31 |
A |
1080 |
1033 |
25.91 |
|
|
|
| 32 |
A |
1050 |
1004 |
4.13 |
|
|
|
| 33 |
A |
1034 |
990 |
0.67 |
|
|
|
| 34 |
A |
992 |
949 |
50.57 |
|
|
|
| 35 |
A |
943 |
902 |
9.69 |
|
|
|
| 36 |
A |
870 |
832 |
5.36 |
|
|
|
| 37 |
A |
786 |
752 |
1.63 |
|
|
|
| 38 |
A |
774 |
741 |
0.27 |
|
|
|
| 39 |
A |
497 |
476 |
2.87 |
|
|
|
| 40 |
A |
477 |
456 |
9.16 |
|
|
|
| 41 |
A |
397 |
380 |
3.74 |
|
|
|
| 42 |
A |
310 |
297 |
42.10 |
|
|
|
| 43 |
A |
304 |
291 |
55.81 |
|
|
|
| 44 |
A |
238 |
228 |
28.96 |
|
|
|
| 45 |
A |
215 |
206 |
3.88 |
|
|
|
| 46 |
A |
187 |
179 |
0.44 |
|
|
|
| 47 |
A |
107 |
103 |
1.50 |
|
|
|
| 48 |
A |
93 |
89 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36478.6 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 34899.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.104 |
|
|
|
| 2 |
C |
-0.310 |
|
|
|
| 3 |
C |
-0.308 |
|
|
|
| 4 |
C |
-0.513 |
|
|
|
| 5 |
C |
-0.534 |
|
|
|
| 6 |
O |
-0.635 |
|
|
|
| 7 |
H |
0.128 |
|
|
|
| 8 |
H |
0.155 |
|
|
|
| 9 |
H |
0.168 |
|
|
|
| 10 |
H |
0.168 |
|
|
|
| 11 |
H |
0.155 |
|
|
|
| 12 |
H |
0.162 |
|
|
|
| 13 |
H |
0.157 |
|
|
|
| 14 |
H |
0.191 |
|
|
|
| 15 |
H |
0.177 |
|
|
|
| 16 |
H |
0.162 |
|
|
|
| 17 |
H |
0.164 |
|
|
|
| 18 |
H |
0.406 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.127 |
-0.432 |
0.939 |
1.529 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.819 |
2.123 |
0.256 |
| y |
2.123 |
-38.538 |
1.555 |
| z |
0.256 |
1.555 |
-39.624 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.262 |
2.123 |
0.256 |
| y |
2.123 |
0.683 |
1.555 |
| z |
0.256 |
1.555 |
-0.945 |
|
| Polar |
|---|
| 3z2-r2 | -1.890 |
|---|
| x2-y2 | -0.281 |
|---|
| xy | 2.123 |
|---|
| xz | 0.256 |
|---|
| yz | 1.555 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.921 |
0.151 |
0.076 |
| y |
0.151 |
8.120 |
0.053 |
| z |
0.076 |
0.053 |
7.699 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |