return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHCHCH3 (2-Butene, (Z)-)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-156.674350
Energy at 298.15K-156.682009
HF Energy-156.106468
Nuclear repulsion energy117.027888
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3180 3028 68.17      
2 A1 3165 3014 0.95      
3 A1 3062 2916 11.57      
4 A1 1774 1689 2.03      
5 A1 1550 1476 5.78      
6 A1 1474 1404 4.39      
7 A1 1326 1263 0.04      
8 A1 1056 1005 0.73      
9 A1 890 848 0.54      
10 A1 303 288 0.00      
11 A2 3114 2966 0.00      
12 A2 1533 1460 0.00      
13 A2 1086 1034 0.00      
14 A2 997 950 0.00      
15 A2 385 367 0.00      
16 A2 114 109 0.00      
17 B1 3115 2966 51.35      
18 B1 1538 1465 10.00      
19 B1 1099 1047 2.09      
20 B1 708 674 34.51      
21 B1 110 105 0.58      
22 B2 3160 3009 25.40      
23 B2 3146 2996 1.09      
24 B2 3058 2912 39.07      
25 B2 1541 1467 3.15      
26 B2 1485 1414 4.94      
27 B2 1435 1367 2.85      
28 B2 1185 1128 0.72      
29 B2 1011 963 8.64      
30 B2 573 546 5.55      

Unscaled Zero Point Vibrational Energy (zpe) 24086.1 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 22937.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.54143 0.16792 0.13460

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.676 0.638
C2 0.000 -0.676 0.638
C3 0.000 1.662 -0.507
C4 0.000 -1.662 -0.507
H5 0.000 1.156 1.619
H6 0.000 -1.156 1.619
H7 0.000 1.204 -1.495
H8 0.000 -1.204 -1.495
H9 -0.880 2.318 -0.455
H10 0.880 2.318 -0.455
H11 0.880 -2.318 -0.455
H12 -0.880 -2.318 -0.455

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.35111.51122.60301.09192.07722.19722.84262.16032.16033.30633.3063
C21.35112.60301.51122.07721.09192.84262.19723.30633.30632.16032.1603
C31.51122.60303.32462.18503.52951.08943.03151.09921.09924.07724.0772
C42.60301.51123.32463.52952.18503.03151.08944.07724.07721.09921.0992
H51.09192.07722.18503.52952.31103.11403.90642.53482.53484.14024.1402
H62.07721.09193.52952.18502.31103.90643.11404.14024.14022.53482.5348
H72.19722.84261.08943.03153.11403.90642.40731.76071.76073.77653.7765
H82.84262.19723.03151.08943.90643.11402.40733.77653.77651.76071.7607
H92.16033.30631.09924.07722.53484.14021.76073.77651.76084.95984.6367
H102.16033.30631.09924.07722.53484.14021.76073.77651.76084.63674.9598
H113.30632.16034.07721.09924.14022.53483.77651.76074.95984.63671.7608
H123.30632.16034.07721.09924.14022.53483.77651.76074.63674.95981.7608

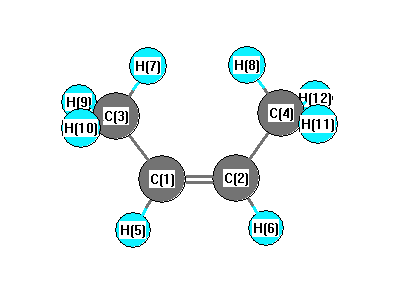
picture of 2-Butene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 130.762 C1 C2 H6 116.074
C1 C3 H7 114.340 C1 C3 H9 110.713
C1 C3 H10 110.713 C2 C1 C3 130.762
C2 C1 H5 116.074 C2 C4 H8 114.340
C2 C4 H11 110.713 C2 C4 H12 110.713
C3 C1 H5 113.164 C4 C2 H6 113.164
H7 C3 H9 107.123 H7 C3 H10 107.123
H8 C4 H11 107.123 H8 C4 H12 107.123
H9 C3 H10 106.439 H11 C4 H12 106.439
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability