return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CCHCH3 (1,2-Butadiene)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-155.448494
Energy at 298.15K-155.453443
HF Energy-154.897660
Nuclear repulsion energy101.908887
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3170 3019 21.69      
2 A' 3167 3016 7.03      
3 A' 3154 3003 6.14      
4 A' 3062 2916 32.17      
5 A' 2079 1979 15.06      
6 A' 1551 1477 2.60      
7 A' 1519 1446 3.39      
8 A' 1463 1393 2.79      
9 A' 1405 1338 6.06      
10 A' 1175 1119 0.05      
11 A' 1124 1070 3.42      
12 A' 901 858 4.45      
13 A' 882 840 50.01      
14 A' 568 540 5.20      
15 A' 209 199 0.74      
16 A" 3242 3088 6.88      
17 A" 3124 2975 24.51      
18 A" 1534 1461 5.39      
19 A" 1094 1041 0.63      
20 A" 1052 1002 0.08      
21 A" 904 861 15.99      
22 A" 540 514 4.78      
23 A" 332 316 4.67      
24 A" 154 147 0.41      

Unscaled Zero Point Vibrational Energy (zpe) 18701.3 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 17809.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
1.13403 0.13867 0.12987

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.686 1.840 0.000
C2 0.000 0.715 0.000
C3 0.687 -0.409 0.000
C4 0.072 -1.790 0.000
H5 -0.985 2.327 0.928
H6 -0.985 2.327 -0.928
H7 1.778 -0.356 0.000
H8 -1.022 -1.729 0.000
H9 0.389 -2.356 -0.886
H10 0.389 -2.356 0.886

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10
C11.31772.63523.70831.08961.08963.30113.58544.42174.4217
C21.31771.31752.50602.10422.10422.07602.64963.22033.2203
C32.63521.31751.51113.33763.33761.09292.15902.15962.1596
C43.70832.50601.51114.35014.35012.22851.09541.09821.0982
H51.08962.10423.33764.35011.85633.96134.16125.20654.8806
H61.08962.10423.33764.35011.85633.96134.16124.88065.2065
H73.30112.07601.09292.22853.96133.96133.11862.59122.5912
H83.58542.64962.15901.09544.16124.16123.11861.77991.7799
H94.42173.22032.15961.09825.20654.88062.59121.77991.7713
H104.42173.22032.15961.09824.88065.20652.59121.77991.7713

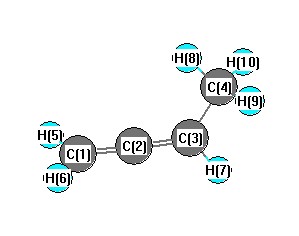
picture of 1,2-Butadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 179.977 C2 C1 H5 121.588
C2 C1 H6 121.588 C2 C3 C4 124.597
C2 C3 H7 118.624 C3 C4 H8 110.842
C3 C4 H9 110.724 C3 C4 H10 110.724
C4 C3 H7 116.779 H5 C1 H6 116.824
H8 C4 H9 108.470 H8 C4 H10 108.470
H9 C4 H10 107.504
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability