return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,4-Pentadiene)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-195.045806
Energy at 298.15K-195.053345
HF Energy-194.834278
Nuclear repulsion energy155.110407
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3273 3106 17.30      
2 A 3193 3030 2.32      
3 A 3182 3021 7.94      
4 A 3056 2901 22.98      
5 A 1737 1649 0.51      
6 A 1523 1446 5.28      
7 A 1485 1410 0.05      
8 A 1343 1275 0.02      
9 A 1289 1223 0.08      
10 A 1105 1049 1.62      
11 A 1041 988 11.15      
12 A 943 895 36.05      
13 A 930 883 4.51      
14 A 685 651 7.90      
15 A 375 356 0.44      
16 A 303 288 0.11      
17 A 89 85 0.01      
18 B 3272 3106 21.45      
19 B 3192 3030 9.52      
20 B 3182 3021 28.91      
21 B 3100 2942 16.94      
22 B 1725 1637 14.27      
23 B 1489 1413 2.57      
24 B 1356 1287 0.52      
25 B 1331 1263 6.20      
26 B 1194 1134 2.70      
27 B 1043 990 14.89      
28 B 986 936 2.23      
29 B 942 894 33.92      
30 B 915 868 7.89      
31 B 617 586 9.98      
32 B 458 435 2.65      
33 B 103 97 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 25228.3 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 23946.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.66180 0.07828 0.07768

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.669
C2 0.000 1.249 -0.177
C3 0.000 -1.249 -0.177
C4 0.970 2.166 -0.179
C5 -0.970 -2.166 -0.179
H6 -0.883 -0.010 1.321
H7 0.883 0.010 1.321
H8 -0.859 1.373 -0.834
H9 0.859 -1.373 -0.834
H10 0.924 3.044 -0.815
H11 -0.924 -3.044 -0.815
H12 1.844 2.072 0.459
H13 -1.844 -2.072 0.459

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.50781.50782.52032.52031.09731.09732.20952.20953.51003.51002.78142.7814
C21.50782.49721.33503.54982.14582.13401.08872.83622.11754.43692.11683.8509
C31.50782.49723.54981.33502.13402.14582.83621.08874.43692.11753.85092.1168
C42.52031.33503.54984.74663.22762.62802.09763.60121.08465.57971.08635.1269
C52.52033.54981.33504.74662.62803.22763.60122.09765.57971.08465.12691.0863
H61.09732.14582.13403.22762.62801.76542.56073.08764.14133.71033.53702.4326
H71.09732.13402.14582.62803.22761.76543.08762.56073.71034.14132.43263.5370
H82.20951.08872.83622.09763.60122.56073.08763.23922.44294.41763.07613.8096
H92.20952.83621.08873.60122.09763.08762.56073.23924.41762.44293.80963.0761
H103.51002.11754.43691.08465.57974.14133.71032.44294.41766.36181.84765.9543
H113.51004.43692.11755.57971.08463.71034.14134.41762.44296.36185.95431.8476
H122.78142.11683.85091.08635.12693.53702.43263.07613.80961.84765.95435.5471
H132.78143.85092.11685.12691.08632.43263.53703.80963.07615.95431.84765.5471

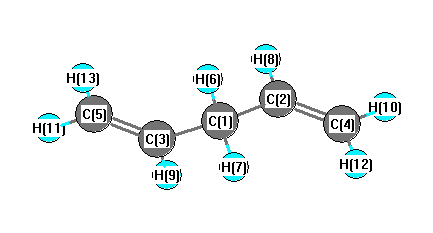
picture of 1,4-Pentadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 124.780 C1 C2 H8 115.689
C1 C3 C5 124.780 C1 C3 H9 115.689
C2 C1 C3 111.810 C2 C1 H6 109.915
C2 C1 H7 108.989 C2 C4 H10 121.781
C2 C4 H12 121.567 C3 C1 H6 108.989
C3 C1 H7 109.915 C3 C5 H11 121.781
C3 C5 H13 121.567 C4 C2 H8 119.522
C5 C3 H9 119.522 H6 C1 H7 107.110
H10 C4 H12 116.651 H11 C5 H13 116.651
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.368      
2 C -0.076      
3 C -0.076      
4 C -0.366      
5 C -0.366      
6 H 0.162      
7 H 0.162      
8 H 0.156      
9 H 0.156      
10 H 0.157      
11 H 0.157      
12 H 0.151      
13 H 0.151      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 7.783 2.991 0.000
y 2.991 9.633 0.000
z 0.000 0.000 5.321


<r2> (average value of r2) Å2
<r2> 158.453
(<r2>)1/2 12.588