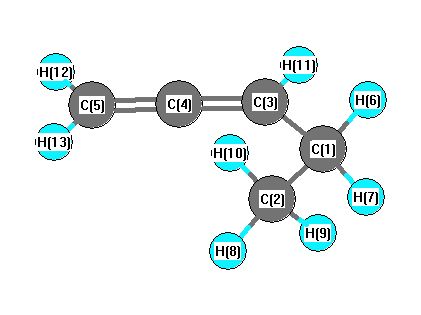
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3222 |
3082 |
5.34 |
|
|
|
| 2 |
A |
3151 |
3014 |
7.33 |
|
|
|
| 3 |
A |
3146 |
3009 |
13.21 |
|
|
|
| 4 |
A |
3143 |
3007 |
29.15 |
|
|
|
| 5 |
A |
3142 |
3006 |
30.13 |
|
|
|
| 6 |
A |
3065 |
2932 |
20.62 |
|
|
|
| 7 |
A |
3060 |
2928 |
20.47 |
|
|
|
| 8 |
A |
3032 |
2901 |
38.29 |
|
|
|
| 9 |
A |
2083 |
1993 |
27.70 |
|
|
|
| 10 |
A |
1528 |
1461 |
5.41 |
|
|
|
| 11 |
A |
1519 |
1454 |
7.48 |
|
|
|
| 12 |
A |
1512 |
1447 |
0.27 |
|
|
|
| 13 |
A |
1497 |
1433 |
3.32 |
|
|
|
| 14 |
A |
1438 |
1376 |
4.99 |
|
|
|
| 15 |
A |
1389 |
1329 |
15.02 |
|
|
|
| 16 |
A |
1362 |
1303 |
2.41 |
|
|
|
| 17 |
A |
1297 |
1241 |
0.02 |
|
|
|
| 18 |
A |
1168 |
1118 |
1.68 |
|
|
|
| 19 |
A |
1111 |
1063 |
2.19 |
|
|
|
| 20 |
A |
1102 |
1054 |
3.07 |
|
|
|
| 21 |
A |
1033 |
989 |
0.00 |
|
|
|
| 22 |
A |
1009 |
965 |
0.70 |
|
|
|
| 23 |
A |
907 |
868 |
13.64 |
|
|
|
| 24 |
A |
871 |
833 |
49.17 |
|
|
|
| 25 |
A |
863 |
826 |
11.87 |
|
|
|
| 26 |
A |
796 |
761 |
3.77 |
|
|
|
| 27 |
A |
643 |
615 |
1.75 |
|
|
|
| 28 |
A |
527 |
504 |
4.68 |
|
|
|
| 29 |
A |
364 |
348 |
2.77 |
|
|
|
| 30 |
A |
348 |
332 |
5.57 |
|
|
|
| 31 |
A |
248 |
237 |
0.07 |
|
|
|
| 32 |
A |
152 |
146 |
0.22 |
|
|
|
| 33 |
A |
99 |
95 |
0.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24913.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 23834.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.328 |
|
|
0.196 |
| 2 |
C |
-0.514 |
|
|
-0.342 |
| 3 |
C |
-0.336 |
|
|
-0.314 |
| 4 |
C |
0.340 |
|
|
0.161 |
| 5 |
C |
-0.582 |
|
|
-0.559 |
| 6 |
H |
0.175 |
|
|
0.008 |
| 7 |
H |
0.175 |
|
|
0.008 |
| 8 |
H |
0.177 |
|
|
0.099 |
| 9 |
H |
0.166 |
|
|
0.077 |
| 10 |
H |
0.177 |
|
|
0.099 |
| 11 |
H |
0.179 |
|
|
0.139 |
| 12 |
H |
0.186 |
|
|
0.214 |
| 13 |
H |
0.186 |
|
|
0.214 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.422 |
0.076 |
0.000 |
0.428 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.385 |
0.056 |
0.000 |
0.389 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.693 |
1.770 |
0.000 |
| y |
1.770 |
-30.840 |
0.001 |
| z |
0.000 |
0.001 |
-31.363 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.409 |
1.770 |
0.000 |
| y |
1.770 |
-0.313 |
0.001 |
| z |
0.000 |
0.001 |
-1.096 |
|
| Polar |
|---|
| 3z2-r2 | -2.192 |
|---|
| x2-y2 | 1.148 |
|---|
| xy | 1.770 |
|---|
| xz | 0.000 |
|---|
| yz | 0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
144.961 |
| (<r2>)1/2 |
12.040 |