return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,2-Pentadiene)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-195.035669
Energy at 298.15K 
HF Energy-194.823344
Nuclear repulsion energy156.408324
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3174 3013 6.48      
2 A' 3168 3007 12.88      
3 A' 3163 3002 31.89      
4 A' 3088 2931 19.25      
5 A' 3058 2903 37.72      
6 A' 2071 1966 19.68      
7 A' 1553 1474 3.86      
8 A' 1537 1459 0.19      
9 A' 1519 1442 2.07      
10 A' 1462 1388 3.66      
11 A' 1411 1339 17.17      
12 A' 1377 1307 1.71      
13 A' 1173 1114 0.98      
14 A' 1114 1057 3.03      
15 A' 1015 964 0.35      
16 A' 881 836 50.31      
17 A' 866 822 5.01      
18 A' 643 611 1.75      
19 A' 367 348 2.46      
20 A' 155 147 0.17      
21 A" 3248 3083 5.49      
22 A" 3165 3005 30.46      
23 A" 3086 2930 21.70      
24 A" 1546 1467 6.15      
25 A" 1317 1250 0.05      
26 A" 1133 1075 2.03      
27 A" 1050 997 0.03      
28 A" 918 871 12.57      
29 A" 810 768 3.01      
30 A" 529 503 4.35      
31 A" 343 326 4.69      
32 A" 258 245 0.05      
33 A" 97 92 0.20      

Unscaled Zero Point Vibrational Energy (zpe) 25146.9 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 23869.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.401 0.447 0.000
C2 -1.460 -1.079 0.000
C3 0.000 1.019 0.000
C4 1.108 0.318 0.000
C5 2.220 -0.376 0.000
H6 -1.935 0.841 0.874
H7 -1.935 0.841 -0.874
H8 -0.962 -1.490 -0.882
H9 -2.497 -1.425 0.000
H10 -0.962 -1.490 0.882
H11 0.078 2.106 0.000
H12 2.703 -0.678 -0.925
H13 2.703 -0.678 0.925

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.52661.51302.51203.71231.09761.09762.17282.16882.17282.22274.35434.3543
C21.52662.55562.92323.74552.16212.16211.09311.09321.09313.53674.28314.2831
C31.51302.55561.31112.62162.13082.13082.82813.49362.82811.08993.32293.3229
C42.51202.92321.31111.31053.20913.20912.88664.00402.88662.06332.09562.0956
C53.71233.74552.62161.31054.41664.41663.48424.83123.48423.27821.08651.0865
H61.09762.16212.13083.20914.41661.74763.07622.49242.52602.53345.20154.8809
H71.09762.16212.13083.20914.41661.74762.52602.49243.07622.53344.88095.2015
H82.17281.09312.82812.88663.48423.07622.52601.77131.76403.84603.75424.1663
H92.16881.09323.49364.00404.83122.49242.49241.77131.77134.37005.33405.3340
H102.17281.09312.82812.88663.48422.52603.07621.76401.77133.84604.16633.7542
H112.22273.53671.08992.06333.27822.53342.53343.84604.37003.84603.93643.9364
H124.35434.28313.32292.09561.08655.20154.88093.75425.33404.16633.93641.8500
H134.35434.28313.32292.09561.08654.88095.20154.16635.33403.75423.93641.8500

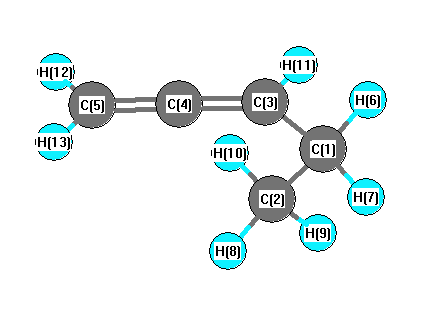
picture of 1,2-Pentadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H8 110.997 C1 C2 H9 110.670
C1 C2 H10 110.997 C1 C3 C4 125.462
C1 C3 H11 116.345 C2 C1 C3 114.446
C2 C1 H6 109.880 C2 C1 H7 109.880
C3 C1 H6 108.362 C3 C1 H7 108.362
C3 C4 C5 179.690 C4 C3 H11 118.193
C4 C5 H12 121.639 C4 C5 H13 121.639
H6 C1 H7 105.515 H8 C2 H9 108.232
H8 C2 H10 107.591 H9 C2 H10 108.232
H12 C5 H13 116.721
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.291      
2 C -0.465      
3 C -0.301      
4 C 0.270      
5 C -0.526      
6 H 0.158      
7 H 0.158      
8 H 0.161      
9 H 0.150      
10 H 0.161      
11 H 0.172      
12 H 0.176      
13 H 0.176      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 10.843 -2.191 0.000
y -2.191 6.953 0.000
z 0.000 0.000 5.398


<r2> (average value of r2) Å2
<r2> 144.992
(<r2>)1/2 12.041