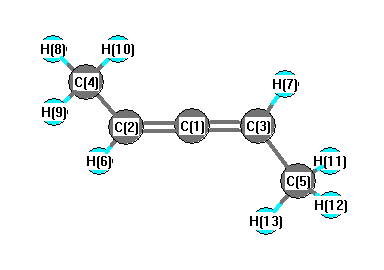
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3172 |
3017 |
15.58 |
|
|
|
| 2 |
A |
3153 |
2999 |
3.73 |
|
|
|
| 3 |
A |
3120 |
2967 |
21.49 |
|
|
|
| 4 |
A |
3060 |
2910 |
7.48 |
|
|
|
| 5 |
A |
1543 |
1467 |
2.83 |
|
|
|
| 6 |
A |
1510 |
1436 |
6.15 |
|
|
|
| 7 |
A |
1475 |
1403 |
0.26 |
|
|
|
| 8 |
A |
1428 |
1358 |
2.04 |
|
|
|
| 9 |
A |
1186 |
1128 |
0.08 |
|
|
|
| 10 |
A |
1108 |
1054 |
1.17 |
|
|
|
| 11 |
A |
1068 |
1016 |
0.00 |
|
|
|
| 12 |
A |
911 |
866 |
22.65 |
|
|
|
| 13 |
A |
842 |
801 |
0.02 |
|
|
|
| 14 |
A |
553 |
526 |
2.75 |
|
|
|
| 15 |
A |
295 |
281 |
1.34 |
|
|
|
| 16 |
A |
179 |
170 |
0.00 |
|
|
|
| 17 |
A |
140 |
133 |
1.53 |
|
|
|
| 18 |
A |
3172 |
3017 |
34.14 |
|
|
|
| 19 |
A |
3152 |
2997 |
3.35 |
|
|
|
| 20 |
A |
3120 |
2967 |
21.47 |
|
|
|
| 21 |
A |
3059 |
2909 |
71.64 |
|
|
|
| 22 |
A |
2106 |
2003 |
11.87 |
|
|
|
| 23 |
A |
1519 |
1445 |
7.17 |
|
|
|
| 24 |
A |
1510 |
1436 |
6.64 |
|
|
|
| 25 |
A |
1426 |
1356 |
4.28 |
|
|
|
| 26 |
A |
1329 |
1264 |
22.71 |
|
|
|
| 27 |
A |
1120 |
1065 |
2.64 |
|
|
|
| 28 |
A |
1065 |
1013 |
0.34 |
|
|
|
| 29 |
A |
981 |
933 |
16.04 |
|
|
|
| 30 |
A |
715 |
680 |
18.25 |
|
|
|
| 31 |
A |
580 |
552 |
11.63 |
|
|
|
| 32 |
A |
231 |
220 |
0.60 |
|
|
|
| 33 |
A |
161 |
153 |
1.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24995.4 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 23770.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.282 |
|
|
|
| 2 |
C |
-0.327 |
|
|
|
| 3 |
C |
-0.327 |
|
|
|
| 4 |
C |
-0.542 |
|
|
|
| 5 |
C |
-0.542 |
|
|
|
| 6 |
H |
0.179 |
|
|
|
| 7 |
H |
0.179 |
|
|
|
| 8 |
H |
0.182 |
|
|
|
| 9 |
H |
0.181 |
|
|
|
| 10 |
H |
0.186 |
|
|
|
| 11 |
H |
0.182 |
|
|
|
| 12 |
H |
0.181 |
|
|
|
| 13 |
H |
0.186 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.192 |
0.192 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.521 |
-0.180 |
0.000 |
| y |
-0.180 |
-27.479 |
0.000 |
| z |
0.000 |
0.000 |
-31.694 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.935 |
-0.180 |
0.000 |
| y |
-0.180 |
4.129 |
0.000 |
| z |
0.000 |
0.000 |
-2.194 |
|
| Polar |
|---|
| 3z2-r2 | -4.387 |
|---|
| x2-y2 | -4.042 |
|---|
| xy | -0.180 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.763 |
-0.996 |
0.000 |
| y |
-0.996 |
13.233 |
0.000 |
| z |
0.000 |
0.000 |
5.661 |
<r2> (average value of r
2) Å
2
| <r2> |
171.551 |
| (<r2>)1/2 |
13.098 |