Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -615.589310 |
| Energy at 298.15K | -615.596534 |
| HF Energy | -615.008891 |
| Nuclear repulsion energy | 206.065141 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3304 |
3052 |
6.79 |
|
|
|
| 2 |
A' |
3258 |
3009 |
17.68 |
|
|
|
| 3 |
A' |
3239 |
2992 |
16.60 |
|
|
|
| 4 |
A' |
3202 |
2958 |
19.26 |
|
|
|
| 5 |
A' |
3153 |
2912 |
26.59 |
|
|
|
| 6 |
A' |
1870 |
1727 |
0.88 |
|
|
|
| 7 |
A' |
1589 |
1468 |
10.21 |
|
|
|
| 8 |
A' |
1576 |
1456 |
2.83 |
|
|
|
| 9 |
A' |
1516 |
1401 |
3.94 |
|
|
|
| 10 |
A' |
1433 |
1323 |
32.35 |
|
|
|
| 11 |
A' |
1408 |
1301 |
4.70 |
|
|
|
| 12 |
A' |
1387 |
1281 |
1.33 |
|
|
|
| 13 |
A' |
1186 |
1096 |
0.25 |
|
|
|
| 14 |
A' |
1112 |
1027 |
9.63 |
|
|
|
| 15 |
A' |
971 |
897 |
7.60 |
|
|
|
| 16 |
A' |
769 |
710 |
20.92 |
|
|
|
| 17 |
A' |
618 |
570 |
3.92 |
|
|
|
| 18 |
A' |
352 |
325 |
1.09 |
|
|
|
| 19 |
A' |
184 |
170 |
0.76 |
|
|
|
| 20 |
A" |
3253 |
3005 |
7.89 |
|
|
|
| 21 |
A" |
3216 |
2971 |
22.24 |
|
|
|
| 22 |
A" |
1575 |
1454 |
6.55 |
|
|
|
| 23 |
A" |
1284 |
1186 |
2.48 |
|
|
|
| 24 |
A" |
1141 |
1054 |
1.04 |
|
|
|
| 25 |
A" |
1055 |
974 |
36.12 |
|
|
|
| 26 |
A" |
1020 |
942 |
1.19 |
|
|
|
| 27 |
A" |
774 |
715 |
0.27 |
|
|
|
| 28 |
A" |
248 |
229 |
4.96 |
|
|
|
| 29 |
A" |
204 |
188 |
0.80 |
|
|
|
| 30 |
A" |
107 |
98 |
1.67 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23001.0 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 21246.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.311 |
0.282 |
0.000 |
| C2 |
0.000 |
1.007 |
0.000 |
| C3 |
1.216 |
0.484 |
0.000 |
| C4 |
2.478 |
1.291 |
0.000 |
| Cl5 |
-1.186 |
-1.498 |
0.000 |
| H6 |
-1.893 |
0.550 |
0.878 |
| H7 |
-1.893 |
0.550 |
-0.878 |
| H8 |
-0.118 |
2.086 |
0.000 |
| H9 |
1.331 |
-0.592 |
0.000 |
| H10 |
2.269 |
2.360 |
0.000 |
| H11 |
3.085 |
1.062 |
0.877 |
| H12 |
3.085 |
1.062 |
-0.877 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4981 | 2.5351 | 3.9211 | 1.7839 | 1.0870 | 1.0870 | 2.1627 | 2.7830 | 4.1395 | 4.5499 | 4.5499 |
C2 | 1.4981 | | 1.3238 | 2.4943 | 2.7713 | 2.1360 | 2.1360 | 1.0855 | 2.0803 | 2.6419 | 3.2075 | 3.2075 | C3 | 2.5351 | 1.3238 | | 1.4980 | 3.1142 | 3.2313 | 3.2313 | 2.0848 | 1.0817 | 2.1511 | 2.1438 | 2.1438 | C4 | 3.9211 | 2.4943 | 1.4980 | | 4.6048 | 4.5194 | 4.5194 | 2.7152 | 2.2044 | 1.0886 | 1.0906 | 1.0906 | Cl5 | 1.7839 | 2.7713 | 3.1142 | 4.6048 | | 2.3376 | 2.3376 | 3.7393 | 2.6757 | 5.1787 | 5.0562 | 5.0562 | H6 | 1.0870 | 2.1360 | 3.2313 | 4.5194 | 2.3376 | | 1.7561 | 2.5057 | 3.5314 | 4.6226 | 5.0040 | 5.3028 | H7 | 1.0870 | 2.1360 | 3.2313 | 4.5194 | 2.3376 | 1.7561 | | 2.5057 | 3.5314 | 4.6226 | 5.3028 | 5.0040 | H8 | 2.1627 | 1.0855 | 2.0848 | 2.7152 | 3.7393 | 2.5057 | 2.5057 | | 3.0447 | 2.4032 | 3.4751 | 3.4751 | H9 | 2.7830 | 2.0803 | 1.0817 | 2.2044 | 2.6757 | 3.5314 | 3.5314 | 3.0447 | | 3.0966 | 2.5651 | 2.5651 | H10 | 4.1395 | 2.6419 | 2.1511 | 1.0886 | 5.1787 | 4.6226 | 4.6226 | 2.4032 | 3.0966 | | 1.7654 | 1.7654 | H11 | 4.5499 | 3.2075 | 2.1438 | 1.0906 | 5.0562 | 5.0040 | 5.3028 | 3.4751 | 2.5651 | 1.7654 | | 1.7538 | H12 | 4.5499 | 3.2075 | 2.1438 | 1.0906 | 5.0562 | 5.3028 | 5.0040 | 3.4751 | 2.5651 | 1.7654 | 1.7538 | |
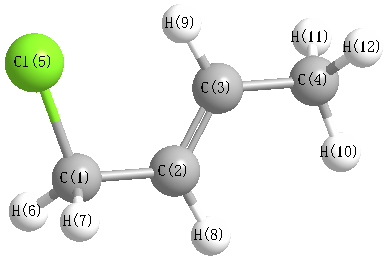 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.787 |
|
C1 |
C2 |
H8 |
112.691 |
| C2 |
C1 |
Cl5 |
114.935 |
|
C2 |
C1 |
H6 |
110.423 |
| C2 |
C1 |
H7 |
110.423 |
|
C2 |
C3 |
C4 |
124.128 |
| C2 |
C3 |
H9 |
119.385 |
|
C3 |
C2 |
H8 |
119.522 |
| C3 |
C4 |
H10 |
111.550 |
|
C3 |
C4 |
H11 |
110.840 |
| C3 |
C4 |
H12 |
110.840 |
|
C4 |
C3 |
H9 |
116.486 |
| Cl5 |
C1 |
H6 |
106.485 |
|
Cl5 |
C1 |
H7 |
106.485 |
| H6 |
C1 |
H7 |
107.751 |
|
H10 |
C4 |
H11 |
108.207 |
| H10 |
C4 |
H12 |
108.207 |
|
H11 |
C4 |
H12 |
107.033 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -615.591427 |
| Energy at 298.15K | -615.598628 |
| HF Energy | -615.011510 |
| Nuclear repulsion energy | 201.974519 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3290 |
3039 |
17.25 |
|
|
|
| 2 |
A |
3277 |
3027 |
1.94 |
|
|
|
| 3 |
A |
3264 |
3015 |
24.29 |
|
|
|
| 4 |
A |
3242 |
2995 |
7.85 |
|
|
|
| 5 |
A |
3218 |
2973 |
17.57 |
|
|
|
| 6 |
A |
3217 |
2972 |
20.94 |
|
|
|
| 7 |
A |
3154 |
2913 |
24.20 |
|
|
|
| 8 |
A |
1858 |
1717 |
7.92 |
|
|
|
| 9 |
A |
1587 |
1466 |
11.43 |
|
|
|
| 10 |
A |
1579 |
1459 |
2.72 |
|
|
|
| 11 |
A |
1574 |
1454 |
7.41 |
|
|
|
| 12 |
A |
1515 |
1400 |
2.76 |
|
|
|
| 13 |
A |
1433 |
1324 |
16.18 |
|
|
|
| 14 |
A |
1409 |
1301 |
2.27 |
|
|
|
| 15 |
A |
1378 |
1273 |
36.94 |
|
|
|
| 16 |
A |
1281 |
1183 |
1.52 |
|
|
|
| 17 |
A |
1195 |
1104 |
0.72 |
|
|
|
| 18 |
A |
1155 |
1067 |
2.53 |
|
|
|
| 19 |
A |
1140 |
1053 |
1.22 |
|
|
|
| 20 |
A |
1058 |
977 |
36.31 |
|
|
|
| 21 |
A |
1011 |
934 |
7.39 |
|
|
|
| 22 |
A |
947 |
874 |
1.75 |
|
|
|
| 23 |
A |
852 |
787 |
19.56 |
|
|
|
| 24 |
A |
757 |
699 |
46.39 |
|
|
|
| 25 |
A |
515 |
476 |
1.67 |
|
|
|
| 26 |
A |
376 |
347 |
2.40 |
|
|
|
| 27 |
A |
289 |
267 |
2.15 |
|
|
|
| 28 |
A |
209 |
193 |
0.95 |
|
|
|
| 29 |
A |
165 |
152 |
1.90 |
|
|
|
| 30 |
A |
89 |
82 |
0.44 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23016.3 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 21260.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.779 |
0.813 |
0.126 |
| C2 |
0.517 |
0.152 |
0.447 |
| C3 |
1.595 |
0.250 |
-0.319 |
| C4 |
2.919 |
-0.380 |
-0.016 |
| Cl5 |
-2.093 |
-0.391 |
-0.109 |
| H6 |
-1.112 |
1.460 |
0.931 |
| H7 |
-0.713 |
1.393 |
-0.789 |
| H8 |
0.546 |
-0.423 |
1.364 |
| H9 |
1.536 |
0.822 |
-1.239 |
| H10 |
2.887 |
-0.938 |
0.918 |
| H11 |
3.214 |
-1.064 |
-0.813 |
| H12 |
3.702 |
0.376 |
0.064 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4901 | 2.4803 | 3.8881 | 1.7976 | 1.0858 | 1.0849 | 2.1953 | 2.6875 | 4.1386 | 4.5102 | 4.5021 |
C2 | 1.4901 | | 1.3261 | 2.5031 | 2.7236 | 2.1449 | 2.1402 | 1.0833 | 2.0809 | 2.6500 | 3.2147 | 3.2149 | C3 | 2.4803 | 1.3261 | | 1.4968 | 3.7493 | 3.2185 | 2.6182 | 2.0947 | 1.0851 | 2.1465 | 2.1418 | 2.1445 | C4 | 3.8881 | 2.5031 | 1.4968 | | 5.0128 | 4.5313 | 4.1146 | 2.7453 | 2.2028 | 1.0882 | 1.0905 | 1.0906 | Cl5 | 1.7976 | 2.7236 | 3.7493 | 5.0128 | | 2.3393 | 2.3555 | 3.0230 | 3.9898 | 5.1137 | 5.3949 | 5.8475 | H6 | 1.0858 | 2.1449 | 3.2185 | 4.5313 | 2.3393 | | 1.7670 | 2.5468 | 3.4830 | 4.6625 | 5.3030 | 5.0098 | H7 | 1.0849 | 2.1402 | 2.6182 | 4.1146 | 2.3555 | 1.7670 | | 3.0855 | 2.3638 | 4.6152 | 4.6317 | 4.6096 | H8 | 2.1953 | 1.0833 | 2.0947 | 2.7453 | 3.0230 | 2.5468 | 3.0855 | | 3.0511 | 2.4373 | 3.5018 | 3.5049 | H9 | 2.6875 | 2.0809 | 1.0851 | 2.2028 | 3.9898 | 3.4830 | 2.3638 | 3.0511 | | 3.0939 | 2.5595 | 2.5662 | H10 | 4.1386 | 2.6500 | 2.1465 | 1.0882 | 5.1137 | 4.6625 | 4.6152 | 2.4373 | 3.0939 | | 1.7656 | 1.7659 | H11 | 4.5102 | 3.2147 | 2.1418 | 1.0905 | 5.3949 | 5.3030 | 4.6317 | 3.5018 | 2.5595 | 1.7656 | | 1.7548 | H12 | 4.5021 | 3.2149 | 2.1445 | 1.0906 | 5.8475 | 5.0098 | 4.6096 | 3.5049 | 2.5662 | 1.7659 | 1.7548 | |
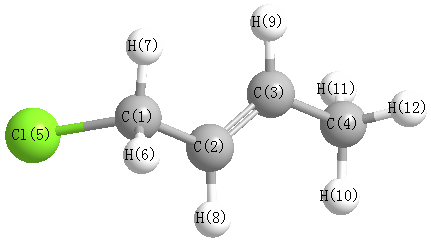 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.357 |
|
C1 |
C2 |
H8 |
116.203 |
| C2 |
C1 |
Cl5 |
111.535 |
|
C2 |
C1 |
H6 |
111.795 |
| C2 |
C1 |
H7 |
111.466 |
|
C2 |
C3 |
C4 |
124.812 |
| C2 |
C3 |
H9 |
118.978 |
|
C3 |
C2 |
H8 |
120.438 |
| C3 |
C4 |
H10 |
111.287 |
|
C3 |
C4 |
H11 |
110.770 |
| C3 |
C4 |
H12 |
110.975 |
|
C4 |
C3 |
H9 |
116.210 |
| Cl5 |
C1 |
H6 |
105.783 |
|
Cl5 |
C1 |
H7 |
106.994 |
| H6 |
C1 |
H7 |
108.991 |
|
H10 |
C4 |
H11 |
108.258 |
| H10 |
C4 |
H12 |
108.277 |
|
H11 |
C4 |
H12 |
107.123 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
