Jump to
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -616.707858 |
| Energy at 298.15K | -616.714920 |
| HF Energy | -616.707858 |
| Nuclear repulsion energy | 205.126364 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3188 |
3050 |
9.34 |
|
|
|
| 2 |
A' |
3146 |
3010 |
16.47 |
|
|
|
| 3 |
A' |
3135 |
3000 |
15.79 |
|
|
|
| 4 |
A' |
3081 |
2948 |
21.24 |
|
|
|
| 5 |
A' |
3045 |
2913 |
29.31 |
|
|
|
| 6 |
A' |
1778 |
1701 |
0.83 |
|
|
|
| 7 |
A' |
1517 |
1451 |
11.72 |
|
|
|
| 8 |
A' |
1497 |
1432 |
2.66 |
|
|
|
| 9 |
A' |
1438 |
1376 |
3.01 |
|
|
|
| 10 |
A' |
1349 |
1291 |
22.37 |
|
|
|
| 11 |
A' |
1339 |
1281 |
16.89 |
|
|
|
| 12 |
A' |
1313 |
1256 |
2.81 |
|
|
|
| 13 |
A' |
1137 |
1088 |
0.17 |
|
|
|
| 14 |
A' |
1061 |
1015 |
12.37 |
|
|
|
| 15 |
A' |
925 |
885 |
10.93 |
|
|
|
| 16 |
A' |
723 |
692 |
21.95 |
|
|
|
| 17 |
A' |
589 |
564 |
5.00 |
|
|
|
| 18 |
A' |
336 |
321 |
1.17 |
|
|
|
| 19 |
A' |
171 |
164 |
0.87 |
|
|
|
| 20 |
A" |
3127 |
2992 |
7.63 |
|
|
|
| 21 |
A" |
3102 |
2968 |
20.10 |
|
|
|
| 22 |
A" |
1504 |
1439 |
7.41 |
|
|
|
| 23 |
A" |
1215 |
1163 |
2.05 |
|
|
|
| 24 |
A" |
1076 |
1030 |
0.03 |
|
|
|
| 25 |
A" |
999 |
956 |
35.46 |
|
|
|
| 26 |
A" |
959 |
917 |
0.00 |
|
|
|
| 27 |
A" |
730 |
699 |
0.43 |
|
|
|
| 28 |
A" |
241 |
231 |
4.59 |
|
|
|
| 29 |
A" |
207 |
198 |
2.11 |
|
|
|
| 30 |
A" |
114 |
109 |
1.54 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22022.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 21068.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.313 |
0.295 |
0.000 |
| C2 |
0.000 |
1.015 |
0.000 |
| C3 |
1.223 |
0.485 |
0.000 |
| C4 |
2.492 |
1.278 |
0.000 |
| Cl5 |
-1.196 |
-1.500 |
0.000 |
| H6 |
-1.903 |
0.562 |
0.883 |
| H7 |
-1.903 |
0.562 |
-0.883 |
| H8 |
-0.116 |
2.101 |
0.000 |
| H9 |
1.323 |
-0.600 |
0.000 |
| H10 |
2.301 |
2.356 |
0.000 |
| H11 |
3.105 |
1.040 |
0.880 |
| H12 |
3.105 |
1.040 |
-0.880 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4968 | 2.5424 | 3.9299 | 1.7988 | 1.0953 | 1.0953 | 2.1662 | 2.7835 | 4.1601 | 4.5652 | 4.5652 |
C2 | 1.4968 | | 1.3323 | 2.5063 | 2.7842 | 2.1460 | 2.1460 | 1.0926 | 2.0871 | 2.6636 | 3.2270 | 3.2270 | C3 | 2.5424 | 1.3323 | | 1.4968 | 3.1286 | 3.2487 | 3.2487 | 2.0984 | 1.0894 | 2.1594 | 2.1504 | 2.1504 | C4 | 3.9299 | 2.5063 | 1.4968 | | 4.6170 | 4.5399 | 4.5399 | 2.7354 | 2.2116 | 1.0951 | 1.0980 | 1.0980 | Cl5 | 1.7988 | 2.7842 | 3.1286 | 4.6170 | | 2.3520 | 2.3520 | 3.7591 | 2.6749 | 5.2053 | 5.0713 | 5.0713 | H6 | 1.0953 | 2.1460 | 3.2487 | 4.5399 | 2.3520 | | 1.7665 | 2.5180 | 3.5407 | 4.6553 | 5.0301 | 5.3302 | H7 | 1.0953 | 2.1460 | 3.2487 | 4.5399 | 2.3520 | 1.7665 | | 2.5180 | 3.5407 | 4.6553 | 5.3302 | 5.0301 | H8 | 2.1662 | 1.0926 | 2.0984 | 2.7354 | 3.7591 | 2.5180 | 2.5180 | | 3.0601 | 2.4308 | 3.5032 | 3.5032 | H9 | 2.7835 | 2.0871 | 1.0894 | 2.2116 | 2.6749 | 3.5407 | 3.5407 | 3.0601 | | 3.1131 | 2.5762 | 2.5762 | H10 | 4.1601 | 2.6636 | 2.1594 | 1.0951 | 5.2053 | 4.6553 | 4.6553 | 2.4308 | 3.1131 | | 1.7751 | 1.7751 | H11 | 4.5652 | 3.2270 | 2.1504 | 1.0980 | 5.0713 | 5.0301 | 5.3302 | 3.5032 | 2.5762 | 1.7751 | | 1.7601 | H12 | 4.5652 | 3.2270 | 2.1504 | 1.0980 | 5.0713 | 5.3302 | 5.0301 | 3.5032 | 2.5762 | 1.7751 | 1.7601 | |
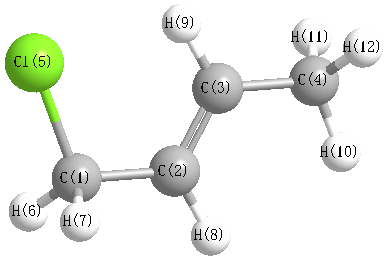 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.862 |
|
C1 |
C2 |
H8 |
112.622 |
| C2 |
C1 |
Cl5 |
114.997 |
|
C2 |
C1 |
H6 |
110.817 |
| C2 |
C1 |
H7 |
110.817 |
|
C2 |
C3 |
C4 |
124.617 |
| C2 |
C3 |
H9 |
118.708 |
|
C3 |
C2 |
H8 |
119.516 |
| C3 |
C4 |
H10 |
111.913 |
|
C3 |
C4 |
H11 |
111.004 |
| C3 |
C4 |
H12 |
111.004 |
|
C4 |
C3 |
H9 |
116.675 |
| Cl5 |
C1 |
H6 |
106.155 |
|
Cl5 |
C1 |
H7 |
106.155 |
| H6 |
C1 |
H7 |
107.488 |
|
H10 |
C4 |
H11 |
108.077 |
| H10 |
C4 |
H12 |
108.077 |
|
H11 |
C4 |
H12 |
106.550 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.467 |
|
|
|
| 2 |
C |
-0.137 |
|
|
|
| 3 |
C |
-0.100 |
|
|
|
| 4 |
C |
-0.560 |
|
|
|
| 5 |
Cl |
-0.071 |
|
|
|
| 6 |
H |
0.224 |
|
|
|
| 7 |
H |
0.224 |
|
|
|
| 8 |
H |
0.162 |
|
|
|
| 9 |
H |
0.182 |
|
|
|
| 10 |
H |
0.176 |
|
|
|
| 11 |
H |
0.184 |
|
|
|
| 12 |
H |
0.184 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.345 |
2.022 |
0.000 |
2.051 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.995 |
-2.610 |
0.000 |
| y |
-2.610 |
-38.098 |
0.000 |
| z |
0.000 |
0.000 |
-38.948 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.528 |
-2.610 |
0.000 |
| y |
-2.610 |
-1.127 |
0.000 |
| z |
0.000 |
0.000 |
-2.401 |
|
| Polar |
|---|
| 3z2-r2 | -4.802 |
|---|
| x2-y2 | 3.103 |
|---|
| xy | -2.610 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.803 |
0.435 |
0.000 |
| y |
0.435 |
8.357 |
0.000 |
| z |
0.000 |
0.000 |
4.871 |
<r2> (average value of r
2) Å
2
| <r2> |
210.164 |
| (<r2>)1/2 |
14.497 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -616.710189 |
| Energy at 298.15K | -616.717210 |
| HF Energy | -616.710189 |
| Nuclear repulsion energy | 200.781984 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3183 |
3045 |
20.33 |
|
|
|
| 2 |
A |
3166 |
3029 |
1.61 |
|
|
|
| 3 |
A |
3156 |
3020 |
29.21 |
|
|
|
| 4 |
A |
3139 |
3003 |
2.85 |
|
|
|
| 5 |
A |
3106 |
2971 |
17.21 |
|
|
|
| 6 |
A |
3103 |
2969 |
17.96 |
|
|
|
| 7 |
A |
3046 |
2914 |
24.92 |
|
|
|
| 8 |
A |
1760 |
1684 |
14.10 |
|
|
|
| 9 |
A |
1514 |
1449 |
14.17 |
|
|
|
| 10 |
A |
1504 |
1439 |
2.59 |
|
|
|
| 11 |
A |
1503 |
1438 |
9.00 |
|
|
|
| 12 |
A |
1436 |
1374 |
2.28 |
|
|
|
| 13 |
A |
1356 |
1298 |
6.58 |
|
|
|
| 14 |
A |
1343 |
1285 |
1.38 |
|
|
|
| 15 |
A |
1297 |
1241 |
44.09 |
|
|
|
| 16 |
A |
1218 |
1165 |
3.46 |
|
|
|
| 17 |
A |
1136 |
1087 |
0.66 |
|
|
|
| 18 |
A |
1112 |
1064 |
2.95 |
|
|
|
| 19 |
A |
1074 |
1028 |
0.35 |
|
|
|
| 20 |
A |
1008 |
965 |
32.72 |
|
|
|
| 21 |
A |
961 |
920 |
12.24 |
|
|
|
| 22 |
A |
898 |
859 |
3.26 |
|
|
|
| 23 |
A |
803 |
768 |
13.39 |
|
|
|
| 24 |
A |
689 |
659 |
67.02 |
|
|
|
| 25 |
A |
493 |
472 |
1.81 |
|
|
|
| 26 |
A |
353 |
338 |
4.92 |
|
|
|
| 27 |
A |
277 |
265 |
2.14 |
|
|
|
| 28 |
A |
213 |
204 |
1.16 |
|
|
|
| 29 |
A |
158 |
152 |
2.64 |
|
|
|
| 30 |
A |
89 |
85 |
0.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22047.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 21092.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.772 |
0.827 |
0.103 |
| C2 |
0.522 |
0.184 |
0.453 |
| C3 |
1.602 |
0.226 |
-0.332 |
| C4 |
2.928 |
-0.382 |
-0.006 |
| Cl5 |
-2.105 |
-0.400 |
-0.099 |
| H6 |
-1.126 |
1.500 |
0.888 |
| H7 |
-0.713 |
1.377 |
-0.839 |
| H8 |
0.560 |
-0.335 |
1.411 |
| H9 |
1.530 |
0.742 |
-1.292 |
| H10 |
2.916 |
-0.883 |
0.967 |
| H11 |
3.218 |
-1.119 |
-0.766 |
| H12 |
3.719 |
0.379 |
0.011 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4870 | 2.4874 | 3.8948 | 1.8222 | 1.0932 | 1.0920 | 2.1996 | 2.6938 | 4.1565 | 4.5242 | 4.5150 |
C2 | 1.4870 | | 1.3360 | 2.5144 | 2.7464 | 2.1536 | 2.1489 | 1.0899 | 2.0916 | 2.6712 | 3.2336 | 3.2340 | C3 | 2.4874 | 1.3360 | | 1.4953 | 3.7658 | 3.2489 | 2.6344 | 2.1063 | 1.0920 | 2.1554 | 2.1472 | 2.1508 | C4 | 3.8948 | 2.5144 | 1.4953 | | 5.0336 | 4.5587 | 4.1290 | 2.7595 | 2.2071 | 1.0946 | 1.0980 | 1.0980 | Cl5 | 1.8222 | 2.7464 | 3.7658 | 5.0336 | | 2.3540 | 2.3745 | 3.0636 | 3.9924 | 5.1551 | 5.4123 | 5.8767 | H6 | 1.0932 | 2.1536 | 3.2489 | 4.5587 | 2.3540 | | 1.7803 | 2.5470 | 3.5196 | 4.6932 | 5.3360 | 5.0505 | H7 | 1.0920 | 2.1489 | 2.6344 | 4.1290 | 2.3745 | 1.7803 | | 3.1007 | 2.3755 | 4.6412 | 4.6573 | 4.6223 | H8 | 2.1996 | 1.0899 | 2.1063 | 2.7595 | 3.0636 | 2.5470 | 3.1007 | | 3.0668 | 2.4586 | 3.5238 | 3.5282 | H9 | 2.6938 | 2.0916 | 1.0920 | 2.2071 | 3.9924 | 3.5196 | 2.3755 | 3.0668 | | 3.1088 | 2.5666 | 2.5730 | H10 | 4.1565 | 2.6712 | 2.1554 | 1.0946 | 5.1551 | 4.6932 | 4.6412 | 2.4586 | 3.1088 | | 1.7756 | 1.7759 | H11 | 4.5242 | 3.2336 | 2.1472 | 1.0980 | 5.4123 | 5.3360 | 4.6573 | 3.5238 | 2.5666 | 1.7756 | | 1.7606 | H12 | 4.5150 | 3.2340 | 2.1508 | 1.0980 | 5.8767 | 5.0505 | 4.6223 | 3.5282 | 2.5730 | 1.7759 | 1.7606 | |
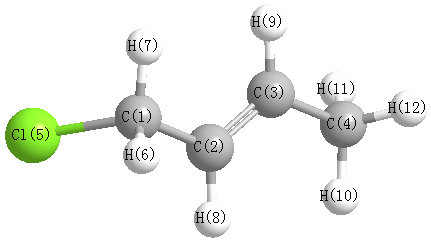 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.463 |
|
C1 |
C2 |
H8 |
116.360 |
| C2 |
C1 |
Cl5 |
111.779 |
|
C2 |
C1 |
H6 |
112.260 |
| C2 |
C1 |
H7 |
111.947 |
|
C2 |
C3 |
C4 |
125.171 |
| C2 |
C3 |
H9 |
118.610 |
|
C3 |
C2 |
H8 |
120.176 |
| C3 |
C4 |
H10 |
111.728 |
|
C3 |
C4 |
H11 |
110.850 |
| C3 |
C4 |
H12 |
111.136 |
|
C4 |
C3 |
H9 |
116.220 |
| Cl5 |
C1 |
H6 |
104.918 |
|
Cl5 |
C1 |
H7 |
106.420 |
| H6 |
C1 |
H7 |
109.119 |
|
H10 |
C4 |
H11 |
108.156 |
| H10 |
C4 |
H12 |
108.182 |
|
H11 |
C4 |
H12 |
106.589 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.473 |
|
|
|
| 2 |
C |
-0.100 |
|
|
|
| 3 |
C |
-0.108 |
|
|
|
| 4 |
C |
-0.559 |
|
|
|
| 5 |
Cl |
-0.089 |
|
|
|
| 6 |
H |
0.224 |
|
|
|
| 7 |
H |
0.219 |
|
|
|
| 8 |
H |
0.170 |
|
|
|
| 9 |
H |
0.160 |
|
|
|
| 10 |
H |
0.183 |
|
|
|
| 11 |
H |
0.189 |
|
|
|
| 12 |
H |
0.185 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.342 |
1.267 |
0.142 |
2.666 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.292 |
-2.083 |
-1.009 |
| y |
-2.083 |
-37.381 |
-0.823 |
| z |
-1.009 |
-0.823 |
-36.673 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.265 |
-2.083 |
-1.009 |
| y |
-2.083 |
0.601 |
-0.823 |
| z |
-1.009 |
-0.823 |
1.664 |
|
| Polar |
|---|
| 3z2-r2 | 3.327 |
|---|
| x2-y2 | -1.911 |
|---|
| xy | -2.083 |
|---|
| xz | -1.009 |
|---|
| yz | -0.823 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.561 |
0.215 |
-0.839 |
| y |
0.215 |
6.039 |
-0.349 |
| z |
-0.839 |
-0.349 |
6.372 |
<r2> (average value of r
2) Å
2
| <r2> |
232.058 |
| (<r2>)1/2 |
15.233 |
