Jump to
S1C2
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -615.709963 |
| Energy at 298.15K | -615.717040 |
| HF Energy | -615.007762 |
| Nuclear repulsion energy | 204.975849 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3204 |
3051 |
7.74 |
|
|
|
| 2 |
A' |
3157 |
3006 |
18.60 |
|
|
|
| 3 |
A' |
3141 |
2991 |
17.00 |
|
|
|
| 4 |
A' |
3101 |
2953 |
20.45 |
|
|
|
| 5 |
A' |
3058 |
2912 |
26.43 |
|
|
|
| 6 |
A' |
1789 |
1703 |
0.60 |
|
|
|
| 7 |
A' |
1545 |
1472 |
9.24 |
|
|
|
| 8 |
A' |
1531 |
1458 |
2.57 |
|
|
|
| 9 |
A' |
1470 |
1400 |
3.72 |
|
|
|
| 10 |
A' |
1387 |
1321 |
28.21 |
|
|
|
| 11 |
A' |
1361 |
1296 |
3.96 |
|
|
|
| 12 |
A' |
1343 |
1279 |
1.70 |
|
|
|
| 13 |
A' |
1153 |
1098 |
0.14 |
|
|
|
| 14 |
A' |
1083 |
1031 |
8.25 |
|
|
|
| 15 |
A' |
944 |
899 |
6.94 |
|
|
|
| 16 |
A' |
743 |
708 |
17.27 |
|
|
|
| 17 |
A' |
596 |
568 |
3.35 |
|
|
|
| 18 |
A' |
341 |
325 |
0.93 |
|
|
|
| 19 |
A' |
180 |
171 |
0.75 |
|
|
|
| 20 |
A" |
3149 |
2998 |
9.24 |
|
|
|
| 21 |
A" |
3120 |
2972 |
22.73 |
|
|
|
| 22 |
A" |
1530 |
1457 |
5.90 |
|
|
|
| 23 |
A" |
1240 |
1181 |
2.24 |
|
|
|
| 24 |
A" |
1097 |
1045 |
0.39 |
|
|
|
| 25 |
A" |
1003 |
955 |
31.09 |
|
|
|
| 26 |
A" |
984 |
937 |
3.32 |
|
|
|
| 27 |
A" |
733 |
698 |
0.23 |
|
|
|
| 28 |
A" |
238 |
226 |
4.67 |
|
|
|
| 29 |
A" |
194 |
184 |
0.72 |
|
|
|
| 30 |
A" |
101 |
96 |
1.55 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22256.5 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 21194.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.319 |
0.292 |
0.000 |
| C2 |
0.000 |
1.015 |
0.000 |
| C3 |
1.224 |
0.477 |
0.000 |
| C4 |
2.495 |
1.281 |
0.000 |
| Cl5 |
-1.195 |
-1.498 |
0.000 |
| H6 |
-1.905 |
0.563 |
0.885 |
| H7 |
-1.905 |
0.563 |
-0.885 |
| H8 |
-0.111 |
2.102 |
0.000 |
| H9 |
1.332 |
-0.607 |
0.000 |
| H10 |
2.288 |
2.358 |
0.000 |
| H11 |
3.105 |
1.049 |
0.883 |
| H12 |
3.105 |
1.049 |
-0.883 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5040 | 2.5499 | 3.9401 | 1.7938 | 1.0953 | 1.0953 | 2.1765 | 2.7991 | 4.1565 | 4.5747 | 4.5747 |
C2 | 1.5040 | | 1.3371 | 2.5092 | 2.7820 | 2.1482 | 2.1482 | 1.0936 | 2.0987 | 2.6530 | 3.2287 | 3.2287 | C3 | 2.5499 | 1.3371 | | 1.5037 | 3.1229 | 3.2528 | 3.2528 | 2.1038 | 1.0897 | 2.1607 | 2.1554 | 2.1554 | C4 | 3.9401 | 2.5092 | 1.5037 | | 4.6193 | 4.5449 | 4.5449 | 2.7326 | 2.2179 | 1.0966 | 1.0984 | 1.0984 | Cl5 | 1.7938 | 2.7820 | 3.1229 | 4.6193 | | 2.3524 | 2.3524 | 3.7598 | 2.6791 | 5.1957 | 5.0754 | 5.0754 | H6 | 1.0953 | 2.1482 | 3.2528 | 4.5449 | 2.3524 | | 1.7697 | 2.5237 | 3.5536 | 4.6455 | 5.0335 | 5.3351 | H7 | 1.0953 | 2.1482 | 3.2528 | 4.5449 | 2.3524 | 1.7697 | | 2.5237 | 3.5536 | 4.6455 | 5.3351 | 5.0335 | H8 | 2.1765 | 1.0936 | 2.1038 | 2.7326 | 3.7598 | 2.5237 | 2.5237 | | 3.0702 | 2.4125 | 3.4979 | 3.4979 | H9 | 2.7991 | 2.0987 | 1.0897 | 2.2179 | 2.6791 | 3.5536 | 3.5536 | 3.0702 | | 3.1155 | 2.5826 | 2.5826 | H10 | 4.1565 | 2.6530 | 2.1607 | 1.0966 | 5.1957 | 4.6455 | 4.6455 | 2.4125 | 3.1155 | | 1.7780 | 1.7780 | H11 | 4.5747 | 3.2287 | 2.1554 | 1.0984 | 5.0754 | 5.0335 | 5.3351 | 3.4979 | 2.5826 | 1.7780 | | 1.7667 | H12 | 4.5747 | 3.2287 | 2.1554 | 1.0984 | 5.0754 | 5.3351 | 5.0335 | 3.4979 | 2.5826 | 1.7780 | 1.7667 | |
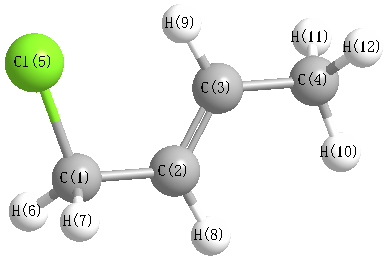 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.571 |
|
C1 |
C2 |
H8 |
112.889 |
| C2 |
C1 |
Cl5 |
114.765 |
|
C2 |
C1 |
H6 |
110.483 |
| C2 |
C1 |
H7 |
110.483 |
|
C2 |
C3 |
C4 |
123.975 |
| C2 |
C3 |
H9 |
119.366 |
|
C3 |
C2 |
H8 |
119.540 |
| C3 |
C4 |
H10 |
111.427 |
|
C3 |
C4 |
H11 |
110.898 |
| C3 |
C4 |
H12 |
110.898 |
|
C4 |
C3 |
H9 |
116.658 |
| Cl5 |
C1 |
H6 |
106.505 |
|
Cl5 |
C1 |
H7 |
106.505 |
| H6 |
C1 |
H7 |
107.770 |
|
H10 |
C4 |
H11 |
108.191 |
| H10 |
C4 |
H12 |
108.191 |
|
H11 |
C4 |
H12 |
107.072 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -615.711837 |
| Energy at 298.15K | -615.718893 |
| HF Energy | -615.010395 |
| Nuclear repulsion energy | 200.807329 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3187 |
3035 |
19.43 |
|
|
|
| 2 |
A |
3173 |
3021 |
2.56 |
|
|
|
| 3 |
A |
3161 |
3011 |
26.73 |
|
|
|
| 4 |
A |
3143 |
2993 |
6.89 |
|
|
|
| 5 |
A |
3122 |
2973 |
20.98 |
|
|
|
| 6 |
A |
3117 |
2968 |
18.42 |
|
|
|
| 7 |
A |
3059 |
2913 |
24.17 |
|
|
|
| 8 |
A |
1779 |
1694 |
6.06 |
|
|
|
| 9 |
A |
1543 |
1470 |
9.92 |
|
|
|
| 10 |
A |
1532 |
1459 |
2.85 |
|
|
|
| 11 |
A |
1530 |
1457 |
6.74 |
|
|
|
| 12 |
A |
1469 |
1399 |
2.70 |
|
|
|
| 13 |
A |
1388 |
1322 |
12.25 |
|
|
|
| 14 |
A |
1362 |
1297 |
2.42 |
|
|
|
| 15 |
A |
1335 |
1271 |
32.99 |
|
|
|
| 16 |
A |
1241 |
1181 |
1.51 |
|
|
|
| 17 |
A |
1159 |
1104 |
0.52 |
|
|
|
| 18 |
A |
1125 |
1071 |
2.30 |
|
|
|
| 19 |
A |
1097 |
1045 |
0.76 |
|
|
|
| 20 |
A |
1006 |
958 |
32.51 |
|
|
|
| 21 |
A |
982 |
935 |
7.43 |
|
|
|
| 22 |
A |
913 |
870 |
1.75 |
|
|
|
| 23 |
A |
811 |
772 |
19.27 |
|
|
|
| 24 |
A |
726 |
691 |
35.55 |
|
|
|
| 25 |
A |
499 |
475 |
1.53 |
|
|
|
| 26 |
A |
362 |
345 |
2.02 |
|
|
|
| 27 |
A |
280 |
267 |
1.97 |
|
|
|
| 28 |
A |
199 |
190 |
0.95 |
|
|
|
| 29 |
A |
160 |
152 |
1.73 |
|
|
|
| 30 |
A |
84 |
80 |
0.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22270.5 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 21208.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.783 |
0.819 |
0.128 |
| C2 |
0.517 |
0.153 |
0.451 |
| C3 |
1.606 |
0.253 |
-0.322 |
| C4 |
2.933 |
-0.382 |
-0.016 |
| Cl5 |
-2.103 |
-0.393 |
-0.111 |
| H6 |
-1.119 |
1.470 |
0.940 |
| H7 |
-0.714 |
1.403 |
-0.794 |
| H8 |
0.545 |
-0.430 |
1.374 |
| H9 |
1.548 |
0.832 |
-1.248 |
| H10 |
2.896 |
-0.946 |
0.923 |
| H11 |
3.230 |
-1.071 |
-0.819 |
| H12 |
3.722 |
0.377 |
0.069 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4956 | 2.4957 | 3.9080 | 1.8080 | 1.0939 | 1.0933 | 2.2073 | 2.7061 | 4.1568 | 4.5355 | 4.5272 |
C2 | 1.4956 | | 1.3392 | 2.5187 | 2.7343 | 2.1567 | 2.1512 | 1.0915 | 2.0997 | 2.6627 | 3.2360 | 3.2361 | C3 | 2.4957 | 1.3392 | | 1.5030 | 3.7706 | 3.2403 | 2.6322 | 2.1136 | 1.0934 | 2.1568 | 2.1534 | 2.1559 | C4 | 3.9080 | 2.5187 | 1.5030 | | 5.0371 | 4.5572 | 4.1348 | 2.7640 | 2.2160 | 1.0963 | 1.0983 | 1.0984 | Cl5 | 1.8080 | 2.7343 | 3.7706 | 5.0371 | | 2.3547 | 2.3709 | 3.0355 | 4.0150 | 5.1344 | 5.4225 | 5.8788 | H6 | 1.0939 | 2.1567 | 3.2403 | 4.5572 | 2.3547 | | 1.7815 | 2.5623 | 3.5077 | 4.6859 | 5.3354 | 5.0393 | H7 | 1.0933 | 2.1512 | 2.6322 | 4.1348 | 2.3709 | 1.7815 | | 3.1049 | 2.3766 | 4.6366 | 4.6560 | 4.6347 | H8 | 2.2073 | 1.0915 | 2.1136 | 2.7640 | 3.0355 | 2.5623 | 3.1049 | | 3.0771 | 2.4490 | 3.5258 | 3.5287 | H9 | 2.7061 | 2.0997 | 1.0934 | 2.2160 | 4.0150 | 3.5077 | 2.3766 | 3.0771 | | 3.1131 | 2.5758 | 2.5825 | H10 | 4.1568 | 2.6627 | 2.1568 | 1.0963 | 5.1344 | 4.6859 | 4.6366 | 2.4490 | 3.1131 | | 1.7784 | 1.7787 | H11 | 4.5355 | 3.2360 | 2.1534 | 1.0983 | 5.4225 | 5.3354 | 4.6560 | 3.5258 | 2.5758 | 1.7784 | | 1.7679 | H12 | 4.5272 | 3.2361 | 2.1559 | 1.0984 | 5.8788 | 5.0393 | 4.6347 | 3.5287 | 2.5825 | 1.7787 | 1.7679 | |
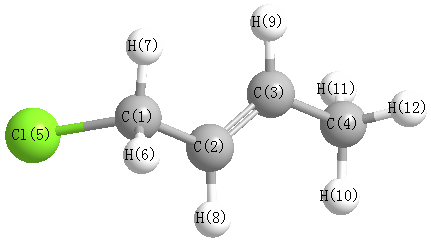 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.280 |
|
C1 |
C2 |
H8 |
116.249 |
| C2 |
C1 |
Cl5 |
111.367 |
|
C2 |
C1 |
H6 |
111.846 |
| C2 |
C1 |
H7 |
111.438 |
|
C2 |
C3 |
C4 |
124.697 |
| C2 |
C3 |
H9 |
119.001 |
|
C3 |
C2 |
H8 |
120.470 |
| C3 |
C4 |
H10 |
111.188 |
|
C3 |
C4 |
H11 |
110.791 |
| C3 |
C4 |
H12 |
110.986 |
|
C4 |
C3 |
H9 |
116.302 |
| Cl5 |
C1 |
H6 |
105.825 |
|
Cl5 |
C1 |
H7 |
107.017 |
| H6 |
C1 |
H7 |
109.076 |
|
H10 |
C4 |
H11 |
108.264 |
| H10 |
C4 |
H12 |
108.283 |
|
H11 |
C4 |
H12 |
107.184 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
