Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -616.485609 |
| Energy at 298.15K | -616.492710 |
| HF Energy | -616.485609 |
| Nuclear repulsion energy | 205.651577 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3201 |
3044 |
8.75 |
|
|
|
| 2 |
A' |
3162 |
3007 |
14.85 |
|
|
|
| 3 |
A' |
3151 |
2996 |
15.27 |
|
|
|
| 4 |
A' |
3094 |
2943 |
20.52 |
|
|
|
| 5 |
A' |
3058 |
2908 |
27.80 |
|
|
|
| 6 |
A' |
1793 |
1705 |
0.85 |
|
|
|
| 7 |
A' |
1519 |
1445 |
11.90 |
|
|
|
| 8 |
A' |
1500 |
1426 |
2.83 |
|
|
|
| 9 |
A' |
1441 |
1370 |
3.33 |
|
|
|
| 10 |
A' |
1353 |
1287 |
24.98 |
|
|
|
| 11 |
A' |
1342 |
1276 |
14.05 |
|
|
|
| 12 |
A' |
1317 |
1253 |
1.83 |
|
|
|
| 13 |
A' |
1145 |
1089 |
0.11 |
|
|
|
| 14 |
A' |
1068 |
1015 |
12.46 |
|
|
|
| 15 |
A' |
930 |
885 |
10.75 |
|
|
|
| 16 |
A' |
733 |
697 |
21.90 |
|
|
|
| 17 |
A' |
594 |
565 |
4.48 |
|
|
|
| 18 |
A' |
338 |
322 |
1.14 |
|
|
|
| 19 |
A' |
174 |
165 |
0.84 |
|
|
|
| 20 |
A" |
3142 |
2988 |
6.90 |
|
|
|
| 21 |
A" |
3118 |
2966 |
18.59 |
|
|
|
| 22 |
A" |
1507 |
1433 |
7.66 |
|
|
|
| 23 |
A" |
1220 |
1160 |
2.08 |
|
|
|
| 24 |
A" |
1080 |
1027 |
0.06 |
|
|
|
| 25 |
A" |
1003 |
954 |
35.46 |
|
|
|
| 26 |
A" |
964 |
916 |
0.00 |
|
|
|
| 27 |
A" |
734 |
698 |
0.42 |
|
|
|
| 28 |
A" |
244 |
232 |
4.36 |
|
|
|
| 29 |
A" |
209 |
199 |
2.41 |
|
|
|
| 30 |
A" |
119 |
113 |
1.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22125.6 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 21041.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.311 |
0.297 |
0.000 |
| C2 |
0.000 |
1.013 |
0.000 |
| C3 |
1.219 |
0.479 |
0.000 |
| C4 |
2.488 |
1.266 |
0.000 |
| Cl5 |
-1.192 |
-1.491 |
0.000 |
| H6 |
-1.901 |
0.565 |
0.883 |
| H7 |
-1.901 |
0.565 |
-0.883 |
| H8 |
-0.111 |
2.100 |
0.000 |
| H9 |
1.313 |
-0.606 |
0.000 |
| H10 |
2.299 |
2.344 |
0.000 |
| H11 |
3.100 |
1.028 |
0.880 |
| H12 |
3.100 |
1.028 |
-0.880 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4943 | 2.5365 | 3.9214 | 1.7923 | 1.0949 | 1.0949 | 2.1658 | 2.7757 | 4.1504 | 4.5568 | 4.5568 |
C2 | 1.4943 | | 1.3305 | 2.5012 | 2.7740 | 2.1432 | 2.1432 | 1.0921 | 2.0849 | 2.6565 | 3.2220 | 3.2220 | C3 | 2.5365 | 1.3305 | | 1.4940 | 3.1137 | 3.2432 | 3.2432 | 2.0962 | 1.0891 | 2.1556 | 2.1478 | 2.1478 | C4 | 3.9214 | 2.5012 | 1.4940 | | 4.5992 | 4.5318 | 4.5318 | 2.7297 | 2.2104 | 1.0947 | 1.0974 | 1.0974 | Cl5 | 1.7923 | 2.7740 | 3.1137 | 4.5992 | | 2.3477 | 2.3477 | 3.7504 | 2.6576 | 5.1868 | 5.0537 | 5.0537 | H6 | 1.0949 | 2.1432 | 3.2432 | 4.5318 | 2.3477 | | 1.7653 | 2.5174 | 3.5332 | 4.6458 | 5.0219 | 5.3221 | H7 | 1.0949 | 2.1432 | 3.2432 | 4.5318 | 2.3477 | 1.7653 | | 2.5174 | 3.5332 | 4.6458 | 5.3221 | 5.0219 | H8 | 2.1658 | 1.0921 | 2.0962 | 2.7297 | 3.7504 | 2.5174 | 2.5174 | | 3.0577 | 2.4222 | 3.4972 | 3.4972 | H9 | 2.7757 | 2.0849 | 1.0891 | 2.2104 | 2.6576 | 3.5332 | 3.5332 | 3.0577 | | 3.1106 | 2.5755 | 2.5755 | H10 | 4.1504 | 2.6565 | 2.1556 | 1.0947 | 5.1868 | 4.6458 | 4.6458 | 2.4222 | 3.1106 | | 1.7745 | 1.7745 | H11 | 4.5568 | 3.2220 | 2.1478 | 1.0974 | 5.0537 | 5.0219 | 5.3221 | 3.4972 | 2.5755 | 1.7745 | | 1.7592 | H12 | 4.5568 | 3.2220 | 2.1478 | 1.0974 | 5.0537 | 5.3221 | 5.0219 | 3.4972 | 2.5755 | 1.7745 | 1.7592 | |
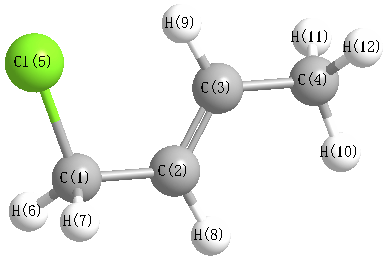 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.685 |
|
C1 |
C2 |
H8 |
112.810 |
| C2 |
C1 |
Cl5 |
114.836 |
|
C2 |
C1 |
H6 |
110.799 |
| C2 |
C1 |
H7 |
110.799 |
|
C2 |
C3 |
C4 |
124.530 |
| C2 |
C3 |
H9 |
118.665 |
|
C3 |
C2 |
H8 |
119.505 |
| C3 |
C4 |
H10 |
111.830 |
|
C3 |
C4 |
H11 |
111.027 |
| C3 |
C4 |
H12 |
111.027 |
|
C4 |
C3 |
H9 |
116.805 |
| Cl5 |
C1 |
H6 |
106.285 |
|
Cl5 |
C1 |
H7 |
106.285 |
| H6 |
C1 |
H7 |
107.442 |
|
H10 |
C4 |
H11 |
108.095 |
| H10 |
C4 |
H12 |
108.095 |
|
H11 |
C4 |
H12 |
106.554 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.473 |
|
|
|
| 2 |
C |
-0.140 |
|
|
|
| 3 |
C |
-0.103 |
|
|
|
| 4 |
C |
-0.565 |
|
|
|
| 5 |
Cl |
-0.067 |
|
|
|
| 6 |
H |
0.225 |
|
|
|
| 7 |
H |
0.225 |
|
|
|
| 8 |
H |
0.164 |
|
|
|
| 9 |
H |
0.184 |
|
|
|
| 10 |
H |
0.177 |
|
|
|
| 11 |
H |
0.186 |
|
|
|
| 12 |
H |
0.186 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.337 |
2.001 |
0.000 |
2.030 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.099 |
-2.603 |
0.000 |
| y |
-2.603 |
-38.157 |
0.000 |
| z |
0.000 |
0.000 |
-39.039 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.499 |
-2.603 |
0.000 |
| y |
-2.603 |
-1.088 |
0.000 |
| z |
0.000 |
0.000 |
-2.411 |
|
| Polar |
|---|
| 3z2-r2 | -4.822 |
|---|
| x2-y2 | 3.058 |
|---|
| xy | -2.603 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.763 |
0.408 |
0.000 |
| y |
0.408 |
8.304 |
0.000 |
| z |
0.000 |
0.000 |
4.861 |
<r2> (average value of r
2) Å
2
| <r2> |
209.022 |
| (<r2>)1/2 |
14.458 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -616.487572 |
| Energy at 298.15K | -616.494623 |
| HF Energy | -616.487572 |
| Nuclear repulsion energy | 201.232358 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3197 |
3040 |
18.46 |
|
|
|
| 2 |
A |
3179 |
3023 |
1.77 |
|
|
|
| 3 |
A |
3171 |
3016 |
27.98 |
|
|
|
| 4 |
A |
3154 |
3000 |
2.45 |
|
|
|
| 5 |
A |
3120 |
2967 |
15.48 |
|
|
|
| 6 |
A |
3118 |
2965 |
17.80 |
|
|
|
| 7 |
A |
3060 |
2910 |
23.59 |
|
|
|
| 8 |
A |
1775 |
1688 |
13.59 |
|
|
|
| 9 |
A |
1517 |
1443 |
14.34 |
|
|
|
| 10 |
A |
1506 |
1433 |
2.84 |
|
|
|
| 11 |
A |
1505 |
1431 |
9.17 |
|
|
|
| 12 |
A |
1439 |
1368 |
2.49 |
|
|
|
| 13 |
A |
1359 |
1292 |
7.89 |
|
|
|
| 14 |
A |
1345 |
1279 |
1.63 |
|
|
|
| 15 |
A |
1300 |
1237 |
42.03 |
|
|
|
| 16 |
A |
1223 |
1163 |
3.23 |
|
|
|
| 17 |
A |
1142 |
1086 |
0.70 |
|
|
|
| 18 |
A |
1122 |
1067 |
2.63 |
|
|
|
| 19 |
A |
1077 |
1025 |
0.34 |
|
|
|
| 20 |
A |
1012 |
963 |
32.78 |
|
|
|
| 21 |
A |
964 |
917 |
12.39 |
|
|
|
| 22 |
A |
904 |
860 |
3.02 |
|
|
|
| 23 |
A |
808 |
768 |
15.05 |
|
|
|
| 24 |
A |
702 |
668 |
62.97 |
|
|
|
| 25 |
A |
495 |
471 |
1.76 |
|
|
|
| 26 |
A |
357 |
340 |
4.50 |
|
|
|
| 27 |
A |
279 |
265 |
2.20 |
|
|
|
| 28 |
A |
216 |
205 |
1.15 |
|
|
|
| 29 |
A |
160 |
152 |
2.60 |
|
|
|
| 30 |
A |
90 |
85 |
0.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22146.9 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 21061.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.773 |
0.822 |
0.112 |
| C2 |
0.519 |
0.172 |
0.452 |
| C3 |
1.600 |
0.234 |
-0.327 |
| C4 |
2.922 |
-0.381 |
-0.010 |
| Cl5 |
-2.100 |
-0.396 |
-0.103 |
| H6 |
-1.122 |
1.487 |
0.906 |
| H7 |
-0.710 |
1.384 |
-0.822 |
| H8 |
0.555 |
-0.370 |
1.397 |
| H9 |
1.530 |
0.772 |
-1.275 |
| H10 |
2.904 |
-0.903 |
0.951 |
| H11 |
3.216 |
-1.100 |
-0.785 |
| H12 |
3.713 |
0.379 |
0.029 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4851 | 2.4832 | 3.8877 | 1.8138 | 1.0929 | 1.0919 | 2.1983 | 2.6887 | 4.1470 | 4.5173 | 4.5084 |
C2 | 1.4851 | | 1.3340 | 2.5093 | 2.7363 | 2.1510 | 2.1451 | 1.0896 | 2.0892 | 2.6636 | 3.2285 | 3.2290 | C3 | 2.4832 | 1.3340 | | 1.4927 | 3.7592 | 3.2403 | 2.6271 | 2.1044 | 1.0916 | 2.1515 | 2.1448 | 2.1483 | C4 | 3.8877 | 2.5093 | 1.4927 | | 5.0230 | 4.5481 | 4.1191 | 2.7540 | 2.2058 | 1.0942 | 1.0974 | 1.0974 | Cl5 | 1.8138 | 2.7363 | 3.7592 | 5.0230 | | 2.3487 | 2.3698 | 3.0491 | 3.9892 | 5.1386 | 5.4053 | 5.8658 | H6 | 1.0929 | 2.1510 | 3.2403 | 4.5481 | 2.3487 | | 1.7792 | 2.5494 | 3.5073 | 4.6824 | 5.3262 | 5.0376 | H7 | 1.0919 | 2.1451 | 2.6271 | 4.1191 | 2.3698 | 1.7792 | | 3.0980 | 2.3661 | 4.6299 | 4.6457 | 4.6148 | H8 | 2.1983 | 1.0896 | 2.1044 | 2.7540 | 3.0491 | 2.5494 | 3.0980 | | 3.0645 | 2.4499 | 3.5180 | 3.5225 | H9 | 2.6887 | 2.0892 | 1.0916 | 2.2058 | 3.9892 | 3.5073 | 2.3661 | 3.0645 | | 3.1060 | 2.5659 | 2.5724 | H10 | 4.1470 | 2.6636 | 2.1515 | 1.0942 | 5.1386 | 4.6824 | 4.6299 | 2.4499 | 3.1060 | | 1.7749 | 1.7752 | H11 | 4.5173 | 3.2285 | 2.1448 | 1.0974 | 5.4053 | 5.3262 | 4.6457 | 3.5180 | 2.5659 | 1.7749 | | 1.7597 | H12 | 4.5084 | 3.2290 | 2.1483 | 1.0974 | 5.8658 | 5.0376 | 4.6148 | 3.5225 | 2.5724 | 1.7752 | 1.7597 | |
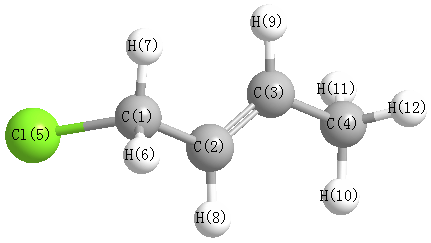 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.400 |
|
C1 |
C2 |
H8 |
116.412 |
| C2 |
C1 |
Cl5 |
111.698 |
|
C2 |
C1 |
H6 |
112.209 |
| C2 |
C1 |
H7 |
111.789 |
|
C2 |
C3 |
C4 |
125.079 |
| C2 |
C3 |
H9 |
118.582 |
|
C3 |
C2 |
H8 |
120.186 |
| C3 |
C4 |
H10 |
111.624 |
|
C3 |
C4 |
H11 |
110.883 |
| C3 |
C4 |
H12 |
111.165 |
|
C4 |
C3 |
H9 |
116.339 |
| Cl5 |
C1 |
H6 |
105.095 |
|
Cl5 |
C1 |
H7 |
106.633 |
| H6 |
C1 |
H7 |
109.052 |
|
H10 |
C4 |
H11 |
108.174 |
| H10 |
C4 |
H12 |
108.198 |
|
H11 |
C4 |
H12 |
106.601 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.480 |
|
|
|
| 2 |
C |
-0.104 |
|
|
|
| 3 |
C |
-0.111 |
|
|
|
| 4 |
C |
-0.564 |
|
|
|
| 5 |
Cl |
-0.084 |
|
|
|
| 6 |
H |
0.226 |
|
|
|
| 7 |
H |
0.220 |
|
|
|
| 8 |
H |
0.173 |
|
|
|
| 9 |
H |
0.162 |
|
|
|
| 10 |
H |
0.185 |
|
|
|
| 11 |
H |
0.191 |
|
|
|
| 12 |
H |
0.187 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.309 |
1.255 |
0.155 |
2.633 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.353 |
-2.029 |
-1.037 |
| y |
-2.029 |
-37.440 |
-0.852 |
| z |
-1.037 |
-0.852 |
-36.808 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.229 |
-2.029 |
-1.037 |
| y |
-2.029 |
0.640 |
-0.852 |
| z |
-1.037 |
-0.852 |
1.589 |
|
| Polar |
|---|
| 3z2-r2 | 3.177 |
|---|
| x2-y2 | -1.913 |
|---|
| xy | -2.029 |
|---|
| xz | -1.037 |
|---|
| yz | -0.852 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.455 |
0.228 |
-0.841 |
| y |
0.228 |
6.034 |
-0.366 |
| z |
-0.841 |
-0.366 |
6.330 |
<r2> (average value of r
2) Å
2
| <r2> |
231.206 |
| (<r2>)1/2 |
15.205 |
