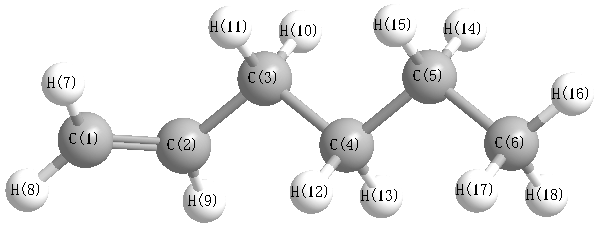
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3246 |
3106 |
21.00 |
|
|
|
| 2 |
A |
3164 |
3027 |
6.43 |
|
|
|
| 3 |
A |
3146 |
3010 |
30.55 |
|
|
|
| 4 |
A |
3130 |
2994 |
39.13 |
|
|
|
| 5 |
A |
3126 |
2990 |
56.98 |
|
|
|
| 6 |
A |
3095 |
2961 |
51.32 |
|
|
|
| 7 |
A |
3076 |
2943 |
3.23 |
|
|
|
| 8 |
A |
3063 |
2931 |
7.39 |
|
|
|
| 9 |
A |
3051 |
2919 |
38.76 |
|
|
|
| 10 |
A |
3041 |
2909 |
61.21 |
|
|
|
| 11 |
A |
3033 |
2902 |
0.71 |
|
|
|
| 12 |
A |
3021 |
2891 |
16.52 |
|
|
|
| 13 |
A |
1739 |
1664 |
11.11 |
|
|
|
| 14 |
A |
1534 |
1468 |
5.55 |
|
|
|
| 15 |
A |
1525 |
1459 |
7.11 |
|
|
|
| 16 |
A |
1522 |
1456 |
1.45 |
|
|
|
| 17 |
A |
1511 |
1445 |
0.43 |
|
|
|
| 18 |
A |
1503 |
1438 |
2.11 |
|
|
|
| 19 |
A |
1469 |
1405 |
0.90 |
|
|
|
| 20 |
A |
1434 |
1372 |
2.54 |
|
|
|
| 21 |
A |
1414 |
1353 |
0.59 |
|
|
|
| 22 |
A |
1367 |
1308 |
0.75 |
|
|
|
| 23 |
A |
1344 |
1286 |
0.65 |
|
|
|
| 24 |
A |
1332 |
1274 |
0.16 |
|
|
|
| 25 |
A |
1323 |
1266 |
0.50 |
|
|
|
| 26 |
A |
1279 |
1223 |
3.50 |
|
|
|
| 27 |
A |
1257 |
1203 |
0.13 |
|
|
|
| 28 |
A |
1207 |
1154 |
0.36 |
|
|
|
| 29 |
A |
1135 |
1086 |
2.82 |
|
|
|
| 30 |
A |
1081 |
1034 |
0.16 |
|
|
|
| 31 |
A |
1060 |
1014 |
7.36 |
|
|
|
| 32 |
A |
1040 |
995 |
1.69 |
|
|
|
| 33 |
A |
1035 |
990 |
9.11 |
|
|
|
| 34 |
A |
951 |
910 |
2.25 |
|
|
|
| 35 |
A |
935 |
895 |
38.49 |
|
|
|
| 36 |
A |
930 |
890 |
4.09 |
|
|
|
| 37 |
A |
913 |
873 |
1.91 |
|
|
|
| 38 |
A |
800 |
766 |
1.04 |
|
|
|
| 39 |
A |
739 |
707 |
3.98 |
|
|
|
| 40 |
A |
650 |
622 |
9.89 |
|
|
|
| 41 |
A |
456 |
436 |
1.15 |
|
|
|
| 42 |
A |
357 |
342 |
0.07 |
|
|
|
| 43 |
A |
352 |
337 |
0.45 |
|
|
|
| 44 |
A |
247 |
236 |
0.00 |
|
|
|
| 45 |
A |
174 |
167 |
0.02 |
|
|
|
| 46 |
A |
124 |
119 |
0.07 |
|
|
|
| 47 |
A |
89 |
86 |
0.02 |
|
|
|
| 48 |
A |
73 |
70 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36545.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 34962.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.391 |
|
|
|
| 2 |
C |
-0.069 |
|
|
|
| 3 |
C |
-0.358 |
|
|
|
| 4 |
C |
-0.303 |
|
|
|
| 5 |
C |
-0.305 |
|
|
|
| 6 |
C |
-0.508 |
|
|
|
| 7 |
H |
0.158 |
|
|
|
| 8 |
H |
0.161 |
|
|
|
| 9 |
H |
0.152 |
|
|
|
| 10 |
H |
0.167 |
|
|
|
| 11 |
H |
0.163 |
|
|
|
| 12 |
H |
0.165 |
|
|
|
| 13 |
H |
0.155 |
|
|
|
| 14 |
H |
0.157 |
|
|
|
| 15 |
H |
0.158 |
|
|
|
| 16 |
H |
0.167 |
|
|
|
| 17 |
H |
0.166 |
|
|
|
| 18 |
H |
0.165 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.376 |
0.056 |
0.109 |
0.396 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.300 |
-0.863 |
-0.454 |
| y |
-0.863 |
-39.912 |
-1.128 |
| z |
-0.454 |
-1.128 |
-38.629 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.029 |
-0.863 |
-0.454 |
| y |
-0.863 |
-0.448 |
-1.128 |
| z |
-0.454 |
-1.128 |
1.477 |
|
| Polar |
|---|
| 3z2-r2 | 2.955 |
|---|
| x2-y2 | -0.387 |
|---|
| xy | -0.863 |
|---|
| xz | -0.454 |
|---|
| yz | -1.128 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.371 |
-0.490 |
-1.132 |
| y |
-0.490 |
7.851 |
-0.554 |
| z |
-1.132 |
-0.554 |
8.466 |
<r2> (average value of r
2) Å
2
| <r2> |
285.531 |
| (<r2>)1/2 |
16.898 |