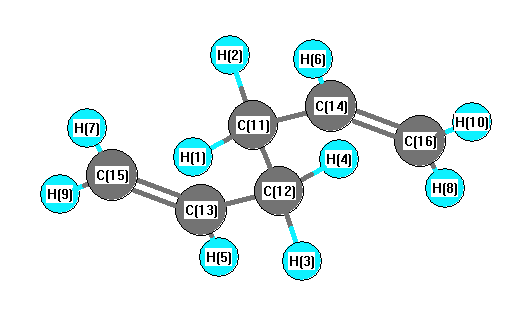
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3248 |
3107 |
9.13 |
|
|
|
| 2 |
A |
3166 |
3029 |
1.83 |
|
|
|
| 3 |
A |
3147 |
3011 |
43.38 |
|
|
|
| 4 |
A |
3104 |
2970 |
22.83 |
|
|
|
| 5 |
A |
3028 |
2897 |
10.26 |
|
|
|
| 6 |
A |
1738 |
1663 |
4.98 |
|
|
|
| 7 |
A |
1503 |
1438 |
2.45 |
|
|
|
| 8 |
A |
1467 |
1403 |
2.17 |
|
|
|
| 9 |
A |
1380 |
1320 |
0.52 |
|
|
|
| 10 |
A |
1332 |
1275 |
1.31 |
|
|
|
| 11 |
A |
1253 |
1199 |
0.19 |
|
|
|
| 12 |
A |
1100 |
1052 |
0.02 |
|
|
|
| 13 |
A |
1047 |
1001 |
0.85 |
|
|
|
| 14 |
A |
1035 |
990 |
9.20 |
|
|
|
| 15 |
A |
956 |
915 |
0.40 |
|
|
|
| 16 |
A |
938 |
897 |
18.67 |
|
|
|
| 17 |
A |
789 |
754 |
0.51 |
|
|
|
| 18 |
A |
636 |
608 |
7.20 |
|
|
|
| 19 |
A |
414 |
396 |
0.00 |
|
|
|
| 20 |
A |
348 |
332 |
0.12 |
|
|
|
| 21 |
A |
99 |
95 |
0.02 |
|
|
|
| 22 |
A |
74 |
71 |
0.02 |
|
|
|
| 23 |
B |
3248 |
3107 |
30.84 |
|
|
|
| 24 |
B |
3165 |
3028 |
10.65 |
|
|
|
| 25 |
B |
3147 |
3010 |
15.88 |
|
|
|
| 26 |
B |
3088 |
2954 |
7.62 |
|
|
|
| 27 |
B |
3031 |
2900 |
41.04 |
|
|
|
| 28 |
B |
1741 |
1665 |
15.40 |
|
|
|
| 29 |
B |
1516 |
1450 |
7.20 |
|
|
|
| 30 |
B |
1469 |
1405 |
0.08 |
|
|
|
| 31 |
B |
1339 |
1281 |
0.74 |
|
|
|
| 32 |
B |
1322 |
1265 |
0.12 |
|
|
|
| 33 |
B |
1281 |
1225 |
3.35 |
|
|
|
| 34 |
B |
1189 |
1137 |
2.53 |
|
|
|
| 35 |
B |
1038 |
993 |
20.07 |
|
|
|
| 36 |
B |
969 |
927 |
5.22 |
|
|
|
| 37 |
B |
942 |
901 |
1.78 |
|
|
|
| 38 |
B |
938 |
898 |
64.67 |
|
|
|
| 39 |
B |
663 |
634 |
18.51 |
|
|
|
| 40 |
B |
435 |
416 |
1.83 |
|
|
|
| 41 |
B |
217 |
207 |
0.19 |
|
|
|
| 42 |
B |
115 |
110 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31326.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 29970.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.361 |
|
|
|
| 2 |
C |
-0.361 |
|
|
|
| 3 |
C |
-0.071 |
|
|
|
| 4 |
C |
-0.071 |
|
|
|
| 5 |
C |
-0.389 |
|
|
|
| 6 |
C |
-0.389 |
|
|
|
| 7 |
H |
0.170 |
|
|
|
| 8 |
H |
0.175 |
|
|
|
| 9 |
H |
0.170 |
|
|
|
| 10 |
H |
0.175 |
|
|
|
| 11 |
H |
0.153 |
|
|
|
| 12 |
H |
0.153 |
|
|
|
| 13 |
H |
0.163 |
|
|
|
| 14 |
H |
0.160 |
|
|
|
| 15 |
H |
0.163 |
|
|
|
| 16 |
H |
0.160 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.255 |
0.255 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.495 |
1.582 |
0.000 |
| y |
1.582 |
-38.665 |
0.000 |
| z |
0.000 |
0.000 |
-36.406 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.960 |
1.582 |
0.000 |
| y |
1.582 |
-1.214 |
0.000 |
| z |
0.000 |
0.000 |
2.174 |
|
| Polar |
|---|
| 3z2-r2 | 4.347 |
|---|
| x2-y2 | 0.169 |
|---|
| xy | 1.582 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.807 |
1.699 |
0.000 |
| y |
1.699 |
12.467 |
0.000 |
| z |
0.000 |
0.000 |
8.804 |
<r2> (average value of r
2) Å
2
| <r2> |
254.188 |
| (<r2>)1/2 |
15.943 |