return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CCl2CH3 (Propane, 2,2-dichloro-)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-1036.780661
Energy at 298.15K 
HF Energy-1036.063005
Nuclear repulsion energy295.054896
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3185 3033 9.05      
2 A1 3087 2940 11.37      
3 A1 1538 1465 2.67      
4 A1 1478 1407 6.53      
5 A1 1231 1172 30.08      
6 A1 966 920 6.91      
7 A1 586 558 14.36      
8 A1 378 360 0.49      
9 A1 268 255 1.20      
10 A2 3162 3011 0.00      
11 A2 1528 1455 0.00      
12 A2 1060 1009 0.00      
13 A2 295 281 0.00      
14 A2 284 271 0.00      
15 B1 3168 3017 21.23      
16 B1 1548 1474 7.99      
17 B1 1189 1132 80.41      
18 B1 715 681 81.55      
19 B1 373 356 1.79      
20 B1 320 304 0.02      
21 B2 3183 3032 5.86      
22 B2 3083 2936 5.25      
23 B2 1525 1452 4.51      
24 B2 1461 1391 14.55      
25 B2 1262 1201 4.86      
26 B2 987 940 0.17      
27 B2 401 382 3.33      

Unscaled Zero Point Vibrational Energy (zpe) 19130.3 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 18217.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.12224 0.08122 0.07140

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.334
Cl2 1.463 0.000 -0.717
Cl3 -1.463 0.000 -0.717
C4 0.000 1.269 1.173
C5 0.000 -1.269 1.173
H6 0.000 2.156 0.533
H7 0.000 -2.156 0.533
H8 -0.893 1.286 1.808
H9 0.893 1.286 1.808
H10 0.893 -1.286 1.808
H11 -0.893 -1.286 1.808

Atom - Atom Distances (Å)
  C1 Cl2 Cl3 C4 C5 H6 H7 H8 H9 H10 H11
C11.80141.80141.52101.52102.16522.16522.15032.15032.15032.1503
Cl21.80142.92572.70582.70582.88992.88993.68502.89042.89043.6850
Cl31.80142.92572.70582.70582.88992.88992.89043.68503.68502.8904
C41.52102.70582.70582.53781.09363.48411.09581.09582.78042.7804
C51.52102.70582.70582.53783.48411.09362.78042.78041.09581.0958
H62.16522.88992.88991.09363.48414.31211.78261.78263.77793.7779
H72.16522.88992.88993.48411.09364.31213.77793.77791.78261.7826
H82.15033.68502.89041.09582.78041.78263.77791.78583.13202.5730
H92.15032.89043.68501.09582.78041.78263.77791.78582.57303.1320
H102.15032.89043.68502.78041.09583.77791.78263.13202.57301.7858
H112.15033.68502.89042.78041.09583.77791.78262.57303.13201.7858

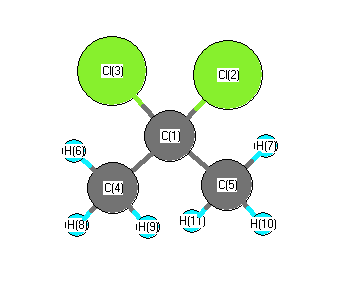
picture of Propane, 2,2-dichloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H6 110.755 C1 C4 H8 109.444
C1 C4 H9 109.444 C1 C5 H7 110.755
C1 C5 H10 109.444 C1 C5 H11 109.444
Cl2 C1 Cl3 108.598 Cl2 C1 C4 108.768
Cl2 C1 C5 108.768 Cl3 C1 C4 108.768
Cl3 C1 C5 108.768 C4 C1 C5 113.081
H6 C4 H8 109.015 H6 C4 H9 109.015
H7 C5 H10 109.015 H7 C5 H11 109.015
H8 C4 H9 109.144 H10 C5 H11 109.144
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability