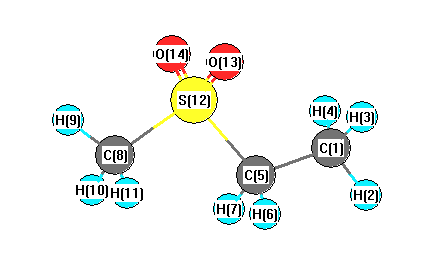
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3204 |
3065 |
1.70 |
|
|
|
| 2 |
A |
3198 |
3060 |
0.87 |
|
|
|
| 3 |
A |
3172 |
3034 |
7.59 |
|
|
|
| 4 |
A |
3150 |
3014 |
8.20 |
|
|
|
| 5 |
A |
3140 |
3004 |
8.96 |
|
|
|
| 6 |
A |
3092 |
2958 |
1.22 |
|
|
|
| 7 |
A |
3090 |
2956 |
4.91 |
|
|
|
| 8 |
A |
3073 |
2940 |
16.39 |
|
|
|
| 9 |
A |
1527 |
1461 |
8.94 |
|
|
|
| 10 |
A |
1522 |
1456 |
10.18 |
|
|
|
| 11 |
A |
1484 |
1420 |
11.90 |
|
|
|
| 12 |
A |
1480 |
1416 |
4.08 |
|
|
|
| 13 |
A |
1474 |
1410 |
4.25 |
|
|
|
| 14 |
A |
1435 |
1373 |
2.80 |
|
|
|
| 15 |
A |
1371 |
1312 |
22.40 |
|
|
|
| 16 |
A |
1350 |
1291 |
146.37 |
|
|
|
| 17 |
A |
1315 |
1258 |
56.13 |
|
|
|
| 18 |
A |
1276 |
1221 |
9.30 |
|
|
|
| 19 |
A |
1145 |
1095 |
146.18 |
|
|
|
| 20 |
A |
1092 |
1045 |
2.10 |
|
|
|
| 21 |
A |
1071 |
1025 |
10.52 |
|
|
|
| 22 |
A |
1004 |
961 |
3.98 |
|
|
|
| 23 |
A |
981 |
939 |
24.91 |
|
|
|
| 24 |
A |
974 |
932 |
14.33 |
|
|
|
| 25 |
A |
805 |
770 |
54.92 |
|
|
|
| 26 |
A |
732 |
700 |
28.64 |
|
|
|
| 27 |
A |
650 |
621 |
13.56 |
|
|
|
| 28 |
A |
493 |
472 |
19.55 |
|
|
|
| 29 |
A |
437 |
419 |
28.24 |
|
|
|
| 30 |
A |
396 |
379 |
7.68 |
|
|
|
| 31 |
A |
309 |
295 |
1.05 |
|
|
|
| 32 |
A |
280 |
268 |
1.43 |
|
|
|
| 33 |
A |
225 |
216 |
1.94 |
|
|
|
| 34 |
A |
205 |
196 |
1.09 |
|
|
|
| 35 |
A |
196 |
188 |
0.09 |
|
|
|
| 36 |
A |
86 |
82 |
2.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25216.7 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 24124.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.533 |
|
|
-0.191 |
| 2 |
H |
0.220 |
|
|
0.081 |
| 3 |
H |
0.193 |
|
|
0.083 |
| 4 |
H |
0.179 |
|
|
0.067 |
| 5 |
C |
-0.527 |
|
|
-0.142 |
| 6 |
H |
0.240 |
|
|
0.115 |
| 7 |
H |
0.215 |
|
|
0.091 |
| 8 |
C |
-0.711 |
|
|
-0.572 |
| 9 |
H |
0.220 |
|
|
0.195 |
| 10 |
H |
0.247 |
|
|
0.222 |
| 11 |
H |
0.222 |
|
|
0.185 |
| 12 |
S |
1.112 |
|
|
0.828 |
| 13 |
O |
-0.541 |
|
|
-0.466 |
| 14 |
O |
-0.537 |
|
|
-0.496 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.069 |
2.825 |
-2.162 |
4.698 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
3.100 |
2.899 |
-2.151 |
4.758 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.604 |
-5.167 |
-1.270 |
| y |
-5.167 |
-41.418 |
-0.594 |
| z |
-1.270 |
-0.594 |
-47.615 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.912 |
-5.167 |
-1.270 |
| y |
-5.167 |
4.192 |
-0.594 |
| z |
-1.270 |
-0.594 |
-5.104 |
|
| Polar |
|---|
| 3z2-r2 | -10.208 |
|---|
| x2-y2 | -2.187 |
|---|
| xy | -5.167 |
|---|
| xz | -1.270 |
|---|
| yz | -0.594 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
192.522 |
| (<r2>)1/2 |
13.875 |