return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H8 ((Z)-hexa-2,3,4-triene)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-233.075870
Energy at 298.15K-233.082205
HF Energy-232.820343
Nuclear repulsion energy190.051318
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3179 3017 33.88      
2 A1 3163 3002 3.69      
3 A1 3067 2912 12.86      
4 A1 2203 2091 0.80      
5 A1 1543 1465 5.27      
6 A1 1452 1378 3.09      
7 A1 1419 1347 0.00      
8 A1 1143 1085 1.70      
9 A1 1098 1042 0.80      
10 A1 775 736 0.30      
11 A1 476 452 5.72      
12 A1 99 94 1.10      
13 A2 3122 2963 0.00      
14 A2 1529 1452 0.00      
15 A2 1075 1021 0.00      
16 A2 861 817 0.00      
17 A2 557 528 0.00      
18 A2 231 220 0.00      
19 A2 144 137 0.00      
20 B1 3122 2963 43.54      
21 B1 1530 1452 10.91      
22 B1 1073 1018 0.54      
23 B1 712 676 39.25      
24 B1 241 229 1.37      
25 B1 129 123 1.85      
26 B2 3179 3017 15.76      
27 B2 3163 3002 0.26      
28 B2 3067 2911 66.07      
29 B2 1729 1641 3.65      
30 B2 1533 1455 9.57      
31 B2 1448 1375 14.34      
32 B2 1316 1249 20.96      
33 B2 1124 1067 2.95      
34 B2 956 907 30.83      
35 B2 571 542 2.80      
36 B2 224 213 0.50      

Unscaled Zero Point Vibrational Energy (zpe) 26125.6 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 24798.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.41917 0.04614 0.04221

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 0.000 2.029 1.603
H2 0.862 3.350 0.798
H3 -0.862 3.350 0.798
H4 0.000 -2.029 1.603
H5 0.862 -3.350 0.798
H6 -0.862 -3.350 0.798
C7 0.000 2.701 0.744
C8 0.000 -2.701 0.744
H9 0.000 2.450 -1.419
H10 0.000 -2.450 -1.419
C11 0.000 -0.617 -0.528
C12 0.000 0.617 -0.528
C13 0.000 1.927 -0.513
C14 0.000 -1.927 -0.513

Atom - Atom Distances (Å)
  H1 H2 H3 H4 H5 H6 C7 C8 H9 H10 C11 C12 C13 C14
H11.77011.77014.05825.50645.50641.08974.80683.05065.40313.39702.55662.11804.4858
H21.77011.72315.50646.69946.91741.08016.11152.54306.26884.27003.15792.11825.5046
H31.77011.72315.50646.91746.69941.08016.11152.54306.26884.27003.15792.11825.5046
H44.05825.50645.50641.77011.77014.80681.08975.40313.05062.55663.39704.48582.1180
H55.50646.69946.91741.77011.72316.11151.08016.26882.54303.15794.27005.50462.1182
H65.50646.91746.69941.77011.72316.11151.08016.26882.54303.15794.27005.50462.1182
C71.08971.08011.08014.80686.11156.11155.40112.17765.58663.55282.44191.47644.7949
C84.80686.11156.11151.08971.08011.08015.40115.58662.17762.44193.55284.79491.4764
H93.05062.54302.54305.40316.26886.26882.17765.58664.90063.19352.03861.04644.4696
H105.40316.26886.26883.05062.54302.54305.58662.17764.90062.03863.19354.46961.0464
C113.39704.27004.27002.55663.15793.15793.55282.44193.19352.03861.23302.54311.3101
C122.55663.15793.15793.39704.27004.27002.44193.55282.03863.19351.23301.31012.5431
C132.11802.11822.11824.48585.50465.50461.47644.79491.04644.46962.54311.31013.8531
C144.48585.50465.50462.11802.11822.11824.79491.47644.46961.04641.31012.54313.8531

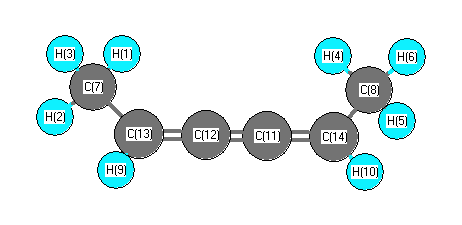
picture of (Z)-hexa-2,3,4-triene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C7 H2 109.331 H1 C7 H3 109.331
H1 C7 C13 110.344 H2 C7 H3 105.818
H2 C7 C13 110.952 H3 C7 C13 110.952
H4 C8 H5 109.331 H4 C8 H6 109.331
H4 C8 C14 110.344 H5 C8 H6 105.818
H5 C8 C14 110.952 H6 C8 C14 110.952
C7 C13 H9 118.350 C7 C13 C12 122.287
C8 C14 H10 118.350 C8 C14 C11 122.287
H9 C13 C12 119.362 H10 C14 C11 119.362
C11 C12 C13 179.329 C12 C11 C14 179.329
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 H 0.174      
2 H 0.167      
3 H 0.167      
4 H 0.174      
5 H 0.167      
6 H 0.167      
7 C -0.486      
8 C -0.486      
9 H 0.176      
10 H 0.176      
11 C 0.137      
12 C 0.137      
13 C -0.336      
14 C -0.336      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 5.119 0.000 0.000
y 0.000 20.587 0.000
z 0.000 0.000 6.900


<r2> (average value of r2) Å2
<r2> 265.181
(<r2>)1/2 16.284