return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,2-Butadiene, 3-methyl-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-195.040446
Energy at 298.15K-195.047559
HF Energy-194.827761
Nuclear repulsion energy156.991060
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3176 3014 19.59      
2 A1 3168 3007 9.06      
3 A1 3067 2911 56.31      
4 A1 2077 1972 16.38      
5 A1 1560 1480 0.08      
6 A1 1516 1439 2.04      
7 A1 1459 1385 2.35      
8 A1 1345 1276 17.18      
9 A1 1048 995 7.09      
10 A1 746 708 0.01      
11 A1 343 325 0.13      
12 A2 3120 2962 0.00      
13 A2 1525 1447 0.00      
14 A2 1029 976 0.00      
15 A2 624 592 0.00      
16 A2 168 160 0.00      
17 B1 3239 3075 7.60      
18 B1 3122 2963 49.91      
19 B1 1540 1461 13.95      
20 B1 1126 1069 0.15      
21 B1 1050 997 0.24      
22 B1 457 433 0.38      
23 B1 267 254 3.99      
24 B1 179 170 1.05      
25 B2 3175 3014 17.95      
26 B2 3063 2908 17.69      
27 B2 1535 1457 5.79      
28 B2 1443 1370 2.29      
29 B2 1242 1179 4.24      
30 B2 991 941 0.26      
31 B2 879 834 50.12      
32 B2 609 578 7.60      
33 B2 184 175 0.13      

Unscaled Zero Point Vibrational Energy (zpe) 25034.3 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 23762.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.27548 0.12007 0.08785

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 2.244
C2 0.000 0.000 0.934
H3 -0.925 0.000 2.816
H4 0.925 0.000 2.816
C5 0.000 0.000 -0.379
C6 0.000 1.286 -1.174
C7 0.000 -1.286 -1.174
H8 0.000 2.161 -0.522
H9 0.000 -2.161 -0.522
H10 0.882 1.337 -1.823
H11 -0.882 1.337 -1.823
H12 -0.882 -1.337 -1.823
H13 0.882 -1.337 -1.823

Atom - Atom Distances (Å)
  C1 C2 H3 H4 C5 C6 C7 H8 H9 H10 H11 H12 H13
C11.31051.08701.08702.62363.65243.65243.51043.51044.37174.37174.37174.3717
C21.31052.09702.09701.31302.46942.46942.60582.60583.18863.18863.18863.1886
H31.08702.09701.84913.32624.29294.29294.08254.08255.15494.82844.82845.1549
H41.08702.09701.84913.32624.29294.29294.08254.08254.82845.15495.15494.8284
C52.62361.31303.32623.32621.51211.51212.16602.16602.15652.15652.15652.1565
C63.65242.46944.29294.29291.51212.57281.09133.50891.09621.09622.84282.8428
C73.65242.46944.29294.29291.51212.57283.50891.09132.84282.84281.09621.0962
H83.51042.60584.08254.08252.16601.09133.50894.32271.77511.77513.83533.8353
H93.51042.60584.08254.08252.16603.50891.09134.32273.83533.83531.77511.7751
H104.37173.18865.15494.82842.15651.09622.84281.77513.83531.76333.20312.6740
H114.37173.18864.82845.15492.15651.09622.84281.77513.83531.76332.67403.2031
H124.37173.18864.82845.15492.15652.84281.09623.83531.77513.20312.67401.7633
H134.37173.18865.15494.82842.15652.84281.09623.83531.77512.67403.20311.7633

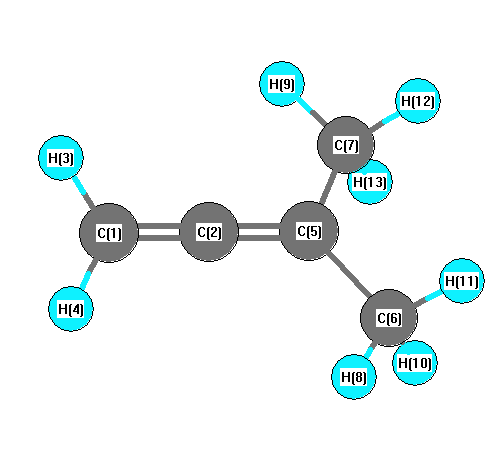
picture of 1,2-Butadiene, 3-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C5 180.000 C2 C1 H3 121.726
C2 C1 H4 121.726 C2 C5 C6 121.710
C2 C5 C7 121.710 H3 C1 H4 116.547
C5 C6 H8 111.590 C5 C6 H10 110.529
C5 C6 H11 110.529 C5 C7 H9 111.590
C5 C7 H12 110.529 C5 C7 H13 110.529
C6 C5 C7 116.580 H8 C6 H10 108.482
H8 C6 H11 108.482 H9 C7 H12 108.482
H9 C7 H13 108.482 H10 C6 H11 107.087
H12 C7 H13 107.087
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.534      
2 C 0.214      
3 H 0.173      
4 H 0.173      
5 C -0.032      
6 C -0.492      
7 C -0.492      
8 H 0.174      
9 H 0.174      
10 H 0.161      
11 H 0.161      
12 H 0.161      
13 H 0.161      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 5.452 0.000 0.000
y 0.000 5.963 0.000
z 0.000 0.000 12.493


<r2> (average value of r2) Å2
<r2> 143.141
(<r2>)1/2 11.964