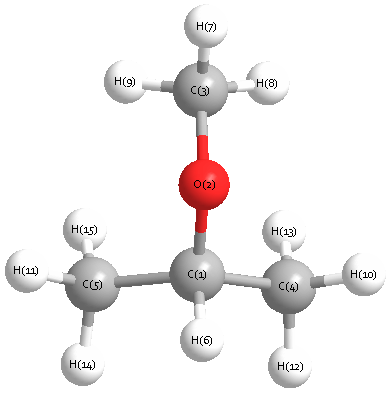
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3147 |
3011 |
37.37 |
|
|
|
| 2 |
A |
3146 |
3009 |
15.91 |
|
|
|
| 3 |
A |
3142 |
3006 |
18.16 |
|
|
|
| 4 |
A |
3137 |
3001 |
78.07 |
|
|
|
| 5 |
A |
3130 |
2995 |
0.06 |
|
|
|
| 6 |
A |
3069 |
2936 |
82.91 |
|
|
|
| 7 |
A |
3068 |
2935 |
21.50 |
|
|
|
| 8 |
A |
3062 |
2930 |
9.70 |
|
|
|
| 9 |
A |
3056 |
2924 |
1.32 |
|
|
|
| 10 |
A |
3013 |
2883 |
52.12 |
|
|
|
| 11 |
A |
1540 |
1473 |
13.97 |
|
|
|
| 12 |
A |
1535 |
1468 |
2.31 |
|
|
|
| 13 |
A |
1527 |
1461 |
1.95 |
|
|
|
| 14 |
A |
1522 |
1456 |
6.22 |
|
|
|
| 15 |
A |
1509 |
1443 |
0.01 |
|
|
|
| 16 |
A |
1506 |
1441 |
1.96 |
|
|
|
| 17 |
A |
1492 |
1428 |
0.49 |
|
|
|
| 18 |
A |
1435 |
1373 |
20.69 |
|
|
|
| 19 |
A |
1418 |
1356 |
20.85 |
|
|
|
| 20 |
A |
1411 |
1350 |
1.32 |
|
|
|
| 21 |
A |
1374 |
1315 |
4.94 |
|
|
|
| 22 |
A |
1242 |
1188 |
37.77 |
|
|
|
| 23 |
A |
1220 |
1167 |
126.90 |
|
|
|
| 24 |
A |
1190 |
1139 |
6.08 |
|
|
|
| 25 |
A |
1186 |
1135 |
1.69 |
|
|
|
| 26 |
A |
1153 |
1103 |
16.84 |
|
|
|
| 27 |
A |
1088 |
1041 |
29.05 |
|
|
|
| 28 |
A |
943 |
902 |
0.11 |
|
|
|
| 29 |
A |
931 |
891 |
14.12 |
|
|
|
| 30 |
A |
924 |
884 |
0.39 |
|
|
|
| 31 |
A |
796 |
762 |
7.52 |
|
|
|
| 32 |
A |
552 |
528 |
2.23 |
|
|
|
| 33 |
A |
417 |
399 |
9.30 |
|
|
|
| 34 |
A |
364 |
348 |
1.24 |
|
|
|
| 35 |
A |
307 |
293 |
0.67 |
|
|
|
| 36 |
A |
257 |
246 |
1.51 |
|
|
|
| 37 |
A |
249 |
238 |
0.05 |
|
|
|
| 38 |
A |
179 |
172 |
1.71 |
|
|
|
| 39 |
A |
53 |
50 |
2.97 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30144.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 28838.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.262 |
|
|
0.018 |
| 2 |
H |
0.149 |
|
|
0.034 |
| 3 |
H |
0.179 |
|
|
0.087 |
| 4 |
H |
0.149 |
|
|
0.034 |
| 5 |
O |
-0.465 |
|
|
-0.447 |
| 6 |
C |
-0.530 |
|
|
-0.547 |
| 7 |
H |
0.168 |
|
|
0.129 |
| 8 |
H |
0.181 |
|
|
0.143 |
| 9 |
H |
0.165 |
|
|
0.147 |
| 10 |
C |
-0.530 |
|
|
-0.547 |
| 11 |
H |
0.168 |
|
|
0.129 |
| 12 |
H |
0.165 |
|
|
0.147 |
| 13 |
H |
0.181 |
|
|
0.143 |
| 14 |
C |
0.118 |
|
|
0.520 |
| 15 |
H |
0.163 |
|
|
0.009 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.586 |
0.001 |
1.147 |
1.288 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.569 |
0.001 |
1.184 |
1.314 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.951 |
0.000 |
2.187 |
| y |
0.000 |
-32.863 |
-0.001 |
| z |
2.187 |
-0.001 |
-33.660 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.311 |
0.000 |
2.187 |
| y |
0.000 |
-0.557 |
-0.001 |
| z |
2.187 |
-0.001 |
-1.753 |
|
| Polar |
|---|
| 3z2-r2 | -3.507 |
|---|
| x2-y2 | 1.912 |
|---|
| xy | 0.000 |
|---|
| xz | 2.187 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
133.808 |
| (<r2>)1/2 |
11.568 |