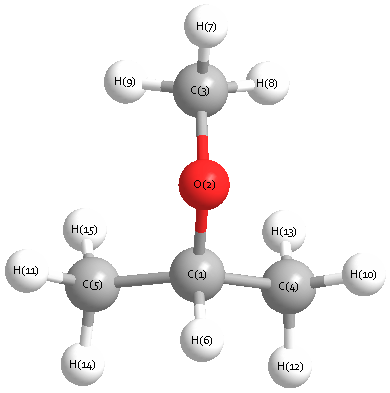
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3163 |
3008 |
33.73 |
|
|
|
| 2 |
A |
3161 |
3007 |
14.91 |
|
|
|
| 3 |
A |
3158 |
3004 |
18.96 |
|
|
|
| 4 |
A |
3152 |
2997 |
73.25 |
|
|
|
| 5 |
A |
3145 |
2991 |
0.25 |
|
|
|
| 6 |
A |
3083 |
2932 |
78.94 |
|
|
|
| 7 |
A |
3082 |
2931 |
21.23 |
|
|
|
| 8 |
A |
3075 |
2924 |
9.18 |
|
|
|
| 9 |
A |
3069 |
2918 |
2.13 |
|
|
|
| 10 |
A |
3026 |
2878 |
49.66 |
|
|
|
| 11 |
A |
1543 |
1468 |
14.02 |
|
|
|
| 12 |
A |
1538 |
1463 |
2.02 |
|
|
|
| 13 |
A |
1530 |
1455 |
2.25 |
|
|
|
| 14 |
A |
1525 |
1450 |
6.66 |
|
|
|
| 15 |
A |
1510 |
1436 |
0.10 |
|
|
|
| 16 |
A |
1508 |
1434 |
1.78 |
|
|
|
| 17 |
A |
1495 |
1422 |
0.48 |
|
|
|
| 18 |
A |
1438 |
1368 |
22.71 |
|
|
|
| 19 |
A |
1422 |
1352 |
23.69 |
|
|
|
| 20 |
A |
1414 |
1345 |
0.80 |
|
|
|
| 21 |
A |
1377 |
1310 |
3.87 |
|
|
|
| 22 |
A |
1250 |
1189 |
67.50 |
|
|
|
| 23 |
A |
1230 |
1170 |
101.88 |
|
|
|
| 24 |
A |
1195 |
1137 |
5.57 |
|
|
|
| 25 |
A |
1191 |
1133 |
1.72 |
|
|
|
| 26 |
A |
1161 |
1104 |
15.92 |
|
|
|
| 27 |
A |
1097 |
1044 |
25.27 |
|
|
|
| 28 |
A |
947 |
901 |
0.03 |
|
|
|
| 29 |
A |
942 |
895 |
13.58 |
|
|
|
| 30 |
A |
927 |
881 |
0.50 |
|
|
|
| 31 |
A |
806 |
767 |
7.47 |
|
|
|
| 32 |
A |
554 |
527 |
2.20 |
|
|
|
| 33 |
A |
420 |
400 |
9.35 |
|
|
|
| 34 |
A |
367 |
349 |
1.29 |
|
|
|
| 35 |
A |
313 |
298 |
0.61 |
|
|
|
| 36 |
A |
265 |
252 |
1.61 |
|
|
|
| 37 |
A |
252 |
240 |
0.07 |
|
|
|
| 38 |
A |
182 |
173 |
1.64 |
|
|
|
| 39 |
A |
50 |
48 |
2.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30281.6 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 28797.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.266 |
|
|
|
| 2 |
H |
0.150 |
|
|
|
| 3 |
H |
0.181 |
|
|
|
| 4 |
H |
0.150 |
|
|
|
| 5 |
O |
-0.466 |
|
|
|
| 6 |
C |
-0.535 |
|
|
|
| 7 |
H |
0.170 |
|
|
|
| 8 |
H |
0.183 |
|
|
|
| 9 |
H |
0.167 |
|
|
|
| 10 |
C |
-0.535 |
|
|
|
| 11 |
H |
0.170 |
|
|
|
| 12 |
H |
0.167 |
|
|
|
| 13 |
H |
0.183 |
|
|
|
| 14 |
C |
0.114 |
|
|
|
| 15 |
H |
0.166 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.588 |
0.002 |
1.144 |
1.287 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.045 |
0.001 |
2.184 |
| y |
0.001 |
-32.958 |
-0.002 |
| z |
2.184 |
-0.002 |
-33.755 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.311 |
0.001 |
2.184 |
| y |
0.001 |
-0.558 |
-0.002 |
| z |
2.184 |
-0.002 |
-1.753 |
|
| Polar |
|---|
| 3z2-r2 | -3.506 |
|---|
| x2-y2 | 1.913 |
|---|
| xy | 0.001 |
|---|
| xz | 2.184 |
|---|
| yz | -0.002 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.662 |
0.000 |
0.232 |
| y |
0.000 |
6.847 |
-0.000 |
| z |
0.232 |
-0.000 |
6.284 |
<r2> (average value of r
2) Å
2
| <r2> |
133.073 |
| (<r2>)1/2 |
11.536 |