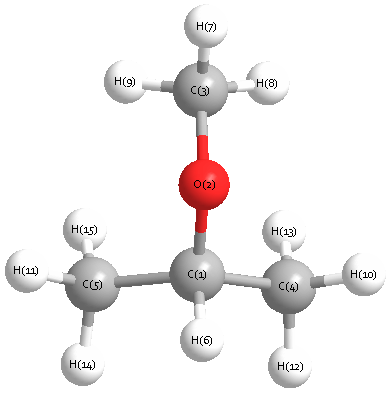
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3073 |
3022 |
30.46 |
|
|
|
| 2 |
A |
3072 |
3021 |
15.64 |
|
|
|
| 3 |
A |
3065 |
3014 |
31.87 |
|
|
|
| 4 |
A |
3061 |
3011 |
60.07 |
|
|
|
| 5 |
A |
3056 |
3006 |
0.10 |
|
|
|
| 6 |
A |
2990 |
2940 |
18.49 |
|
|
|
| 7 |
A |
2986 |
2937 |
46.57 |
|
|
|
| 8 |
A |
2984 |
2935 |
12.91 |
|
|
|
| 9 |
A |
2973 |
2924 |
38.30 |
|
|
|
| 10 |
A |
2927 |
2879 |
56.35 |
|
|
|
| 11 |
A |
1497 |
1473 |
12.76 |
|
|
|
| 12 |
A |
1490 |
1465 |
4.57 |
|
|
|
| 13 |
A |
1481 |
1457 |
1.44 |
|
|
|
| 14 |
A |
1479 |
1454 |
5.50 |
|
|
|
| 15 |
A |
1465 |
1441 |
0.23 |
|
|
|
| 16 |
A |
1460 |
1436 |
1.81 |
|
|
|
| 17 |
A |
1446 |
1423 |
0.40 |
|
|
|
| 18 |
A |
1386 |
1363 |
17.67 |
|
|
|
| 19 |
A |
1369 |
1346 |
19.01 |
|
|
|
| 20 |
A |
1364 |
1342 |
2.09 |
|
|
|
| 21 |
A |
1329 |
1307 |
6.14 |
|
|
|
| 22 |
A |
1199 |
1180 |
5.84 |
|
|
|
| 23 |
A |
1165 |
1146 |
135.61 |
|
|
|
| 24 |
A |
1155 |
1136 |
8.56 |
|
|
|
| 25 |
A |
1148 |
1129 |
0.66 |
|
|
|
| 26 |
A |
1119 |
1100 |
16.64 |
|
|
|
| 27 |
A |
1046 |
1028 |
34.81 |
|
|
|
| 28 |
A |
915 |
900 |
0.25 |
|
|
|
| 29 |
A |
897 |
883 |
0.32 |
|
|
|
| 30 |
A |
896 |
881 |
13.52 |
|
|
|
| 31 |
A |
770 |
758 |
7.81 |
|
|
|
| 32 |
A |
536 |
528 |
2.09 |
|
|
|
| 33 |
A |
403 |
396 |
8.81 |
|
|
|
| 34 |
A |
356 |
350 |
1.24 |
|
|
|
| 35 |
A |
300 |
295 |
0.50 |
|
|
|
| 36 |
A |
255 |
251 |
1.36 |
|
|
|
| 37 |
A |
250 |
246 |
0.06 |
|
|
|
| 38 |
A |
186 |
183 |
1.53 |
|
|
|
| 39 |
A |
70 |
69 |
2.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29309.3 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 28825.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.260 |
|
|
|
| 2 |
H |
0.141 |
|
|
|
| 3 |
H |
0.168 |
|
|
|
| 4 |
H |
0.141 |
|
|
|
| 5 |
O |
-0.418 |
|
|
|
| 6 |
C |
-0.501 |
|
|
|
| 7 |
H |
0.158 |
|
|
|
| 8 |
H |
0.170 |
|
|
|
| 9 |
H |
0.157 |
|
|
|
| 10 |
C |
-0.501 |
|
|
|
| 11 |
H |
0.158 |
|
|
|
| 12 |
H |
0.157 |
|
|
|
| 13 |
H |
0.170 |
|
|
|
| 14 |
C |
0.113 |
|
|
|
| 15 |
H |
0.148 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.577 |
0.001 |
1.077 |
1.221 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.211 |
0.000 |
2.053 |
| y |
0.000 |
-33.064 |
-0.001 |
| z |
2.053 |
-0.001 |
-33.815 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.229 |
0.000 |
2.053 |
| y |
0.000 |
-0.551 |
-0.001 |
| z |
2.053 |
-0.001 |
-1.678 |
|
| Polar |
|---|
| 3z2-r2 | -3.356 |
|---|
| x2-y2 | 1.853 |
|---|
| xy | 0.000 |
|---|
| xz | 2.053 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.031 |
0.000 |
0.262 |
| y |
0.000 |
7.104 |
0.000 |
| z |
0.262 |
0.000 |
6.483 |
<r2> (average value of r
2) Å
2
| <r2> |
135.156 |
| (<r2>)1/2 |
11.626 |