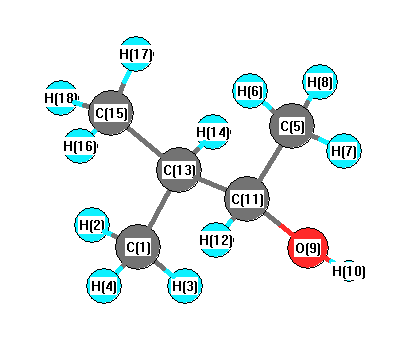
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3635 |
3575 |
3.74 |
|
|
|
| 2 |
A |
3090 |
3039 |
14.57 |
|
|
|
| 3 |
A |
3072 |
3021 |
24.52 |
|
|
|
| 4 |
A |
3062 |
3012 |
32.06 |
|
|
|
| 5 |
A |
3049 |
2999 |
75.14 |
|
|
|
| 6 |
A |
3045 |
2994 |
17.58 |
|
|
|
| 7 |
A |
3043 |
2993 |
7.33 |
|
|
|
| 8 |
A |
2980 |
2931 |
29.68 |
|
|
|
| 9 |
A |
2976 |
2927 |
40.39 |
|
|
|
| 10 |
A |
2975 |
2926 |
14.00 |
|
|
|
| 11 |
A |
2957 |
2908 |
16.76 |
|
|
|
| 12 |
A |
2871 |
2823 |
56.30 |
|
|
|
| 13 |
A |
1499 |
1475 |
6.66 |
|
|
|
| 14 |
A |
1489 |
1464 |
2.79 |
|
|
|
| 15 |
A |
1480 |
1455 |
13.70 |
|
|
|
| 16 |
A |
1477 |
1453 |
3.59 |
|
|
|
| 17 |
A |
1470 |
1446 |
3.37 |
|
|
|
| 18 |
A |
1469 |
1445 |
2.51 |
|
|
|
| 19 |
A |
1402 |
1379 |
8.96 |
|
|
|
| 20 |
A |
1386 |
1363 |
8.41 |
|
|
|
| 21 |
A |
1381 |
1359 |
10.50 |
|
|
|
| 22 |
A |
1373 |
1350 |
2.42 |
|
|
|
| 23 |
A |
1337 |
1315 |
6.59 |
|
|
|
| 24 |
A |
1315 |
1294 |
5.88 |
|
|
|
| 25 |
A |
1308 |
1287 |
14.74 |
|
|
|
| 26 |
A |
1245 |
1224 |
25.95 |
|
|
|
| 27 |
A |
1169 |
1150 |
3.95 |
|
|
|
| 28 |
A |
1154 |
1135 |
10.70 |
|
|
|
| 29 |
A |
1127 |
1109 |
4.36 |
|
|
|
| 30 |
A |
1075 |
1057 |
79.06 |
|
|
|
| 31 |
A |
1059 |
1041 |
8.31 |
|
|
|
| 32 |
A |
968 |
952 |
5.89 |
|
|
|
| 33 |
A |
949 |
934 |
3.78 |
|
|
|
| 34 |
A |
929 |
914 |
13.41 |
|
|
|
| 35 |
A |
912 |
897 |
1.51 |
|
|
|
| 36 |
A |
879 |
865 |
7.81 |
|
|
|
| 37 |
A |
772 |
759 |
2.12 |
|
|
|
| 38 |
A |
512 |
504 |
3.56 |
|
|
|
| 39 |
A |
448 |
441 |
8.02 |
|
|
|
| 40 |
A |
399 |
393 |
0.83 |
|
|
|
| 41 |
A |
359 |
353 |
6.38 |
|
|
|
| 42 |
A |
352 |
347 |
2.18 |
|
|
|
| 43 |
A |
281 |
276 |
82.45 |
|
|
|
| 44 |
A |
276 |
271 |
22.28 |
|
|
|
| 45 |
A |
229 |
225 |
0.46 |
|
|
|
| 46 |
A |
222 |
219 |
0.09 |
|
|
|
| 47 |
A |
207 |
203 |
0.30 |
|
|
|
| 48 |
A |
78 |
77 |
1.55 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35370.9 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 34787.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.484 |
|
|
|
| 2 |
H |
0.148 |
|
|
|
| 3 |
H |
0.182 |
|
|
|
| 4 |
H |
0.144 |
|
|
|
| 5 |
C |
-0.507 |
|
|
|
| 6 |
H |
0.161 |
|
|
|
| 7 |
H |
0.144 |
|
|
|
| 8 |
H |
0.167 |
|
|
|
| 9 |
O |
-0.593 |
|
|
|
| 10 |
H |
0.381 |
|
|
|
| 11 |
C |
0.117 |
|
|
|
| 12 |
H |
0.115 |
|
|
|
| 13 |
C |
-0.096 |
|
|
|
| 14 |
H |
0.144 |
|
|
|
| 15 |
C |
-0.492 |
|
|
|
| 16 |
H |
0.152 |
|
|
|
| 17 |
H |
0.157 |
|
|
|
| 18 |
H |
0.159 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.858 |
0.857 |
0.752 |
1.427 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.906 |
-0.944 |
1.883 |
| y |
-0.944 |
-41.342 |
-1.177 |
| z |
1.883 |
-1.177 |
-39.541 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.535 |
-0.944 |
1.883 |
| y |
-0.944 |
-3.618 |
-1.177 |
| z |
1.883 |
-1.177 |
-0.917 |
|
| Polar |
|---|
| 3z2-r2 | -1.834 |
|---|
| x2-y2 | 5.436 |
|---|
| xy | -0.944 |
|---|
| xz | 1.883 |
|---|
| yz | -1.177 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.797 |
0.021 |
0.138 |
| y |
0.021 |
8.929 |
0.018 |
| z |
0.138 |
0.018 |
7.765 |
<r2> (average value of r
2) Å
2
| <r2> |
196.908 |
| (<r2>)1/2 |
14.032 |