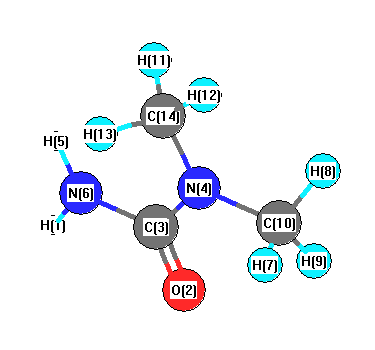
Vibrational Frequencies calculated at B3LYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3665 |
3520 |
21.87 |
|
|
|
| 2 |
A |
3558 |
3417 |
16.39 |
|
|
|
| 3 |
A |
3190 |
3063 |
0.78 |
|
|
|
| 4 |
A |
3141 |
3017 |
11.61 |
|
|
|
| 5 |
A |
3087 |
2965 |
26.32 |
|
|
|
| 6 |
A |
3069 |
2947 |
51.68 |
|
|
|
| 7 |
A |
3025 |
2905 |
70.11 |
|
|
|
| 8 |
A |
3000 |
2881 |
67.75 |
|
|
|
| 9 |
A |
1793 |
1722 |
363.85 |
|
|
|
| 10 |
A |
1661 |
1595 |
123.81 |
|
|
|
| 11 |
A |
1564 |
1502 |
44.23 |
|
|
|
| 12 |
A |
1538 |
1477 |
14.09 |
|
|
|
| 13 |
A |
1529 |
1468 |
14.04 |
|
|
|
| 14 |
A |
1528 |
1467 |
12.46 |
|
|
|
| 15 |
A |
1501 |
1441 |
58.12 |
|
|
|
| 16 |
A |
1455 |
1397 |
7.55 |
|
|
|
| 17 |
A |
1445 |
1388 |
171.21 |
|
|
|
| 18 |
A |
1307 |
1256 |
42.52 |
|
|
|
| 19 |
A |
1284 |
1233 |
43.15 |
|
|
|
| 20 |
A |
1184 |
1137 |
3.16 |
|
|
|
| 21 |
A |
1143 |
1098 |
3.41 |
|
|
|
| 22 |
A |
1117 |
1073 |
68.23 |
|
|
|
| 23 |
A |
1096 |
1053 |
11.75 |
|
|
|
| 24 |
A |
1037 |
996 |
35.06 |
|
|
|
| 25 |
A |
782 |
750 |
93.10 |
|
|
|
| 26 |
A |
774 |
743 |
2.35 |
|
|
|
| 27 |
A |
616 |
591 |
145.98 |
|
|
|
| 28 |
A |
595 |
571 |
40.49 |
|
|
|
| 29 |
A |
515 |
494 |
8.57 |
|
|
|
| 30 |
A |
446 |
428 |
29.70 |
|
|
|
| 31 |
A |
393 |
377 |
13.68 |
|
|
|
| 32 |
A |
319 |
306 |
9.29 |
|
|
|
| 33 |
A |
194 |
186 |
6.05 |
|
|
|
| 34 |
A |
153 |
147 |
1.42 |
|
|
|
| 35 |
A |
133 |
128 |
5.01 |
|
|
|
| 36 |
A |
88 |
84 |
1.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26460.8 cm
-1
Scaled (by 0.9603) Zero Point Vibrational Energy (zpe) 25410.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.338 |
0.366 |
|
0.373 |
| 2 |
O |
-0.531 |
-0.550 |
|
-0.537 |
| 3 |
C |
0.696 |
0.664 |
|
0.612 |
| 4 |
N |
-0.407 |
-0.177 |
|
-0.056 |
| 5 |
H |
0.321 |
0.341 |
|
0.343 |
| 6 |
N |
-0.762 |
-0.841 |
|
-0.843 |
| 7 |
H |
0.148 |
0.073 |
|
0.117 |
| 8 |
H |
0.145 |
0.063 |
|
0.108 |
| 9 |
H |
0.206 |
0.094 |
|
0.139 |
| 10 |
C |
-0.316 |
-0.143 |
|
-0.326 |
| 11 |
H |
0.148 |
0.039 |
|
0.074 |
| 12 |
H |
0.164 |
0.072 |
|
0.107 |
| 13 |
H |
0.178 |
0.055 |
|
0.094 |
| 14 |
C |
-0.329 |
-0.056 |
|
-0.205 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.697 |
3.155 |
1.203 |
3.779 |
| CHELPG |
-1.726 |
3.153 |
1.223 |
3.796 |
| AIM |
|
|
|
|
| ESP |
-1.731 |
3.158 |
1.220 |
3.802 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.334 |
4.549 |
2.273 |
| y |
4.549 |
-37.228 |
1.365 |
| z |
2.273 |
1.365 |
-37.024 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.792 |
4.549 |
2.273 |
| y |
4.549 |
-2.049 |
1.365 |
| z |
2.273 |
1.365 |
-1.743 |
|
| Polar |
|---|
| 3z2-r2 | -3.486 |
|---|
| x2-y2 | 3.894 |
|---|
| xy | 4.549 |
|---|
| xz | 2.273 |
|---|
| yz | 1.365 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.661 |
-0.055 |
-0.045 |
| y |
-0.055 |
7.820 |
0.246 |
| z |
-0.045 |
0.246 |
4.905 |
<r2> (average value of r
2) Å
2
| <r2> |
166.378 |
| (<r2>)1/2 |
12.899 |