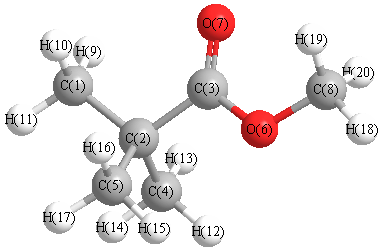
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3191 |
3053 |
16.56 |
|
|
|
| 2 |
A |
3160 |
3023 |
22.38 |
|
|
|
| 3 |
A |
3157 |
3020 |
20.39 |
|
|
|
| 4 |
A |
3156 |
3019 |
35.01 |
|
|
|
| 5 |
A |
3153 |
3016 |
0.36 |
|
|
|
| 6 |
A |
3145 |
3009 |
32.04 |
|
|
|
| 7 |
A |
3131 |
2995 |
25.22 |
|
|
|
| 8 |
A |
3129 |
2994 |
12.11 |
|
|
|
| 9 |
A |
3079 |
2946 |
32.90 |
|
|
|
| 10 |
A |
3068 |
2935 |
24.15 |
|
|
|
| 11 |
A |
3059 |
2926 |
12.98 |
|
|
|
| 12 |
A |
3057 |
2924 |
29.88 |
|
|
|
| 13 |
A |
1834 |
1755 |
199.89 |
|
|
|
| 14 |
A |
1547 |
1480 |
27.85 |
|
|
|
| 15 |
A |
1528 |
1462 |
10.85 |
|
|
|
| 16 |
A |
1524 |
1458 |
6.64 |
|
|
|
| 17 |
A |
1521 |
1455 |
6.90 |
|
|
|
| 18 |
A |
1514 |
1449 |
0.18 |
|
|
|
| 19 |
A |
1509 |
1443 |
0.12 |
|
|
|
| 20 |
A |
1508 |
1443 |
7.04 |
|
|
|
| 21 |
A |
1502 |
1437 |
0.02 |
|
|
|
| 22 |
A |
1488 |
1424 |
7.72 |
|
|
|
| 23 |
A |
1451 |
1388 |
11.12 |
|
|
|
| 24 |
A |
1423 |
1362 |
5.52 |
|
|
|
| 25 |
A |
1417 |
1355 |
6.00 |
|
|
|
| 26 |
A |
1331 |
1273 |
92.78 |
|
|
|
| 27 |
A |
1271 |
1216 |
6.86 |
|
|
|
| 28 |
A |
1247 |
1193 |
2.20 |
|
|
|
| 29 |
A |
1229 |
1175 |
191.38 |
|
|
|
| 30 |
A |
1200 |
1148 |
127.58 |
|
|
|
| 31 |
A |
1184 |
1133 |
0.66 |
|
|
|
| 32 |
A |
1072 |
1026 |
10.61 |
|
|
|
| 33 |
A |
1059 |
1013 |
0.37 |
|
|
|
| 34 |
A |
1032 |
987 |
4.04 |
|
|
|
| 35 |
A |
976 |
934 |
0.05 |
|
|
|
| 36 |
A |
961 |
919 |
3.03 |
|
|
|
| 37 |
A |
955 |
914 |
1.74 |
|
|
|
| 38 |
A |
887 |
849 |
13.38 |
|
|
|
| 39 |
A |
805 |
771 |
1.78 |
|
|
|
| 40 |
A |
779 |
746 |
5.08 |
|
|
|
| 41 |
A |
591 |
566 |
2.27 |
|
|
|
| 42 |
A |
497 |
476 |
2.67 |
|
|
|
| 43 |
A |
378 |
362 |
0.93 |
|
|
|
| 44 |
A |
362 |
346 |
11.12 |
|
|
|
| 45 |
A |
340 |
325 |
0.46 |
|
|
|
| 46 |
A |
320 |
306 |
7.21 |
|
|
|
| 47 |
A |
289 |
276 |
2.45 |
|
|
|
| 48 |
A |
264 |
253 |
0.06 |
|
|
|
| 49 |
A |
256 |
245 |
1.81 |
|
|
|
| 50 |
A |
212 |
203 |
0.27 |
|
|
|
| 51 |
A |
199 |
190 |
1.73 |
|
|
|
| 52 |
A |
133 |
127 |
1.45 |
|
|
|
| 53 |
A |
118 |
113 |
0.86 |
|
|
|
| 54 |
A |
36 |
34 |
0.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38615.7 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 36943.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.493 |
|
|
|
| 2 |
C |
-0.062 |
|
|
|
| 3 |
C |
0.657 |
|
|
|
| 4 |
C |
-0.508 |
|
|
|
| 5 |
C |
-0.508 |
|
|
|
| 6 |
O |
-0.459 |
|
|
|
| 7 |
O |
-0.488 |
|
|
|
| 8 |
C |
-0.282 |
|
|
|
| 9 |
H |
0.180 |
|
|
|
| 10 |
H |
0.180 |
|
|
|
| 11 |
H |
0.159 |
|
|
|
| 12 |
H |
0.186 |
|
|
|
| 13 |
H |
0.174 |
|
|
|
| 14 |
H |
0.170 |
|
|
|
| 15 |
H |
0.186 |
|
|
|
| 16 |
H |
0.174 |
|
|
|
| 17 |
H |
0.170 |
|
|
|
| 18 |
H |
0.184 |
|
|
|
| 19 |
H |
0.190 |
|
|
|
| 20 |
H |
0.190 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.161 |
-1.744 |
0.000 |
1.751 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.971 |
1.743 |
0.001 |
| y |
1.743 |
-54.168 |
0.000 |
| z |
0.001 |
0.000 |
-48.215 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
8.220 |
1.743 |
0.001 |
| y |
1.743 |
-8.575 |
0.000 |
| z |
0.001 |
0.000 |
0.355 |
|
| Polar |
|---|
| 3z2-r2 | 0.709 |
|---|
| x2-y2 | 11.197 |
|---|
| xy | 1.743 |
|---|
| xz | 0.001 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.444 |
0.156 |
-0.000 |
| y |
0.156 |
10.021 |
0.000 |
| z |
-0.000 |
0.000 |
8.917 |
<r2> (average value of r
2) Å
2
| <r2> |
296.446 |
| (<r2>)1/2 |
17.218 |