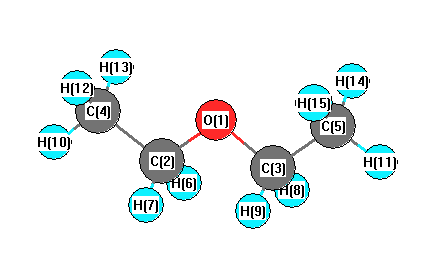
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3148 |
3012 |
35.82 |
|
|
|
| 2 |
A1 |
3069 |
2936 |
7.59 |
|
|
|
| 3 |
A1 |
2982 |
2852 |
110.51 |
|
|
|
| 4 |
A1 |
1566 |
1498 |
0.72 |
|
|
|
| 5 |
A1 |
1528 |
1462 |
4.18 |
|
|
|
| 6 |
A1 |
1472 |
1408 |
5.98 |
|
|
|
| 7 |
A1 |
1420 |
1359 |
0.53 |
|
|
|
| 8 |
A1 |
1197 |
1145 |
23.40 |
|
|
|
| 9 |
A1 |
1076 |
1029 |
11.51 |
|
|
|
| 10 |
A1 |
873 |
835 |
5.69 |
|
|
|
| 11 |
A1 |
443 |
424 |
0.04 |
|
|
|
| 12 |
A1 |
193 |
185 |
0.71 |
|
|
|
| 13 |
A2 |
3153 |
3016 |
0.00 |
|
|
|
| 14 |
A2 |
3003 |
2873 |
0.00 |
|
|
|
| 15 |
A2 |
1509 |
1444 |
0.00 |
|
|
|
| 16 |
A2 |
1306 |
1250 |
0.00 |
|
|
|
| 17 |
A2 |
1182 |
1131 |
0.00 |
|
|
|
| 18 |
A2 |
824 |
789 |
0.00 |
|
|
|
| 19 |
A2 |
255 |
244 |
0.00 |
|
|
|
| 20 |
A2 |
105 |
100 |
0.00 |
|
|
|
| 21 |
B1 |
3153 |
3016 |
54.63 |
|
|
|
| 22 |
B1 |
3000 |
2870 |
140.73 |
|
|
|
| 23 |
B1 |
1510 |
1444 |
11.16 |
|
|
|
| 24 |
B1 |
1313 |
1256 |
3.07 |
|
|
|
| 25 |
B1 |
1215 |
1162 |
13.01 |
|
|
|
| 26 |
B1 |
837 |
800 |
1.37 |
|
|
|
| 27 |
B1 |
259 |
248 |
1.51 |
|
|
|
| 28 |
B1 |
107 |
103 |
4.12 |
|
|
|
| 29 |
B2 |
3148 |
3012 |
17.92 |
|
|
|
| 30 |
B2 |
3068 |
2935 |
28.26 |
|
|
|
| 31 |
B2 |
2969 |
2840 |
10.41 |
|
|
|
| 32 |
B2 |
1544 |
1477 |
3.86 |
|
|
|
| 33 |
B2 |
1525 |
1459 |
2.96 |
|
|
|
| 34 |
B2 |
1436 |
1374 |
42.23 |
|
|
|
| 35 |
B2 |
1395 |
1335 |
47.81 |
|
|
|
| 36 |
B2 |
1195 |
1143 |
205.54 |
|
|
|
| 37 |
B2 |
1111 |
1063 |
3.30 |
|
|
|
| 38 |
B2 |
960 |
918 |
3.65 |
|
|
|
| 39 |
B2 |
434 |
415 |
3.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30240.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 28930.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.465 |
|
|
-0.411 |
| 2 |
C |
-0.050 |
|
|
0.306 |
| 3 |
C |
-0.050 |
|
|
0.306 |
| 4 |
C |
-0.522 |
|
|
-0.341 |
| 5 |
C |
-0.522 |
|
|
-0.341 |
| 6 |
H |
0.139 |
|
|
-0.020 |
| 7 |
H |
0.139 |
|
|
-0.022 |
| 8 |
H |
0.139 |
|
|
-0.020 |
| 9 |
H |
0.139 |
|
|
-0.022 |
| 10 |
H |
0.165 |
|
|
0.084 |
| 11 |
H |
0.165 |
|
|
0.084 |
| 12 |
H |
0.181 |
|
|
0.100 |
| 13 |
H |
0.181 |
|
|
0.099 |
| 14 |
H |
0.181 |
|
|
0.099 |
| 15 |
H |
0.181 |
|
|
0.100 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.035 |
1.035 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.008 |
0.000 |
-1.019 |
1.019 |
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
176.020 |
| (<r2>)1/2 |
13.267 |