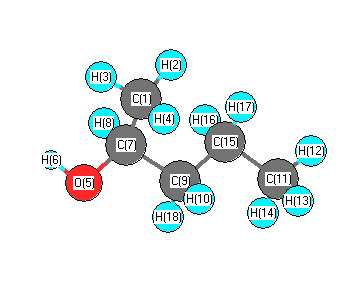
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3780 |
3617 |
8.07 |
|
|
|
| 2 |
A |
3143 |
3007 |
28.63 |
|
|
|
| 3 |
A |
3131 |
2995 |
37.50 |
|
|
|
| 4 |
A |
3127 |
2992 |
61.34 |
|
|
|
| 5 |
A |
3120 |
2985 |
32.05 |
|
|
|
| 6 |
A |
3099 |
2965 |
32.47 |
|
|
|
| 7 |
A |
3072 |
2939 |
3.66 |
|
|
|
| 8 |
A |
3053 |
2921 |
42.80 |
|
|
|
| 9 |
A |
3049 |
2917 |
19.99 |
|
|
|
| 10 |
A |
3048 |
2916 |
38.10 |
|
|
|
| 11 |
A |
3037 |
2906 |
9.67 |
|
|
|
| 12 |
A |
2970 |
2841 |
59.19 |
|
|
|
| 13 |
A |
1535 |
1469 |
6.05 |
|
|
|
| 14 |
A |
1526 |
1460 |
4.98 |
|
|
|
| 15 |
A |
1525 |
1459 |
1.78 |
|
|
|
| 16 |
A |
1524 |
1458 |
7.78 |
|
|
|
| 17 |
A |
1519 |
1453 |
1.40 |
|
|
|
| 18 |
A |
1508 |
1442 |
0.45 |
|
|
|
| 19 |
A |
1448 |
1385 |
19.59 |
|
|
|
| 20 |
A |
1437 |
1375 |
3.89 |
|
|
|
| 21 |
A |
1432 |
1370 |
19.35 |
|
|
|
| 22 |
A |
1408 |
1347 |
9.75 |
|
|
|
| 23 |
A |
1391 |
1331 |
3.94 |
|
|
|
| 24 |
A |
1340 |
1282 |
1.20 |
|
|
|
| 25 |
A |
1325 |
1268 |
21.81 |
|
|
|
| 26 |
A |
1296 |
1240 |
6.00 |
|
|
|
| 27 |
A |
1258 |
1203 |
16.95 |
|
|
|
| 28 |
A |
1196 |
1145 |
15.21 |
|
|
|
| 29 |
A |
1163 |
1112 |
35.81 |
|
|
|
| 30 |
A |
1112 |
1064 |
47.34 |
|
|
|
| 31 |
A |
1104 |
1056 |
1.58 |
|
|
|
| 32 |
A |
1058 |
1012 |
2.82 |
|
|
|
| 33 |
A |
1015 |
971 |
4.65 |
|
|
|
| 34 |
A |
976 |
934 |
9.56 |
|
|
|
| 35 |
A |
925 |
885 |
20.30 |
|
|
|
| 36 |
A |
884 |
846 |
0.81 |
|
|
|
| 37 |
A |
847 |
810 |
0.29 |
|
|
|
| 38 |
A |
753 |
720 |
3.50 |
|
|
|
| 39 |
A |
492 |
470 |
8.58 |
|
|
|
| 40 |
A |
450 |
430 |
1.28 |
|
|
|
| 41 |
A |
412 |
394 |
9.23 |
|
|
|
| 42 |
A |
320 |
306 |
1.00 |
|
|
|
| 43 |
A |
310 |
297 |
120.66 |
|
|
|
| 44 |
A |
267 |
255 |
1.01 |
|
|
|
| 45 |
A |
248 |
237 |
0.53 |
|
|
|
| 46 |
A |
183 |
175 |
1.59 |
|
|
|
| 47 |
A |
115 |
110 |
0.01 |
|
|
|
| 48 |
A |
81 |
78 |
0.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36506.8 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 34926.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.531 |
|
|
-0.433 |
| 2 |
H |
0.170 |
|
|
0.099 |
| 3 |
H |
0.155 |
|
|
0.106 |
| 4 |
H |
0.178 |
|
|
0.128 |
| 5 |
O |
-0.637 |
|
|
-0.652 |
| 6 |
H |
0.398 |
|
|
0.415 |
| 7 |
C |
0.110 |
|
|
0.298 |
| 8 |
H |
0.133 |
|
|
0.004 |
| 9 |
C |
-0.305 |
|
|
-0.148 |
| 10 |
H |
0.163 |
|
|
0.084 |
| 11 |
C |
-0.507 |
|
|
-0.321 |
| 12 |
H |
0.166 |
|
|
0.083 |
| 13 |
H |
0.167 |
|
|
0.078 |
| 14 |
H |
0.167 |
|
|
0.083 |
| 15 |
C |
-0.316 |
|
|
0.073 |
| 16 |
H |
0.157 |
|
|
0.010 |
| 17 |
H |
0.160 |
|
|
0.021 |
| 18 |
H |
0.172 |
|
|
0.077 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.411 |
1.294 |
-0.726 |
1.539 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.408 |
1.292 |
-0.713 |
1.531 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.178 |
-1.721 |
1.810 |
| y |
-1.721 |
-41.786 |
0.775 |
| z |
1.810 |
0.775 |
-39.239 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.335 |
-1.721 |
1.810 |
| y |
-1.721 |
-4.578 |
0.775 |
| z |
1.810 |
0.775 |
-0.757 |
|
| Polar |
|---|
| 3z2-r2 | -1.514 |
|---|
| x2-y2 | 6.608 |
|---|
| xy | -1.721 |
|---|
| xz | 1.810 |
|---|
| yz | 0.775 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
239.273 |
| (<r2>)1/2 |
15.468 |