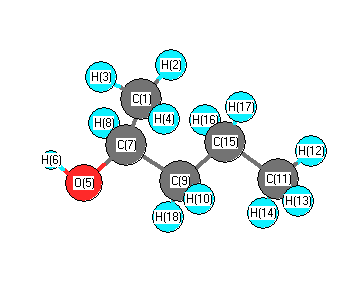
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3808 |
3619 |
10.17 |
|
|
|
| 2 |
A |
3160 |
3003 |
25.60 |
|
|
|
| 3 |
A |
3149 |
2993 |
32.72 |
|
|
|
| 4 |
A |
3145 |
2988 |
55.30 |
|
|
|
| 5 |
A |
3137 |
2981 |
29.78 |
|
|
|
| 6 |
A |
3115 |
2960 |
31.57 |
|
|
|
| 7 |
A |
3088 |
2935 |
2.91 |
|
|
|
| 8 |
A |
3068 |
2915 |
40.94 |
|
|
|
| 9 |
A |
3063 |
2911 |
16.59 |
|
|
|
| 10 |
A |
3063 |
2910 |
37.61 |
|
|
|
| 11 |
A |
3053 |
2901 |
7.66 |
|
|
|
| 12 |
A |
2986 |
2837 |
57.26 |
|
|
|
| 13 |
A |
1537 |
1461 |
6.48 |
|
|
|
| 14 |
A |
1528 |
1452 |
2.58 |
|
|
|
| 15 |
A |
1528 |
1452 |
4.27 |
|
|
|
| 16 |
A |
1526 |
1450 |
8.83 |
|
|
|
| 17 |
A |
1521 |
1445 |
1.39 |
|
|
|
| 18 |
A |
1509 |
1434 |
0.47 |
|
|
|
| 19 |
A |
1456 |
1384 |
20.67 |
|
|
|
| 20 |
A |
1440 |
1368 |
3.05 |
|
|
|
| 21 |
A |
1436 |
1365 |
24.25 |
|
|
|
| 22 |
A |
1413 |
1342 |
8.49 |
|
|
|
| 23 |
A |
1395 |
1326 |
3.91 |
|
|
|
| 24 |
A |
1343 |
1276 |
1.40 |
|
|
|
| 25 |
A |
1328 |
1262 |
25.71 |
|
|
|
| 26 |
A |
1300 |
1236 |
5.12 |
|
|
|
| 27 |
A |
1261 |
1198 |
15.30 |
|
|
|
| 28 |
A |
1202 |
1142 |
16.99 |
|
|
|
| 29 |
A |
1173 |
1115 |
36.64 |
|
|
|
| 30 |
A |
1120 |
1064 |
48.11 |
|
|
|
| 31 |
A |
1117 |
1061 |
0.81 |
|
|
|
| 32 |
A |
1071 |
1017 |
2.36 |
|
|
|
| 33 |
A |
1019 |
968 |
4.55 |
|
|
|
| 34 |
A |
984 |
935 |
7.02 |
|
|
|
| 35 |
A |
932 |
886 |
18.71 |
|
|
|
| 36 |
A |
886 |
842 |
0.86 |
|
|
|
| 37 |
A |
854 |
811 |
0.33 |
|
|
|
| 38 |
A |
755 |
717 |
3.60 |
|
|
|
| 39 |
A |
494 |
470 |
8.66 |
|
|
|
| 40 |
A |
452 |
430 |
1.19 |
|
|
|
| 41 |
A |
416 |
395 |
9.62 |
|
|
|
| 42 |
A |
323 |
307 |
1.08 |
|
|
|
| 43 |
A |
314 |
298 |
120.89 |
|
|
|
| 44 |
A |
269 |
256 |
1.07 |
|
|
|
| 45 |
A |
250 |
238 |
0.56 |
|
|
|
| 46 |
A |
185 |
176 |
1.63 |
|
|
|
| 47 |
A |
116 |
111 |
0.01 |
|
|
|
| 48 |
A |
82 |
78 |
0.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36684.0 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 34860.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.542 |
|
|
|
| 2 |
H |
0.175 |
|
|
|
| 3 |
H |
0.160 |
|
|
|
| 4 |
H |
0.182 |
|
|
|
| 5 |
O |
-0.639 |
|
|
|
| 6 |
H |
0.402 |
|
|
|
| 7 |
C |
0.102 |
|
|
|
| 8 |
H |
0.138 |
|
|
|
| 9 |
C |
-0.315 |
|
|
|
| 10 |
H |
0.168 |
|
|
|
| 11 |
C |
-0.519 |
|
|
|
| 12 |
H |
0.171 |
|
|
|
| 13 |
H |
0.171 |
|
|
|
| 14 |
H |
0.172 |
|
|
|
| 15 |
C |
-0.329 |
|
|
|
| 16 |
H |
0.163 |
|
|
|
| 17 |
H |
0.165 |
|
|
|
| 18 |
H |
0.177 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.419 |
1.293 |
-0.719 |
1.538 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.253 |
-1.726 |
1.781 |
| y |
-1.726 |
-41.877 |
0.761 |
| z |
1.781 |
0.761 |
-39.338 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.354 |
-1.726 |
1.781 |
| y |
-1.726 |
-4.582 |
0.761 |
| z |
1.781 |
0.761 |
-0.773 |
|
| Polar |
|---|
| 3z2-r2 | -1.545 |
|---|
| x2-y2 | 6.624 |
|---|
| xy | -1.726 |
|---|
| xz | 1.781 |
|---|
| yz | 0.761 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.959 |
-0.094 |
0.047 |
| y |
-0.094 |
8.060 |
-0.074 |
| z |
0.047 |
-0.074 |
7.609 |
<r2> (average value of r
2) Å
2
| <r2> |
238.040 |
| (<r2>)1/2 |
15.429 |