return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12O (2-Pentanol)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-272.639854
Energy at 298.15K 
HF Energy-272.364665
Nuclear repulsion energy254.360947
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3761 3570 8.28      
2 A 3166 3005 28.10      
3 A 3152 2992 41.20      
4 A 3150 2990 59.10      
5 A 3143 2983 31.36      
6 A 3121 2963 34.03      
7 A 3097 2940 3.02      
8 A 3077 2921 37.49      
9 A 3073 2917 19.08      
10 A 3072 2916 38.09      
11 A 3062 2907 7.55      
12 A 3005 2852 54.10      
13 A 1562 1482 4.13      
14 A 1551 1473 3.55      
15 A 1550 1471 4.27      
16 A 1548 1470 4.54      
17 A 1545 1466 1.61      
18 A 1534 1456 0.35      
19 A 1467 1393 19.94      
20 A 1461 1387 2.22      
21 A 1456 1382 13.36      
22 A 1430 1358 10.88      
23 A 1410 1339 4.28      
24 A 1358 1289 0.85      
25 A 1343 1274 17.19      
26 A 1314 1247 6.85      
27 A 1276 1211 19.59      
28 A 1216 1154 15.64      
29 A 1170 1111 27.85      
30 A 1125 1068 44.64      
31 A 1109 1053 3.48      
32 A 1063 1009 2.99      
33 A 1029 977 4.53      
34 A 985 935 13.67      
35 A 932 885 21.37      
36 A 897 852 0.58      
37 A 855 811 0.19      
38 A 763 724 2.46      
39 A 498 473 9.20      
40 A 455 432 1.24      
41 A 417 395 9.20      
42 A 326 309 0.40      
43 A 317 301 122.21      
44 A 271 257 0.70      
45 A 251 238 0.37      
46 A 187 178 1.69      
47 A 115 109 0.01      
48 A 80 76 0.54      

Unscaled Zero Point Vibrational Energy (zpe) 36871.7 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 34998.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
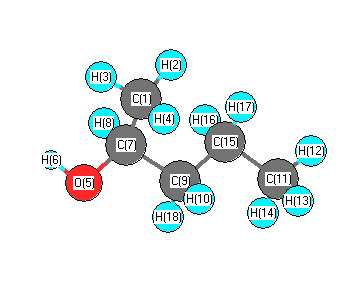
C1 -1.532 1.291 0.207
H2 -0.872 2.099 -0.121
H3 -2.542 1.528 -0.146
H4 -1.549 1.272 1.301
O5 -1.979 -1.096 0.086
H6 -2.866 -0.848 -0.216
C7 -1.078 -0.064 -0.330
H8 -1.049 -0.020 -1.431
C9 0.296 -0.493 0.174
H10 0.269 -0.518 1.271
C11 2.815 -0.151 0.145
H12 3.631 0.489 -0.203
H13 2.874 -0.205 1.237
H14 2.988 -1.159 -0.246
C15 1.455 0.385 -0.304
H16 1.433 0.452 -1.399
H17 1.330 1.407 0.069
H18 0.457 -1.526 -0.157

Atom - Atom Distances (Å)
  C1 H2 H3 H4 O5 H6 C7 H8 C9 H10 C11 H12 H13 H14 C15 H16 H17 H18
C11.09331.09581.09382.43172.55641.52612.15292.55442.76494.57995.24034.76505.16103.16243.47492.86753.4669
H21.09331.76431.77843.38763.55932.18232.49722.85843.17594.32764.78274.60265.05282.89583.10842.31613.8606
H31.09581.76431.77302.69392.39922.17002.50523.49863.75395.62106.25935.85166.14894.15954.30433.87954.2798
H41.09381.77841.77302.69652.92102.16003.06302.79122.55164.73345.45014.66345.37493.51934.10573.13453.7385
O52.43173.38762.69392.69650.96941.43202.07942.35552.60624.88675.83665.06664.97913.75984.03044.14972.4858
H62.55643.55932.39922.92100.96941.95572.33753.20613.48565.73516.63315.95595.86314.49414.64464.77263.3924
C71.52612.18232.17002.16001.43201.95571.10221.52562.14083.92304.74294.25364.21242.57242.77772.85022.1271
H82.15292.49722.50523.06302.07942.33751.10222.14683.04674.17474.86464.74724.35882.77482.52633.15372.4815
C92.55442.85843.49862.79122.35553.20611.52562.14681.09712.54203.49652.80292.80501.53032.15902.16631.0959
H102.76493.17593.75392.55162.60623.48562.14083.04671.09712.80773.80612.62383.17902.16833.07002.50561.7572
C114.57994.32765.62104.73344.88675.73513.92304.17472.54202.80771.09381.09451.09441.52962.15852.15402.7456
H125.24034.78276.25935.45015.83666.63314.74294.86463.49653.80611.09381.76821.76872.18092.50272.49203.7591
H134.76504.60265.85164.66345.06665.95594.25364.74722.80292.62381.09451.76821.76642.17643.07522.51943.0862
H145.16105.05286.14895.37494.97915.86314.21244.35882.80503.17901.09441.76871.76642.17662.51853.07142.5593
C153.16242.89584.15953.51933.75984.49412.57242.77481.53032.16831.52962.18092.17642.17661.09741.09582.1600
H163.47493.10844.30434.10574.03044.64462.77772.52632.15903.07002.15852.50273.07522.51851.09741.75512.5309
H172.86752.31613.87953.13454.14974.77262.85023.15372.16632.50562.15402.49202.51943.07141.09581.75513.0684
H183.46693.86064.27983.73852.48583.39242.12712.48151.09591.75722.74563.75913.08622.55932.16002.53093.0684

picture of 2-Pentanol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C7 O5 110.542 C1 C7 H8 108.927
C1 C7 C9 113.654 H2 C1 H3 107.401
H2 C1 H4 108.798 H2 C1 C7 111.781
H3 C1 H4 108.138 H3 C1 C7 110.641
H4 C1 C7 109.970 O5 C7 H8 109.597
O5 C7 C9 105.538 H6 O5 C7 107.496
C7 C9 H10 108.317 C7 C9 C15 114.654
C7 C9 H18 107.327 H8 C7 C9 108.487
C9 C15 C11 112.354 C9 C15 H16 109.397
C9 C15 H17 110.057 H10 C9 C15 110.138
H10 C9 H18 106.504 C11 C15 H16 109.401
C11 C15 H17 109.145 H12 C11 H13 107.814
H12 C11 H14 107.858 H12 C11 C15 111.392
H13 C11 H14 107.611 H13 C11 C15 110.991
H14 C11 C15 111.011 C15 C9 H18 109.555
H16 C15 H17 106.304
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.483      
2 H 0.156      
3 H 0.140      
4 H 0.163      
5 O -0.685      
6 H 0.405      
7 C 0.158      
8 H 0.121      
9 C -0.277      
10 H 0.149      
11 C -0.459      
12 H 0.151      
13 H 0.151      
14 H 0.152      
15 C -0.287      
16 H 0.141      
17 H 0.145      
18 H 0.158      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 9.846 -0.060 0.049
y -0.060 7.973 -0.074
z 0.049 -0.074 7.530


<r2> (average value of r2) Å2
<r2> 239.602
(<r2>)1/2 15.479