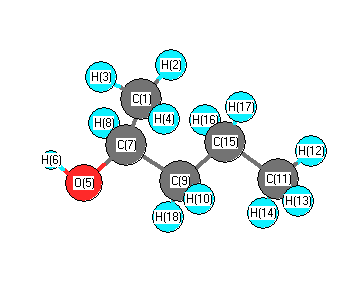
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3801 |
3614 |
9.53 |
|
|
|
| 2 |
A |
3158 |
3003 |
26.89 |
|
|
|
| 3 |
A |
3148 |
2993 |
34.00 |
|
|
|
| 4 |
A |
3142 |
2988 |
57.93 |
|
|
|
| 5 |
A |
3136 |
2982 |
30.65 |
|
|
|
| 6 |
A |
3113 |
2960 |
32.63 |
|
|
|
| 7 |
A |
3086 |
2935 |
3.17 |
|
|
|
| 8 |
A |
3066 |
2916 |
41.49 |
|
|
|
| 9 |
A |
3062 |
2912 |
18.40 |
|
|
|
| 10 |
A |
3061 |
2911 |
37.85 |
|
|
|
| 11 |
A |
3050 |
2901 |
8.26 |
|
|
|
| 12 |
A |
2983 |
2837 |
58.27 |
|
|
|
| 13 |
A |
1538 |
1463 |
6.17 |
|
|
|
| 14 |
A |
1529 |
1454 |
2.65 |
|
|
|
| 15 |
A |
1528 |
1453 |
4.22 |
|
|
|
| 16 |
A |
1526 |
1451 |
8.47 |
|
|
|
| 17 |
A |
1521 |
1446 |
1.40 |
|
|
|
| 18 |
A |
1509 |
1435 |
0.46 |
|
|
|
| 19 |
A |
1456 |
1384 |
20.84 |
|
|
|
| 20 |
A |
1440 |
1370 |
3.01 |
|
|
|
| 21 |
A |
1436 |
1366 |
23.20 |
|
|
|
| 22 |
A |
1413 |
1344 |
8.54 |
|
|
|
| 23 |
A |
1395 |
1327 |
4.05 |
|
|
|
| 24 |
A |
1344 |
1278 |
1.38 |
|
|
|
| 25 |
A |
1328 |
1263 |
24.99 |
|
|
|
| 26 |
A |
1300 |
1236 |
5.34 |
|
|
|
| 27 |
A |
1261 |
1199 |
15.82 |
|
|
|
| 28 |
A |
1202 |
1143 |
16.32 |
|
|
|
| 29 |
A |
1172 |
1114 |
36.00 |
|
|
|
| 30 |
A |
1120 |
1065 |
48.34 |
|
|
|
| 31 |
A |
1115 |
1061 |
1.32 |
|
|
|
| 32 |
A |
1069 |
1017 |
2.49 |
|
|
|
| 33 |
A |
1019 |
969 |
4.65 |
|
|
|
| 34 |
A |
983 |
935 |
7.54 |
|
|
|
| 35 |
A |
932 |
886 |
19.01 |
|
|
|
| 36 |
A |
887 |
843 |
0.79 |
|
|
|
| 37 |
A |
854 |
812 |
0.29 |
|
|
|
| 38 |
A |
755 |
718 |
3.45 |
|
|
|
| 39 |
A |
495 |
470 |
8.63 |
|
|
|
| 40 |
A |
453 |
430 |
1.18 |
|
|
|
| 41 |
A |
416 |
396 |
9.63 |
|
|
|
| 42 |
A |
323 |
308 |
1.06 |
|
|
|
| 43 |
A |
313 |
298 |
120.59 |
|
|
|
| 44 |
A |
270 |
257 |
1.18 |
|
|
|
| 45 |
A |
251 |
238 |
0.57 |
|
|
|
| 46 |
A |
185 |
176 |
1.62 |
|
|
|
| 47 |
A |
117 |
111 |
0.01 |
|
|
|
| 48 |
A |
83 |
79 |
0.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36668.5 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 34871.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.536 |
|
|
|
| 2 |
H |
0.172 |
|
|
|
| 3 |
H |
0.157 |
|
|
|
| 4 |
H |
0.179 |
|
|
|
| 5 |
O |
-0.638 |
|
|
|
| 6 |
H |
0.401 |
|
|
|
| 7 |
C |
0.107 |
|
|
|
| 8 |
H |
0.135 |
|
|
|
| 9 |
C |
-0.309 |
|
|
|
| 10 |
H |
0.165 |
|
|
|
| 11 |
C |
-0.512 |
|
|
|
| 12 |
H |
0.168 |
|
|
|
| 13 |
H |
0.169 |
|
|
|
| 14 |
H |
0.169 |
|
|
|
| 15 |
C |
-0.322 |
|
|
|
| 16 |
H |
0.160 |
|
|
|
| 17 |
H |
0.162 |
|
|
|
| 18 |
H |
0.174 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.417 |
1.295 |
-0.716 |
1.537 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.262 |
-1.737 |
1.770 |
| y |
-1.737 |
-41.893 |
0.759 |
| z |
1.770 |
0.759 |
-39.360 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.365 |
-1.737 |
1.770 |
| y |
-1.737 |
-4.582 |
0.759 |
| z |
1.770 |
0.759 |
-0.783 |
|
| Polar |
|---|
| 3z2-r2 | -1.566 |
|---|
| x2-y2 | 6.631 |
|---|
| xy | -1.737 |
|---|
| xz | 1.770 |
|---|
| yz | 0.759 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.980 |
-0.094 |
0.046 |
| y |
-0.094 |
8.072 |
-0.075 |
| z |
0.046 |
-0.075 |
7.619 |
<r2> (average value of r
2) Å
2
| <r2> |
238.008 |
| (<r2>)1/2 |
15.428 |