Jump to
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -190.576777 |
| Energy at 298.15K | |
| HF Energy | -190.576777 |
| Nuclear repulsion energy | 88.165386 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3102 |
2968 |
31.06 |
|
|
|
| 2 |
A1 |
2355 |
2253 |
849.73 |
|
|
|
| 3 |
A1 |
1820 |
1741 |
24.59 |
|
|
|
| 4 |
A1 |
1499 |
1434 |
4.69 |
|
|
|
| 5 |
A1 |
941 |
900 |
6.62 |
|
|
|
| 6 |
B1 |
1012 |
968 |
25.65 |
|
|
|
| 7 |
B1 |
653 |
625 |
21.53 |
|
|
|
| 8 |
B1 |
236 |
226 |
0.02 |
|
|
|
| 9 |
B2 |
3170 |
3032 |
13.90 |
|
|
|
| 10 |
B2 |
1073 |
1027 |
1.02 |
|
|
|
| 11 |
B2 |
447 |
428 |
15.29 |
|
|
|
| 12 |
B2 |
79i |
76i |
15.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8113.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 7762.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.876 |
| C2 |
0.000 |
0.000 |
-0.558 |
| C3 |
0.000 |
0.000 |
0.720 |
| O4 |
0.000 |
0.000 |
1.901 |
| H5 |
0.000 |
0.924 |
-2.460 |
| H6 |
0.000 |
-0.924 |
-2.460 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3180 | 2.5966 | 3.7775 | 1.0932 | 1.0932 |
C2 | 1.3180 | | 1.2785 | 2.4595 | 2.1143 | 2.1143 | C3 | 2.5966 | 1.2785 | | 1.1810 | 3.3117 | 3.3117 | O4 | 3.7775 | 2.4595 | 1.1810 | | 4.4579 | 4.4579 | H5 | 1.0932 | 2.1143 | 3.3117 | 4.4579 | | 1.8490 | H6 | 1.0932 | 2.1143 | 3.3117 | 4.4579 | 1.8490 | |
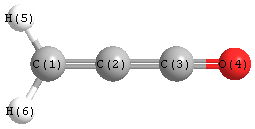 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
122.256 |
| C2 |
C1 |
H6 |
122.256 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
115.488 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.472 |
|
|
|
| 2 |
C |
0.299 |
|
|
|
| 3 |
C |
0.164 |
|
|
|
| 4 |
O |
-0.396 |
|
|
|
| 5 |
H |
0.203 |
|
|
|
| 6 |
H |
0.203 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.714 |
2.714 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.117 |
0.000 |
0.000 |
| y |
0.000 |
-21.903 |
0.000 |
| z |
0.000 |
0.000 |
-21.299 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.516 |
0.000 |
0.000 |
| y |
0.000 |
-0.195 |
0.000 |
| z |
0.000 |
0.000 |
0.711 |
|
| Polar |
|---|
| 3z2-r2 | 1.421 |
|---|
| x2-y2 | -0.214 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.863 |
0.000 |
0.000 |
| y |
0.000 |
2.733 |
0.000 |
| z |
0.000 |
0.000 |
10.151 |
<r2> (average value of r
2) Å
2
| <r2> |
82.433 |
| (<r2>)1/2 |
9.079 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -190.577223 |
| Energy at 298.15K | |
| HF Energy | -190.577223 |
| Nuclear repulsion energy | 88.319368 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3186 |
3048 |
13.35 |
|
|
|
| 2 |
A' |
3110 |
2976 |
29.40 |
|
|
|
| 3 |
A' |
2275 |
2176 |
851.42 |
|
|
|
| 4 |
A' |
1796 |
1718 |
10.18 |
|
|
|
| 5 |
A' |
1503 |
1438 |
1.93 |
|
|
|
| 6 |
A' |
1075 |
1028 |
11.97 |
|
|
|
| 7 |
A' |
960 |
918 |
4.08 |
|
|
|
| 8 |
A' |
482 |
461 |
14.67 |
|
|
|
| 9 |
A' |
139 |
133 |
19.61 |
|
|
|
| 10 |
A" |
1018 |
974 |
25.57 |
|
|
|
| 11 |
A" |
691 |
661 |
16.45 |
|
|
|
| 12 |
A" |
263 |
252 |
1.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8247.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 7890.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.907 |
-1.598 |
0.000 |
| C2 |
0.000 |
-0.631 |
0.000 |
| C3 |
-0.304 |
0.630 |
0.000 |
| O4 |
-0.777 |
1.707 |
0.000 |
| H5 |
1.983 |
-1.407 |
0.000 |
| H6 |
0.617 |
-2.650 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3255 | 2.5360 | 3.7095 | 1.0932 | 1.0912 |
C2 | 1.3255 | | 1.2976 | 2.4642 | 2.1296 | 2.1110 | C3 | 2.5360 | 1.2976 | | 1.1763 | 3.0629 | 3.4071 | O4 | 3.7095 | 2.4642 | 1.1763 | | 4.1618 | 4.5748 | H5 | 1.0932 | 2.1296 | 3.0629 | 4.1618 | | 1.8471 | H6 | 1.0912 | 2.1110 | 3.4071 | 4.5748 | 1.8471 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
150.366 |
|
C2 |
C1 |
H5 |
123.111 |
| C2 |
C1 |
H6 |
121.426 |
|
C2 |
C3 |
O4 |
169.776 |
| H5 |
C1 |
H6 |
115.463 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.392 |
|
|
|
| 2 |
C |
0.024 |
|
|
|
| 3 |
C |
0.344 |
|
|
|
| 4 |
O |
-0.376 |
|
|
|
| 5 |
H |
0.200 |
|
|
|
| 6 |
H |
0.199 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.711 |
-1.850 |
0.000 |
2.520 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.422 |
-0.829 |
0.000 |
| y |
-0.829 |
-22.060 |
0.000 |
| z |
0.000 |
0.000 |
-22.210 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.713 |
-0.829 |
0.000 |
| y |
-0.829 |
-0.243 |
0.000 |
| z |
0.000 |
0.000 |
-0.469 |
|
| Polar |
|---|
| 3z2-r2 | -0.939 |
|---|
| x2-y2 | 0.638 |
|---|
| xy | -0.829 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.391 |
-2.648 |
0.000 |
| y |
-2.648 |
8.517 |
0.000 |
| z |
0.000 |
0.000 |
1.915 |
<r2> (average value of r
2) Å
2
| <r2> |
80.726 |
| (<r2>)1/2 |
8.985 |
