Jump to
S1C2
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -190.102789 |
| Energy at 298.15K | |
| HF Energy | -189.532732 |
| Nuclear repulsion energy | 87.756299 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3133 |
2984 |
25.33 |
|
|
|
| 2 |
A1 |
2326 |
2215 |
761.69 |
|
|
|
| 3 |
A1 |
1790 |
1705 |
33.69 |
|
|
|
| 4 |
A1 |
1513 |
1440 |
6.12 |
|
|
|
| 5 |
A1 |
930 |
885 |
4.11 |
|
|
|
| 6 |
B1 |
1009 |
961 |
25.38 |
|
|
|
| 7 |
B1 |
629 |
599 |
22.83 |
|
|
|
| 8 |
B1 |
220 |
209 |
0.06 |
|
|
|
| 9 |
B2 |
3206 |
3053 |
12.17 |
|
|
|
| 10 |
B2 |
1086 |
1034 |
1.54 |
|
|
|
| 11 |
B2 |
421 |
401 |
14.01 |
|
|
|
| 12 |
B2 |
138i |
131i |
17.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8061.7 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 7677.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.886 |
| C2 |
0.000 |
0.000 |
-0.562 |
| C3 |
0.000 |
0.000 |
0.722 |
| O4 |
0.000 |
0.000 |
1.911 |
| H5 |
0.000 |
0.925 |
-2.466 |
| H6 |
0.000 |
-0.925 |
-2.466 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3244 | 2.6076 | 3.7972 | 1.0921 | 1.0921 |
C2 | 1.3244 | | 1.2832 | 2.4727 | 2.1173 | 2.1173 | C3 | 2.6076 | 1.2832 | | 1.1895 | 3.3191 | 3.3191 | O4 | 3.7972 | 2.4727 | 1.1895 | | 4.4738 | 4.4738 | H5 | 1.0921 | 2.1173 | 3.3191 | 4.4738 | | 1.8507 | H6 | 1.0921 | 2.1173 | 3.3191 | 4.4738 | 1.8507 | |
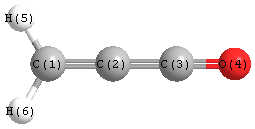 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
122.076 |
| C2 |
C1 |
H6 |
122.076 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
115.848 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -190.104232 |
| Energy at 298.15K | |
| HF Energy | -189.531488 |
| Nuclear repulsion energy | 87.995065 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3222 |
3068 |
11.37 |
|
|
|
| 2 |
A' |
3142 |
2993 |
23.04 |
|
|
|
| 3 |
A' |
2199 |
2094 |
776.63 |
|
|
|
| 4 |
A' |
1764 |
1680 |
10.68 |
|
|
|
| 5 |
A' |
1514 |
1442 |
2.85 |
|
|
|
| 6 |
A' |
1089 |
1037 |
23.91 |
|
|
|
| 7 |
A' |
951 |
906 |
1.18 |
|
|
|
| 8 |
A' |
480 |
457 |
12.63 |
|
|
|
| 9 |
A' |
179 |
170 |
21.69 |
|
|
|
| 10 |
A" |
1004 |
956 |
27.33 |
|
|
|
| 11 |
A" |
697 |
664 |
13.60 |
|
|
|
| 12 |
A" |
259 |
247 |
3.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8250.0 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 7856.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.040 |
-1.503 |
0.000 |
| C2 |
0.000 |
-0.667 |
0.000 |
| C3 |
-0.338 |
0.603 |
0.000 |
| O4 |
-0.898 |
1.644 |
0.000 |
| H5 |
2.078 |
-1.166 |
0.000 |
| H6 |
0.891 |
-2.583 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3341 | 2.5172 | 3.6956 | 1.0920 | 1.0902 |
C2 | 1.3341 | | 1.3150 | 2.4793 | 2.1375 | 2.1127 | C3 | 2.5172 | 1.3150 | | 1.1813 | 2.9951 | 3.4152 | O4 | 3.6956 | 2.4793 | 1.1813 | | 4.0930 | 4.5897 | H5 | 1.0920 | 2.1375 | 2.9951 | 4.0930 | | 1.8490 | H6 | 1.0902 | 2.1127 | 3.4152 | 4.5897 | 1.8490 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
143.689 |
|
C2 |
C1 |
H5 |
123.222 |
| C2 |
C1 |
H6 |
120.934 |
|
C2 |
C3 |
O4 |
166.619 |
| H5 |
C1 |
H6 |
115.844 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
