Jump to
S1C2
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -190.463582 |
| Energy at 298.15K | |
| HF Energy | -190.276177 |
| Nuclear repulsion energy | 87.941864 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3127 |
2968 |
30.31 |
192.61 |
0.11 |
0.20 |
| 2 |
A1 |
2328 |
2210 |
763.94 |
28.95 |
0.34 |
0.51 |
| 3 |
A1 |
1793 |
1702 |
19.12 |
3.57 |
0.41 |
0.58 |
| 4 |
A1 |
1521 |
1443 |
6.57 |
18.43 |
0.51 |
0.68 |
| 5 |
A1 |
930 |
883 |
5.95 |
40.98 |
0.28 |
0.44 |
| 6 |
B1 |
1020 |
968 |
22.63 |
0.04 |
0.75 |
0.86 |
| 7 |
B1 |
643 |
611 |
19.52 |
2.00 |
0.75 |
0.86 |
| 8 |
B1 |
234 |
222 |
0.02 |
4.29 |
0.75 |
0.86 |
| 9 |
B2 |
3195 |
3033 |
14.46 |
126.82 |
0.75 |
0.86 |
| 10 |
B2 |
1089 |
1033 |
1.31 |
0.36 |
0.75 |
0.86 |
| 11 |
B2 |
439 |
417 |
13.59 |
1.57 |
0.75 |
0.86 |
| 12 |
B2 |
119i |
113i |
15.63 |
0.14 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8099.7 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 7688.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.881 |
| C2 |
0.000 |
0.000 |
-0.560 |
| C3 |
0.000 |
0.000 |
0.719 |
| O4 |
0.000 |
0.000 |
1.907 |
| H5 |
0.000 |
0.922 |
-2.464 |
| H6 |
0.000 |
-0.922 |
-2.464 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3213 | 2.6006 | 3.7884 | 1.0907 | 1.0907 |
C2 | 1.3213 | | 1.2793 | 2.4671 | 2.1156 | 2.1156 | C3 | 2.6006 | 1.2793 | | 1.1878 | 3.3142 | 3.3142 | O4 | 3.7884 | 2.4671 | 1.1878 | | 4.4674 | 4.4674 | H5 | 1.0907 | 2.1156 | 3.3142 | 4.4674 | | 1.8438 | H6 | 1.0907 | 2.1156 | 3.3142 | 4.4674 | 1.8438 | |
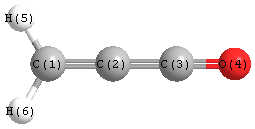 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
122.302 |
| C2 |
C1 |
H6 |
122.302 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
115.395 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.419 |
|
|
|
| 2 |
C |
0.256 |
|
|
|
| 3 |
C |
0.160 |
|
|
|
| 4 |
O |
-0.370 |
|
|
|
| 5 |
H |
0.186 |
|
|
|
| 6 |
H |
0.186 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.690 |
2.690 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.237 |
0.000 |
0.000 |
| y |
0.000 |
-22.138 |
0.000 |
| z |
0.000 |
0.000 |
-21.657 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.339 |
0.000 |
0.000 |
| y |
0.000 |
-0.191 |
0.000 |
| z |
0.000 |
0.000 |
0.530 |
|
| Polar |
|---|
| 3z2-r2 | 1.060 |
|---|
| x2-y2 | -0.098 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.843 |
0.000 |
0.000 |
| y |
0.000 |
2.721 |
0.000 |
| z |
0.000 |
0.000 |
10.255 |
<r2> (average value of r
2) Å
2
| <r2> |
82.899 |
| (<r2>)1/2 |
9.105 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -190.464860 |
| Energy at 298.15K | |
| HF Energy | -190.275677 |
| Nuclear repulsion energy | 88.166815 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3218 |
3054 |
13.21 |
114.34 |
0.66 |
0.80 |
| 2 |
A' |
3139 |
2979 |
27.47 |
143.29 |
0.12 |
0.22 |
| 3 |
A' |
2211 |
2099 |
749.48 |
37.67 |
0.36 |
0.53 |
| 4 |
A' |
1757 |
1668 |
7.17 |
1.32 |
0.75 |
0.86 |
| 5 |
A' |
1523 |
1446 |
1.88 |
19.53 |
0.54 |
0.70 |
| 6 |
A' |
1093 |
1037 |
17.67 |
5.06 |
0.16 |
0.28 |
| 7 |
A' |
954 |
906 |
2.61 |
33.31 |
0.21 |
0.34 |
| 8 |
A' |
492 |
467 |
12.95 |
4.58 |
0.61 |
0.76 |
| 9 |
A' |
173 |
164 |
19.45 |
5.61 |
0.75 |
0.86 |
| 10 |
A" |
1024 |
972 |
23.10 |
0.19 |
0.75 |
0.86 |
| 11 |
A" |
701 |
666 |
12.31 |
2.71 |
0.75 |
0.86 |
| 12 |
A" |
273 |
259 |
3.17 |
6.44 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8278.4 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 7857.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.045 |
-1.493 |
0.000 |
| C2 |
0.000 |
-0.667 |
0.000 |
| C3 |
-0.349 |
0.594 |
0.000 |
| O4 |
-0.896 |
1.640 |
0.000 |
| H5 |
2.081 |
-1.152 |
0.000 |
| H6 |
0.909 |
-2.573 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3325 | 2.5102 | 3.6864 | 1.0905 | 1.0882 |
C2 | 1.3325 | | 1.3086 | 2.4754 | 2.1369 | 2.1116 | C3 | 2.5102 | 1.3086 | | 1.1808 | 2.9922 | 3.4078 | O4 | 3.6864 | 2.4754 | 1.1808 | | 4.0820 | 4.5838 | H5 | 1.0905 | 2.1369 | 2.9922 | 4.0820 | | 1.8419 | H6 | 1.0882 | 2.1116 | 3.4078 | 4.5838 | 1.8419 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
143.778 |
|
C2 |
C1 |
H5 |
123.440 |
| C2 |
C1 |
H6 |
121.131 |
|
C2 |
C3 |
O4 |
167.818 |
| H5 |
C1 |
H6 |
115.429 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.336 |
|
|
|
| 2 |
C |
-0.030 |
|
|
|
| 3 |
C |
0.347 |
|
|
|
| 4 |
O |
-0.345 |
|
|
|
| 5 |
H |
0.182 |
|
|
|
| 6 |
H |
0.182 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.886 |
-1.480 |
0.000 |
2.397 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.507 |
-0.869 |
-0.001 |
| y |
-0.869 |
-22.675 |
0.003 |
| z |
-0.001 |
0.003 |
-22.369 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.016 |
-0.869 |
-0.001 |
| y |
-0.869 |
-0.737 |
0.003 |
| z |
-0.001 |
0.003 |
-0.279 |
|
| Polar |
|---|
| 3z2-r2 | -0.557 |
|---|
| x2-y2 | 1.168 |
|---|
| xy | -0.869 |
|---|
| xz | -0.001 |
|---|
| yz | 0.003 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.913 |
-2.861 |
0.000 |
| y |
-2.861 |
8.104 |
0.000 |
| z |
0.000 |
0.000 |
1.919 |
<r2> (average value of r
2) Å
2
| <r2> |
80.367 |
| (<r2>)1/2 |
8.965 |
