return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6 (Methylenecyclopropane)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-155.437980
Energy at 298.15K-155.443761
HF Energy-154.885961
Nuclear repulsion energy108.951502
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3167 3016 13.00      
2 A1 3153 3003 10.47      
3 A1 1874 1785 1.13      
4 A1 1534 1460 0.05      
5 A1 1485 1414 0.16      
6 A1 1098 1046 0.00      
7 A1 1081 1029 7.17      
8 A1 762 726 5.36      
9 A2 3224 3070 0.00      
10 A2 1202 1145 0.00      
11 A2 980 933 0.00      
12 A2 633 602 0.00      
13 B1 3236 3082 20.55      
14 B1 1129 1075 0.99      
15 B1 908 865 39.19      
16 B1 782 744 0.64      
17 B1 285 271 3.12      
18 B2 3248 3093 14.15      
19 B2 3150 2999 15.20      
20 B2 1499 1428 0.65      
21 B2 1178 1122 7.56      
22 B2 1112 1059 3.32      
23 B2 932 888 11.15      
24 B2 359 342 0.26      

Unscaled Zero Point Vibrational Energy (zpe) 19004.4 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 18097.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.65057 0.22713 0.18052

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.655
C2 0.000 0.000 0.307
H3 0.000 0.896 2.175
H4 0.000 -0.896 2.175
C5 0.000 0.747 -0.894
C6 0.000 -0.747 -0.894
H7 0.919 0.926 -1.348
H8 -0.919 0.926 -1.348
H9 -0.919 -0.926 -1.348
H10 0.919 -0.926 -1.348

Atom - Atom Distances (Å)
  C1 C2 H3 H4 C5 C6 H7 H8 H9 H10
C11.34791.03601.03602.65622.65623.27433.27433.27433.2743
C21.34792.07192.07191.41431.41432.10752.10752.10752.1075
H31.03602.07191.79193.07313.48133.64143.64144.07174.0717
H41.03602.07191.79193.48133.07314.07174.07173.64143.6414
C52.65621.41433.07313.48131.49311.04031.04031.96151.9615
C62.65621.41433.48133.07311.49311.96151.96151.04031.0403
H73.27432.10753.64144.07171.04031.96151.83732.60871.8520
H83.27432.10753.64144.07171.04031.96151.83731.85202.6087
H93.27432.10754.07173.64141.96151.04032.60871.85201.8373
H103.27432.10754.07173.64141.96151.04031.85202.60871.8373

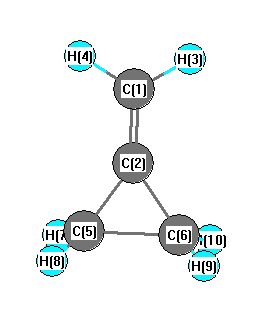
picture of Methylenecyclopropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C5 148.141 C1 C2 C6 148.141
C2 C1 H3 120.145 C2 C1 H4 120.145
C2 C5 C6 58.141 C2 C5 H7 117.502
C2 C5 H8 117.502 C2 C6 C5 58.141
C2 C6 H9 117.502 C2 C6 H10 117.502
H3 C1 H4 119.711 C5 C2 C6 63.718
C5 C6 H9 99.935 C5 C6 H10 99.935
C6 C5 H7 99.935 C6 C5 H8 99.935
H7 C5 H8 124.018 H9 C6 H10 124.018
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability