return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 ((E)-3-methylpent-2-ene)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-235.536368
Energy at 298.15K-235.548399
HF Energy-235.277720
Nuclear repulsion energy236.727869
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3184 3023 41.78      
2 A 3172 3011 34.21      
3 A 3160 2999 33.63      
4 A 3159 2998 6.88      
5 A 3154 2993 37.89      
6 A 3120 2962 29.44      
7 A 3116 2958 23.00      
8 A 3106 2948 15.01      
9 A 3081 2925 27.77      
10 A 3070 2914 12.99      
11 A 3067 2911 48.45      
12 A 3050 2895 34.23      
13 A 1758 1669 1.13      
14 A 1559 1479 2.30      
15 A 1554 1475 8.97      
16 A 1549 1470 2.63      
17 A 1540 1462 10.94      
18 A 1539 1461 3.77      
19 A 1534 1456 2.55      
20 A 1526 1449 0.55      
21 A 1469 1394 3.94      
22 A 1462 1388 0.78      
23 A 1453 1379 1.07      
24 A 1414 1342 1.80      
25 A 1391 1320 5.73      
26 A 1317 1250 0.15      
27 A 1261 1197 3.95      
28 A 1168 1109 5.09      
29 A 1123 1066 7.12      
30 A 1113 1057 1.65      
31 A 1086 1031 0.72      
32 A 1071 1017 0.44      
33 A 1036 984 3.20      
34 A 1012 961 6.44      
35 A 949 900 0.90      
36 A 862 818 11.45      
37 A 805 764 1.68      
38 A 764 726 1.59      
39 A 557 528 3.90      
40 A 495 470 2.44      
41 A 406 386 0.39      
42 A 320 304 0.43      
43 A 315 299 0.38      
44 A 241 229 0.09      
45 A 192 182 0.77      
46 A 127 120 0.12      
47 A 111 105 0.69      
48 A 60 57 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 36788.1 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 34919.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.20666 0.06724 0.05696

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.650 -0.128 0.002
H2 3.216 -0.495 -0.847
H3 3.189 -0.473 0.884
H4 2.694 0.962 -0.024
C5 -0.052 1.491 -0.003
H6 0.920 1.984 -0.016
H7 -0.580 1.851 0.897
H8 -0.611 1.841 -0.862
C9 -2.517 -0.067 -0.023
H10 -2.661 0.589 -0.863
H11 -2.686 0.544 0.893
C12 -1.227 -0.840 0.015
H13 -1.215 -1.486 0.892
H14 -1.190 -1.511 -0.840
C15 1.253 -0.655 -0.022
H16 1.187 -1.755 -0.026
C17 0.077 -0.013 0.020
H18 -3.370 -0.776 -0.024

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 H10 H11 C12 H13 H14 C15 H16 C17 H18
C11.08461.08921.09063.14972.73023.89183.90655.16715.42895.45173.94184.19234.16781.49322.18872.57576.0550
H21.08461.73121.75263.91603.47954.79064.48385.80725.97636.24054.53884.86224.52182.13512.52593.29206.6431
H31.08921.73121.76813.89163.46334.42744.77965.79166.19735.96274.51554.51954.81972.14532.54633.26216.6287
H41.09061.75261.76812.79612.04733.51473.52115.31085.43305.47334.31474.70174.67572.16503.10662.79266.3076
C53.14973.91603.89162.79611.08991.10311.08392.91592.89192.93892.61043.31833.31772.51123.47411.50984.0185
H62.73023.47953.46332.04731.08991.76091.75514.00223.93563.98793.54774.17394.16482.65973.74862.16835.1011
H73.89184.79064.42743.51471.10311.76091.75942.87683.00372.47852.90453.39613.83253.23734.11972.16183.9411
H83.90654.48384.77963.52111.08391.75511.75942.82422.40233.01132.88783.80893.40163.22654.10652.16613.8939
C95.16715.80725.79165.31082.91594.00222.87682.82421.07541.11471.50432.13122.12373.81524.06962.59451.1093
H105.42895.97636.19735.43302.89193.93563.00372.40231.07541.75682.20733.07792.56384.19244.58262.93971.7518
H115.45176.24055.96275.47332.93893.98792.47853.01131.11471.75682.19492.50683.07664.21824.59682.95091.7470
C123.94184.53884.51554.31472.61043.54772.90452.88781.50432.20732.19491.08831.08762.48702.58121.54392.1446
H134.19234.86224.51954.70173.31834.17393.39613.80892.13123.07792.50681.08831.73222.76022.58542.14392.4465
H144.16784.52184.81974.67573.31774.16483.83253.40162.12372.56383.07661.08761.73222.71512.52412.14222.4408
C151.49322.13512.14532.16502.51122.65973.23733.22653.81524.19244.21822.48702.76022.71511.10231.34034.6247
H162.18872.52592.54633.10663.47413.74864.11974.10654.06964.58264.59682.58122.58542.52411.10232.06564.6606
C172.57573.29203.26212.79261.50982.16832.16182.16612.59452.93972.95091.54392.14392.14221.34032.06563.5307
H186.05506.64316.62876.30764.01855.10113.94113.89391.10931.75181.74702.14462.44652.44084.62474.66063.5307

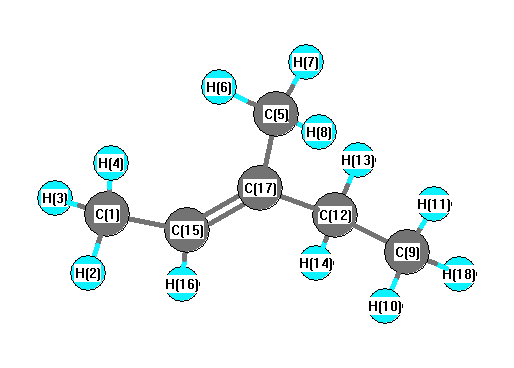
picture of (E)-3-methylpent-2-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C15 H16 114.123 C1 C15 C17 130.661
H2 C1 H3 105.573 H2 C1 H4 107.356
H2 C1 C15 110.842 H3 C1 H4 108.417
H3 C1 C15 111.389 H4 C1 C15 112.913
C5 C17 C12 117.481 C5 C17 C15 123.445
H6 C5 H7 106.820 H6 C5 H8 107.685
H6 C5 C17 112.024 H7 C5 H8 107.115
H7 C5 C17 110.691 H8 C5 C17 112.221
C9 C12 H13 109.541 C9 C12 H14 108.981
C9 C12 C17 116.676 H10 C9 H11 106.656
H10 C9 C12 116.682 H10 C9 H18 106.600
H11 C9 C12 113.032 H11 C9 H18 103.534
C12 C9 H18 109.356 C12 C17 C15 118.987
H13 C12 H14 105.518 H13 C12 C17 107.820
H14 C12 C17 107.726 H16 C15 C17 115.138
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.500      
2 H 0.160      
3 H 0.160      
4 H 0.159      
5 C -0.522      
6 H 0.162      
7 H 0.157      
8 H 0.164      
9 C -0.461      
10 H 0.155      
11 H 0.152      
12 C -0.329      
13 H 0.145      
14 H 0.150      
15 C -0.184      
16 H 0.135      
17 C 0.145      
18 H 0.151      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 11.885 1.007 -0.452
y 1.007 9.351 0.001
z -0.452 0.001 7.181


<r2> (average value of r2) Å2
<r2> 223.373
(<r2>)1/2 14.946