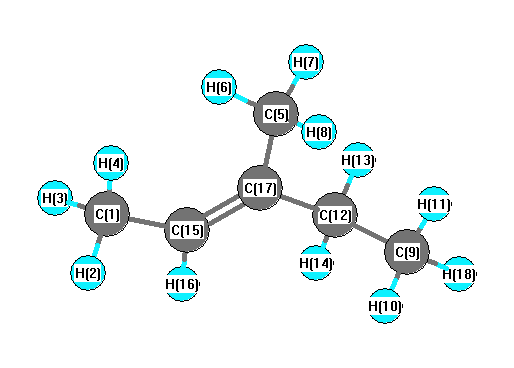
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3174 |
3018 |
40.54 |
|
|
|
| 2 |
A |
3166 |
3011 |
32.58 |
|
|
|
| 3 |
A |
3152 |
2998 |
31.36 |
|
|
|
| 4 |
A |
3151 |
2996 |
9.85 |
|
|
|
| 5 |
A |
3149 |
2995 |
30.81 |
|
|
|
| 6 |
A |
3111 |
2958 |
26.77 |
|
|
|
| 7 |
A |
3108 |
2955 |
19.40 |
|
|
|
| 8 |
A |
3101 |
2949 |
14.05 |
|
|
|
| 9 |
A |
3071 |
2920 |
28.86 |
|
|
|
| 10 |
A |
3058 |
2908 |
14.87 |
|
|
|
| 11 |
A |
3054 |
2905 |
49.88 |
|
|
|
| 12 |
A |
3038 |
2890 |
34.77 |
|
|
|
| 13 |
A |
1779 |
1691 |
1.51 |
|
|
|
| 14 |
A |
1536 |
1461 |
3.38 |
|
|
|
| 15 |
A |
1531 |
1456 |
10.81 |
|
|
|
| 16 |
A |
1524 |
1450 |
3.70 |
|
|
|
| 17 |
A |
1515 |
1441 |
11.27 |
|
|
|
| 18 |
A |
1514 |
1440 |
8.11 |
|
|
|
| 19 |
A |
1509 |
1435 |
2.85 |
|
|
|
| 20 |
A |
1500 |
1427 |
0.61 |
|
|
|
| 21 |
A |
1447 |
1377 |
5.36 |
|
|
|
| 22 |
A |
1444 |
1373 |
2.27 |
|
|
|
| 23 |
A |
1431 |
1361 |
2.33 |
|
|
|
| 24 |
A |
1394 |
1326 |
1.67 |
|
|
|
| 25 |
A |
1377 |
1309 |
3.94 |
|
|
|
| 26 |
A |
1301 |
1238 |
0.26 |
|
|
|
| 27 |
A |
1255 |
1194 |
4.47 |
|
|
|
| 28 |
A |
1160 |
1103 |
6.37 |
|
|
|
| 29 |
A |
1113 |
1059 |
4.77 |
|
|
|
| 30 |
A |
1105 |
1051 |
5.03 |
|
|
|
| 31 |
A |
1071 |
1019 |
0.60 |
|
|
|
| 32 |
A |
1059 |
1007 |
0.48 |
|
|
|
| 33 |
A |
1030 |
980 |
4.50 |
|
|
|
| 34 |
A |
1008 |
959 |
8.14 |
|
|
|
| 35 |
A |
947 |
901 |
1.10 |
|
|
|
| 36 |
A |
861 |
819 |
13.19 |
|
|
|
| 37 |
A |
798 |
758 |
2.35 |
|
|
|
| 38 |
A |
769 |
731 |
1.70 |
|
|
|
| 39 |
A |
554 |
527 |
4.74 |
|
|
|
| 40 |
A |
496 |
472 |
2.73 |
|
|
|
| 41 |
A |
404 |
384 |
0.51 |
|
|
|
| 42 |
A |
321 |
305 |
0.56 |
|
|
|
| 43 |
A |
313 |
298 |
0.40 |
|
|
|
| 44 |
A |
241 |
229 |
0.07 |
|
|
|
| 45 |
A |
192 |
182 |
0.85 |
|
|
|
| 46 |
A |
138 |
131 |
0.11 |
|
|
|
| 47 |
A |
117 |
111 |
0.99 |
|
|
|
| 48 |
A |
64 |
61 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36575.0 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 34782.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.561 |
|
|
|
| 2 |
H |
0.179 |
|
|
|
| 3 |
H |
0.179 |
|
|
|
| 4 |
H |
0.175 |
|
|
|
| 5 |
C |
-0.584 |
|
|
|
| 6 |
H |
0.177 |
|
|
|
| 7 |
H |
0.175 |
|
|
|
| 8 |
H |
0.182 |
|
|
|
| 9 |
C |
-0.514 |
|
|
|
| 10 |
H |
0.174 |
|
|
|
| 11 |
H |
0.169 |
|
|
|
| 12 |
C |
-0.378 |
|
|
|
| 13 |
H |
0.163 |
|
|
|
| 14 |
H |
0.167 |
|
|
|
| 15 |
C |
-0.190 |
|
|
|
| 16 |
H |
0.143 |
|
|
|
| 17 |
C |
0.177 |
|
|
|
| 18 |
H |
0.170 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.036 |
0.152 |
0.038 |
0.160 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.355 |
0.233 |
-0.646 |
| y |
0.233 |
-38.344 |
0.071 |
| z |
-0.646 |
0.071 |
-40.507 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.070 |
0.233 |
-0.646 |
| y |
0.233 |
1.087 |
0.071 |
| z |
-0.646 |
0.071 |
-2.157 |
|
| Polar |
|---|
| 3z2-r2 | -4.314 |
|---|
| x2-y2 | -0.011 |
|---|
| xy | 0.233 |
|---|
| xz | -0.646 |
|---|
| yz | 0.071 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.199 |
1.043 |
-0.473 |
| y |
1.043 |
9.483 |
-0.010 |
| z |
-0.473 |
-0.010 |
7.261 |
<r2> (average value of r
2) Å
2
| <r2> |
222.267 |
| (<r2>)1/2 |
14.909 |