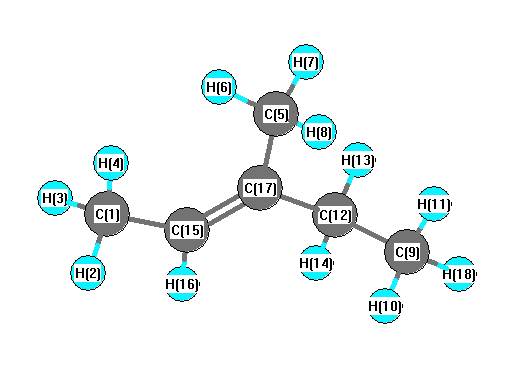
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3080 |
3030 |
37.22 |
|
|
|
| 2 |
A |
3074 |
3023 |
35.82 |
|
|
|
| 3 |
A |
3063 |
3013 |
30.91 |
|
|
|
| 4 |
A |
3060 |
3009 |
37.85 |
|
|
|
| 5 |
A |
3055 |
3004 |
4.60 |
|
|
|
| 6 |
A |
3017 |
2967 |
25.97 |
|
|
|
| 7 |
A |
3013 |
2963 |
20.68 |
|
|
|
| 8 |
A |
3007 |
2957 |
15.74 |
|
|
|
| 9 |
A |
2985 |
2936 |
29.21 |
|
|
|
| 10 |
A |
2968 |
2920 |
17.43 |
|
|
|
| 11 |
A |
2965 |
2916 |
51.93 |
|
|
|
| 12 |
A |
2946 |
2898 |
35.61 |
|
|
|
| 13 |
A |
1703 |
1675 |
1.48 |
|
|
|
| 14 |
A |
1490 |
1466 |
2.96 |
|
|
|
| 15 |
A |
1485 |
1461 |
10.13 |
|
|
|
| 16 |
A |
1478 |
1453 |
2.73 |
|
|
|
| 17 |
A |
1469 |
1445 |
3.75 |
|
|
|
| 18 |
A |
1468 |
1444 |
14.11 |
|
|
|
| 19 |
A |
1463 |
1439 |
2.17 |
|
|
|
| 20 |
A |
1454 |
1430 |
0.40 |
|
|
|
| 21 |
A |
1397 |
1374 |
5.48 |
|
|
|
| 22 |
A |
1394 |
1371 |
1.11 |
|
|
|
| 23 |
A |
1382 |
1359 |
1.30 |
|
|
|
| 24 |
A |
1350 |
1328 |
1.98 |
|
|
|
| 25 |
A |
1324 |
1302 |
4.61 |
|
|
|
| 26 |
A |
1260 |
1239 |
0.26 |
|
|
|
| 27 |
A |
1212 |
1192 |
5.12 |
|
|
|
| 28 |
A |
1122 |
1104 |
7.49 |
|
|
|
| 29 |
A |
1074 |
1056 |
4.76 |
|
|
|
| 30 |
A |
1070 |
1052 |
5.46 |
|
|
|
| 31 |
A |
1031 |
1014 |
0.25 |
|
|
|
| 32 |
A |
1024 |
1007 |
0.23 |
|
|
|
| 33 |
A |
995 |
979 |
5.25 |
|
|
|
| 34 |
A |
976 |
959 |
8.59 |
|
|
|
| 35 |
A |
916 |
901 |
1.16 |
|
|
|
| 36 |
A |
827 |
813 |
12.45 |
|
|
|
| 37 |
A |
773 |
760 |
2.38 |
|
|
|
| 38 |
A |
744 |
732 |
1.86 |
|
|
|
| 39 |
A |
539 |
530 |
4.58 |
|
|
|
| 40 |
A |
481 |
473 |
2.67 |
|
|
|
| 41 |
A |
390 |
384 |
0.56 |
|
|
|
| 42 |
A |
311 |
306 |
0.53 |
|
|
|
| 43 |
A |
304 |
299 |
0.37 |
|
|
|
| 44 |
A |
232 |
228 |
0.11 |
|
|
|
| 45 |
A |
187 |
184 |
0.76 |
|
|
|
| 46 |
A |
141 |
138 |
0.14 |
|
|
|
| 47 |
A |
116 |
114 |
1.12 |
|
|
|
| 48 |
A |
57 |
56 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35435.0 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 34850.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.534 |
|
|
|
| 2 |
H |
0.168 |
|
|
|
| 3 |
H |
0.167 |
|
|
|
| 4 |
H |
0.164 |
|
|
|
| 5 |
C |
-0.566 |
|
|
|
| 6 |
H |
0.165 |
|
|
|
| 7 |
H |
0.164 |
|
|
|
| 8 |
H |
0.170 |
|
|
|
| 9 |
C |
-0.482 |
|
|
|
| 10 |
H |
0.162 |
|
|
|
| 11 |
H |
0.158 |
|
|
|
| 12 |
C |
-0.356 |
|
|
|
| 13 |
H |
0.148 |
|
|
|
| 14 |
H |
0.151 |
|
|
|
| 15 |
C |
-0.167 |
|
|
|
| 16 |
H |
0.121 |
|
|
|
| 17 |
C |
0.209 |
|
|
|
| 18 |
H |
0.157 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.021 |
0.163 |
0.041 |
0.169 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.382 |
0.257 |
-0.630 |
| y |
0.257 |
-38.493 |
0.079 |
| z |
-0.630 |
0.079 |
-40.553 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.141 |
0.257 |
-0.630 |
| y |
0.257 |
0.974 |
0.079 |
| z |
-0.630 |
0.079 |
-2.115 |
|
| Polar |
|---|
| 3z2-r2 | -4.230 |
|---|
| x2-y2 | 0.112 |
|---|
| xy | 0.257 |
|---|
| xz | -0.630 |
|---|
| yz | 0.079 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.706 |
1.027 |
-0.494 |
| y |
1.027 |
9.803 |
0.007 |
| z |
-0.494 |
0.007 |
7.433 |
<r2> (average value of r
2) Å
2
| <r2> |
225.034 |
| (<r2>)1/2 |
15.001 |