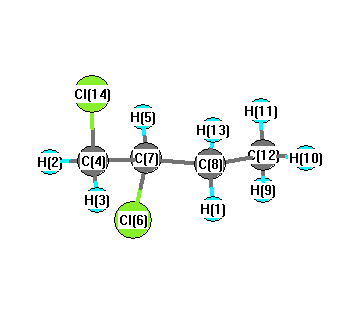
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3206 |
3049 |
2.77 |
|
|
|
| 2 |
A |
3167 |
3012 |
17.32 |
|
|
|
| 3 |
A |
3154 |
3000 |
30.66 |
|
|
|
| 4 |
A |
3139 |
2985 |
12.52 |
|
|
|
| 5 |
A |
3132 |
2978 |
0.74 |
|
|
|
| 6 |
A |
3115 |
2963 |
1.60 |
|
|
|
| 7 |
A |
3077 |
2926 |
25.92 |
|
|
|
| 8 |
A |
3073 |
2922 |
7.35 |
|
|
|
| 9 |
A |
1534 |
1459 |
8.65 |
|
|
|
| 10 |
A |
1525 |
1451 |
8.07 |
|
|
|
| 11 |
A |
1507 |
1434 |
4.74 |
|
|
|
| 12 |
A |
1502 |
1428 |
5.59 |
|
|
|
| 13 |
A |
1443 |
1372 |
4.30 |
|
|
|
| 14 |
A |
1423 |
1354 |
0.62 |
|
|
|
| 15 |
A |
1360 |
1294 |
4.94 |
|
|
|
| 16 |
A |
1348 |
1282 |
4.32 |
|
|
|
| 17 |
A |
1320 |
1255 |
9.26 |
|
|
|
| 18 |
A |
1268 |
1206 |
11.81 |
|
|
|
| 19 |
A |
1227 |
1167 |
11.31 |
|
|
|
| 20 |
A |
1151 |
1095 |
1.26 |
|
|
|
| 21 |
A |
1120 |
1065 |
1.81 |
|
|
|
| 22 |
A |
1087 |
1034 |
3.73 |
|
|
|
| 23 |
A |
1062 |
1010 |
0.95 |
|
|
|
| 24 |
A |
964 |
917 |
11.84 |
|
|
|
| 25 |
A |
853 |
811 |
8.49 |
|
|
|
| 26 |
A |
827 |
786 |
7.95 |
|
|
|
| 27 |
A |
763 |
725 |
57.76 |
|
|
|
| 28 |
A |
680 |
647 |
37.97 |
|
|
|
| 29 |
A |
467 |
444 |
2.01 |
|
|
|
| 30 |
A |
391 |
372 |
1.61 |
|
|
|
| 31 |
A |
300 |
286 |
0.06 |
|
|
|
| 32 |
A |
244 |
232 |
0.59 |
|
|
|
| 33 |
A |
222 |
211 |
2.80 |
|
|
|
| 34 |
A |
192 |
183 |
3.83 |
|
|
|
| 35 |
A |
119 |
113 |
3.80 |
|
|
|
| 36 |
A |
97 |
92 |
1.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25530.1 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 24279.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.184 |
|
|
|
| 2 |
H |
0.243 |
|
|
|
| 3 |
H |
0.233 |
|
|
|
| 4 |
C |
-0.423 |
|
|
|
| 5 |
H |
0.231 |
|
|
|
| 6 |
Cl |
-0.064 |
|
|
|
| 7 |
C |
-0.262 |
|
|
|
| 8 |
C |
-0.312 |
|
|
|
| 9 |
H |
0.195 |
|
|
|
| 10 |
H |
0.180 |
|
|
|
| 11 |
H |
0.174 |
|
|
|
| 12 |
C |
-0.521 |
|
|
|
| 13 |
H |
0.203 |
|
|
|
| 14 |
Cl |
-0.061 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.321 |
0.636 |
0.232 |
0.749 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-57.283 |
3.442 |
-0.264 |
| y |
3.442 |
-50.634 |
-0.195 |
| z |
-0.264 |
-0.195 |
-50.188 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.873 |
3.442 |
-0.264 |
| y |
3.442 |
3.102 |
-0.195 |
| z |
-0.264 |
-0.195 |
3.771 |
|
| Polar |
|---|
| 3z2-r2 | 7.542 |
|---|
| x2-y2 | -6.649 |
|---|
| xy | 3.442 |
|---|
| xz | -0.264 |
|---|
| yz | -0.195 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.079 |
-1.088 |
0.237 |
| y |
-1.088 |
9.407 |
-0.158 |
| z |
0.237 |
-0.158 |
7.114 |
<r2> (average value of r
2) Å
2
| <r2> |
321.171 |
| (<r2>)1/2 |
17.921 |